Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -1072.934601 |
| Energy at 298.15K | -1072.937099 |
| HF Energy | -1072.934601 |
| Nuclear repulsion energy | 263.214489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3176 |
3058 |
1.75 |
|
|
|
| 2 |
A' |
2970 |
2860 |
53.20 |
|
|
|
| 3 |
A' |
1851 |
1782 |
182.05 |
|
|
|
| 4 |
A' |
1412 |
1359 |
4.63 |
|
|
|
| 5 |
A' |
1230 |
1184 |
11.17 |
|
|
|
| 6 |
A' |
1062 |
1023 |
31.75 |
|
|
|
| 7 |
A' |
789 |
759 |
24.06 |
|
|
|
| 8 |
A' |
439 |
423 |
5.27 |
|
|
|
| 9 |
A' |
323 |
311 |
23.79 |
|
|
|
| 10 |
A' |
258 |
248 |
2.46 |
|
|
|
| 11 |
A" |
1273 |
1226 |
22.69 |
|
|
|
| 12 |
A" |
1021 |
983 |
21.51 |
|
|
|
| 13 |
A" |
722 |
695 |
144.61 |
|
|
|
| 14 |
A" |
278 |
267 |
4.03 |
|
|
|
| 15 |
A" |
90 |
86 |
7.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8446.9 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 8131.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.386 |
-0.019 |
0.000 |
| C2 |
-0.171 |
1.400 |
0.000 |
| H3 |
1.472 |
-0.006 |
0.000 |
| Cl4 |
-0.171 |
-0.842 |
1.479 |
| Cl5 |
-0.171 |
-0.842 |
-1.479 |
| O6 |
0.539 |
2.362 |
0.000 |
| H7 |
-1.276 |
1.457 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5239 | 1.0866 | 1.7814 | 1.7814 | 2.3859 | 2.2228 |
C2 | 1.5239 | | 2.1618 | 2.6855 | 2.6855 | 1.1955 | 1.1074 | H3 | 1.0866 | 2.1618 | | 2.3632 | 2.3632 | 2.5447 | 3.1134 | Cl4 | 1.7814 | 2.6855 | 2.3632 | | 2.9576 | 3.5992 | 2.9486 | Cl5 | 1.7814 | 2.6855 | 2.3632 | 2.9576 | | 3.5992 | 2.9486 | O6 | 2.3859 | 1.1955 | 2.5447 | 3.5992 | 3.5992 | | 2.0285 | H7 | 2.2228 | 1.1074 | 3.1134 | 2.9486 | 2.9486 | 2.0285 | |
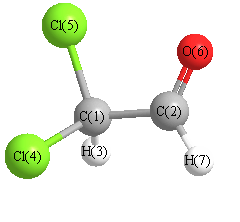 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.185 |
|
C1 |
C2 |
H7 |
114.364 |
| C2 |
C1 |
H3 |
110.695 |
|
C2 |
C1 |
Cl4 |
108.426 |
| C2 |
C1 |
Cl5 |
108.426 |
|
H3 |
C1 |
Cl4 |
108.537 |
| H3 |
C1 |
Cl5 |
108.537 |
|
Cl4 |
C1 |
Cl5 |
112.231 |
| O6 |
C2 |
H7 |
123.452 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.510 |
|
|
|
| 2 |
C |
0.179 |
|
|
|
| 3 |
H |
0.336 |
|
|
|
| 4 |
Cl |
0.017 |
|
|
|
| 5 |
Cl |
0.017 |
|
|
|
| 6 |
O |
-0.249 |
|
|
|
| 7 |
H |
0.211 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.313 |
-0.551 |
0.000 |
0.634 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.090 |
-2.830 |
0.000 |
| y |
-2.830 |
-48.654 |
0.000 |
| z |
0.000 |
0.000 |
-43.205 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.840 |
-2.830 |
0.000 |
| y |
-2.830 |
-6.507 |
0.000 |
| z |
0.000 |
0.000 |
1.667 |
|
| Polar |
|---|
| 3z2-r2 | 3.333 |
|---|
| x2-y2 | 7.564 |
|---|
| xy | -2.830 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.065 |
0.714 |
0.000 |
| y |
0.714 |
7.096 |
0.000 |
| z |
0.000 |
0.000 |
7.622 |
<r2> (average value of r
2) Å
2
| <r2> |
192.816 |
| (<r2>)1/2 |
13.886 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -1072.931547 |
| Energy at 298.15K | |
| HF Energy | -1072.931547 |
| Nuclear repulsion energy | 265.858321 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3117 |
3001 |
6.18 |
|
|
|
| 2 |
A |
2949 |
2839 |
65.70 |
|
|
|
| 3 |
A |
1866 |
1796 |
157.20 |
|
|
|
| 4 |
A |
1406 |
1354 |
16.40 |
|
|
|
| 5 |
A |
1280 |
1232 |
17.99 |
|
|
|
| 6 |
A |
1243 |
1196 |
9.82 |
|
|
|
| 7 |
A |
1031 |
992 |
15.61 |
|
|
|
| 8 |
A |
936 |
901 |
11.43 |
|
|
|
| 9 |
A |
806 |
776 |
88.93 |
|
|
|
| 10 |
A |
645 |
621 |
41.62 |
|
|
|
| 11 |
A |
616 |
593 |
41.36 |
|
|
|
| 12 |
A |
343 |
330 |
2.17 |
|
|
|
| 13 |
A |
278 |
267 |
3.02 |
|
|
|
| 14 |
A |
219 |
211 |
2.46 |
|
|
|
| 15 |
A |
90 |
86 |
10.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8411.2 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 8097.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.096 |
-0.020 |
0.516 |
| C2 |
-0.709 |
-1.284 |
0.201 |
| H3 |
0.228 |
0.068 |
1.596 |
| Cl4 |
1.737 |
-0.253 |
-0.168 |
| Cl5 |
-0.685 |
1.450 |
-0.058 |
| O6 |
-1.780 |
-1.298 |
-0.322 |
| H7 |
-0.186 |
-2.208 |
0.521 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5317 | 1.0910 | 1.7935 | 1.7606 | 2.4201 | 2.2061 |
C2 | 1.5317 | | 2.1568 | 2.6801 | 2.7463 | 1.1919 | 1.1088 | H3 | 1.0910 | 2.1568 | | 2.3434 | 2.3407 | 3.0947 | 2.5501 | Cl4 | 1.7935 | 2.6801 | 2.3434 | | 2.9632 | 3.6723 | 2.8275 | Cl5 | 1.7606 | 2.7463 | 2.3407 | 2.9632 | | 2.9700 | 3.7368 | O6 | 2.4201 | 1.1919 | 3.0947 | 3.6723 | 2.9700 | | 2.0198 | H7 | 2.2061 | 1.1088 | 2.5501 | 2.8275 | 3.7368 | 2.0198 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.916 |
|
C1 |
C2 |
H7 |
112.340 |
| C2 |
C1 |
H3 |
109.490 |
|
C2 |
C1 |
Cl4 |
107.153 |
| C2 |
C1 |
Cl5 |
112.875 |
|
H3 |
C1 |
Cl4 |
106.087 |
| H3 |
C1 |
Cl5 |
108.029 |
|
Cl4 |
C1 |
Cl5 |
112.965 |
| O6 |
C2 |
H7 |
122.743 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.505 |
|
|
|
| 2 |
C |
0.159 |
|
|
|
| 3 |
H |
0.328 |
|
|
|
| 4 |
Cl |
0.005 |
|
|
|
| 5 |
Cl |
0.034 |
|
|
|
| 6 |
O |
-0.230 |
|
|
|
| 7 |
H |
0.209 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.128 |
-0.857 |
2.009 |
2.458 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.855 |
-2.065 |
-1.216 |
| y |
-2.065 |
-41.970 |
-1.578 |
| z |
-1.216 |
-1.578 |
-40.595 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.573 |
-2.065 |
-1.216 |
| y |
-2.065 |
2.255 |
-1.578 |
| z |
-1.216 |
-1.578 |
4.318 |
|
| Polar |
|---|
| 3z2-r2 | 8.636 |
|---|
| x2-y2 | -5.885 |
|---|
| xy | -2.065 |
|---|
| xz | -1.216 |
|---|
| yz | -1.578 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.062 |
-0.847 |
0.138 |
| y |
-0.847 |
6.946 |
-0.401 |
| z |
0.138 |
-0.401 |
4.372 |
<r2> (average value of r
2) Å
2
| <r2> |
186.054 |
| (<r2>)1/2 |
13.640 |
