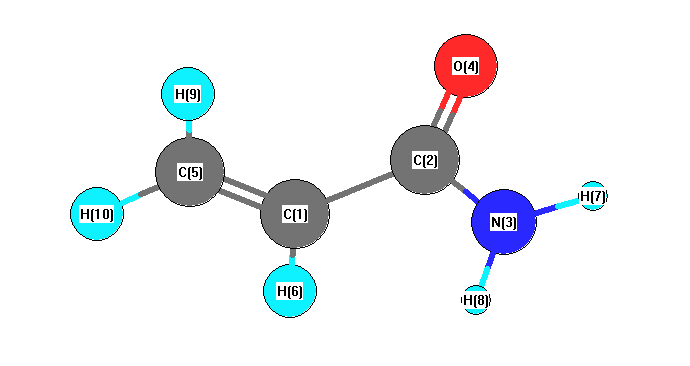
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3994 |
3612 |
41.95 |
|
|
|
| 2 |
A |
3860 |
3491 |
55.87 |
|
|
|
| 3 |
A |
3412 |
3086 |
5.61 |
|
|
|
| 4 |
A |
3333 |
3014 |
17.12 |
|
|
|
| 5 |
A |
3313 |
2996 |
14.58 |
|
|
|
| 6 |
A |
1948 |
1762 |
390.71 |
|
|
|
| 7 |
A |
1846 |
1669 |
59.12 |
|
|
|
| 8 |
A |
1809 |
1636 |
142.50 |
|
|
|
| 9 |
A |
1573 |
1422 |
114.79 |
|
|
|
| 10 |
A |
1468 |
1327 |
66.64 |
|
|
|
| 11 |
A |
1402 |
1268 |
146.81 |
|
|
|
| 12 |
A |
1221 |
1104 |
7.52 |
|
|
|
| 13 |
A |
1138 |
1029 |
7.64 |
|
|
|
| 14 |
A |
1132 |
1023 |
36.97 |
|
|
|
| 15 |
A |
1103 |
998 |
25.76 |
|
|
|
| 16 |
A |
911 |
824 |
45.94 |
|
|
|
| 17 |
A |
881 |
797 |
5.55 |
|
|
|
| 18 |
A |
671 |
607 |
18.01 |
|
|
|
| 19 |
A |
657 |
594 |
5.94 |
|
|
|
| 20 |
A |
511 |
463 |
11.37 |
|
|
|
| 21 |
A |
503 |
455 |
5.79 |
|
|
|
| 22 |
A |
302 |
273 |
7.94 |
|
|
|
| 23 |
A |
132 |
119 |
56.35 |
|
|
|
| 24 |
A |
43 |
39 |
236.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18580.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 16803.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.395 |
-0.341 |
|
-0.402 |
| 2 |
C |
0.687 |
1.009 |
|
0.949 |
| 3 |
N |
-0.915 |
-1.136 |
|
-1.115 |
| 4 |
O |
-0.492 |
-0.669 |
|
-0.630 |
| 5 |
C |
-0.355 |
-0.245 |
|
-0.264 |
| 6 |
H |
0.211 |
0.151 |
|
0.191 |
| 7 |
H |
0.386 |
0.462 |
|
0.464 |
| 8 |
H |
0.380 |
0.445 |
|
0.446 |
| 9 |
H |
0.261 |
0.166 |
|
0.182 |
| 10 |
H |
0.232 |
0.159 |
|
0.178 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.115 |
-3.749 |
0.196 |
3.756 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.092 |
-3.758 |
0.242 |
3.767 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.542 |
-3.213 |
0.397 |
| y |
-3.213 |
-30.576 |
-0.188 |
| z |
0.397 |
-0.188 |
-32.220 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.856 |
-3.213 |
0.397 |
| y |
-3.213 |
-3.695 |
-0.188 |
| z |
0.397 |
-0.188 |
-6.161 |
|
| Polar |
|---|
| 3z2-r2 | -12.322 |
|---|
| x2-y2 | 9.034 |
|---|
| xy | -3.213 |
|---|
| xz | 0.397 |
|---|
| yz | -0.188 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.024 |
-0.848 |
-0.009 |
| y |
-0.848 |
6.049 |
0.013 |
| z |
-0.009 |
0.013 |
3.003 |
<r2> (average value of r
2) Å
2
| <r2> |
113.931 |
| (<r2>)1/2 |
10.674 |