return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCONH2 (Acrylamide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-246.726150
Energy at 298.15K-246.731977
HF Energy-245.874290
Nuclear repulsion energy162.687154
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3776 3576 32.08      
2 A 3640 3448 45.37      
3 A 3301 3126 2.46      
4 A 3209 3039 11.26      
5 A 3192 3023 3.81      
6 A 1813 1717 239.19      
7 A 1701 1611 30.02      
8 A 1688 1599 90.19      
9 A 1466 1388 98.82      
10 A 1377 1304 61.64      
11 A 1311 1242 60.02      
12 A 1143 1082 7.84      
13 A 1064 1008 3.76      
14 A 1010 956 39.03      
15 A 948 898 22.87      
16 A 835 791 8.62      
17 A 809 766 30.02      
18 A 629 596 29.59      
19 A 604 572 12.62      
20 A 478 453 6.61      
21 A 472 447 76.31      
22 A 388 367 209.23      
23 A 284 269 6.16      
24 A 98 93 4.47      

Unscaled Zero Point Vibrational Energy (zpe) 17617.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16685.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.35649 0.14074 0.10103

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.797 -0.656 -0.021
C2 0.477 0.130 -0.005
N3 1.613 -0.640 -0.040
O4 0.511 1.349 -0.004
C5 -1.975 -0.025 0.019
H6 -0.746 -1.741 -0.071
H7 2.479 -0.154 0.130
H8 1.574 -1.607 0.230
H9 -2.003 1.059 0.060
H10 -2.914 -0.567 0.007

Atom - Atom Distances (Å)
  C1 C2 N3 O4 C5 H6 H7 H8 H9 H10
C11.49702.41022.39361.33771.08763.31712.56722.09802.1195
C21.49701.37261.21992.45792.23632.02582.06782.64953.4624
N32.41021.37262.27483.64172.60361.00701.00483.99684.5283
O42.39361.21992.27482.84053.33662.48013.15072.53113.9242
C51.33772.45793.64172.84052.11334.45753.89241.08461.0839
H61.08762.23632.60363.33662.11333.59972.34353.07202.4673
H73.31712.02581.00702.48014.45753.59971.71474.64355.4101
H82.56722.06781.00483.15073.89242.34351.71474.46484.6130
H92.09802.64953.99682.53111.08463.07204.64354.46481.8641
H102.11953.46244.52833.92421.08392.46735.41014.61301.8641

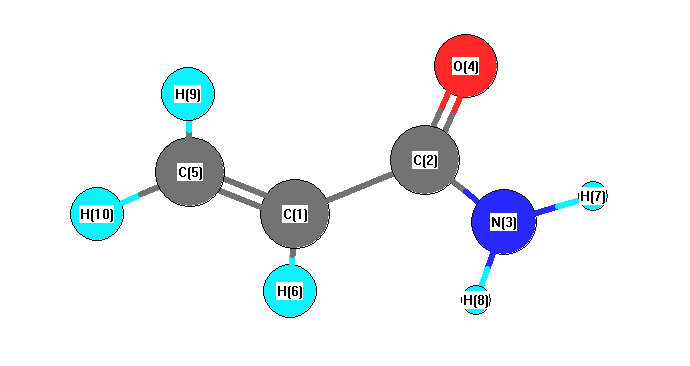
picture of Acrylamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 N3 114.188 C1 C2 O4 123.211
C1 C5 H9 119.658 C1 C5 H10 121.800
C2 C1 C5 120.141 C2 C1 H6 118.968
C2 N3 H7 115.852 C2 N3 H8 120.066
N3 C2 O4 122.562 C5 C1 H6 120.889
H7 N3 H8 116.930 H9 C5 H10 118.542
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability