return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2COOH (Propanoic Acid)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-267.796494
Energy at 298.15K-267.803735
HF Energy-266.906239
Nuclear repulsion energy178.516361
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3778 3578 49.52      
2 A' 3189 3021 18.57      
3 A' 3103 2938 8.92      
4 A' 3100 2936 18.44      
5 A' 1845 1747 229.46      
6 A' 1542 1460 14.52      
7 A' 1505 1425 16.11      
8 A' 1463 1385 4.15      
9 A' 1444 1367 68.19      
10 A' 1340 1269 0.87      
11 A' 1201 1137 261.55      
12 A' 1119 1059 58.94      
13 A' 1038 983 2.27      
14 A' 841 796 9.78      
15 A' 626 592 20.96      
16 A' 477 452 22.80      
17 A' 259 245 2.32      
18 A" 3196 3027 18.87      
19 A" 3148 2981 2.90      
20 A" 1538 1456 9.64      
21 A" 1312 1243 0.05      
22 A" 1140 1079 0.23      
23 A" 829 785 18.89      
24 A" 683 647 110.72      
25 A" 530 502 26.50      
26 A" 227 215 0.00      
27 A" 50 47 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 20258.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19187.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.33785 0.12826 0.09625

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.569 0.000
C2 -0.603 -0.812 0.000
C3 0.454 -1.905 0.000
O4 -0.962 1.522 0.000
O5 1.180 0.832 0.000
H6 -1.260 -0.884 0.872
H7 -1.260 -0.884 -0.872
H8 -0.015 -2.891 0.000
H9 1.094 -1.827 -0.879
H10 1.094 -1.827 0.879
H11 -0.503 2.373 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9 H10 H11
C11.50612.51511.35421.20892.11162.11163.45962.77662.77661.8735
C21.50611.52082.36082.42471.09431.09432.16102.16432.16433.1864
C32.51511.52083.70792.83172.17702.17701.09151.09051.09054.3843
O41.35422.36083.70792.25022.57662.57664.51324.02684.02680.9671
O51.20892.42472.83172.25023.10763.10763.90982.80172.80172.2824
H62.11161.09432.17702.57663.10761.74472.51733.08172.53563.4561
H72.11161.09432.17702.57663.10761.74472.51732.53563.08173.4561
H83.45962.16101.09154.51323.90982.51732.51731.77061.77065.2868
H92.77662.16431.09054.02682.80173.08172.53561.77061.75854.5790
H102.77662.16431.09054.02682.80172.53563.08171.77061.75854.5790
H111.87353.18644.38430.96712.28243.45613.45615.28684.57904.5790

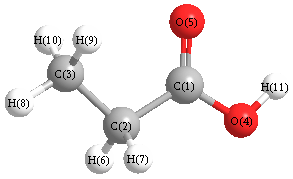
picture of Propanoic Acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.384 C1 C2 H6 107.527
C1 C2 H7 107.527 C1 O4 H11 106.438
C2 C1 O4 111.141 C2 C1 O5 126.171
C2 C3 H8 110.548 C2 C3 H9 110.882
C2 C3 H10 110.882 C3 C2 H6 111.666
C3 C2 H7 111.666 O4 C1 O5 122.688
H6 C2 H7 105.718 H8 C3 H9 108.478
H8 C3 H10 108.478 H9 C3 H10 107.468
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability