return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (2-Propenoic acid)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-266.587938
Energy at 298.15K-266.592613
HF Energy-265.713455
Nuclear repulsion energy162.631065
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3791 3590 66.43      
2 A' 3306 3131 2.86      
3 A' 3246 3074 1.58      
4 A' 3200 3031 2.73      
5 A' 1836 1739 246.63      
6 A' 1687 1598 23.01      
7 A' 1474 1396 35.95      
8 A' 1398 1324 39.71      
9 A' 1324 1254 1.81      
10 A' 1246 1181 196.48      
11 A' 1055 999 82.54      
12 A' 854 808 8.44      
13 A' 593 562 45.14      
14 A' 539 510 6.11      
15 A' 289 274 0.74      
16 A" 1028 974 33.51      
17 A" 958 907 37.57      
18 A" 833 789 41.07      
19 A" 627 594 110.38      
20 A" 488 462 14.66      
21 A" 103 97 0.26      

Unscaled Zero Point Vibrational Energy (zpe) 14936.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14146.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.35883 0.14600 0.10377

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 1.332 0.334 0.000
H2 1.758 1.201 0.000
O3 -0.470 1.685 0.000
C4 0.000 0.569 0.000
C5 -0.812 -0.669 0.000
H6 -1.883 -0.497 0.000
C7 -0.292 -1.903 0.000
H8 0.780 -2.061 0.000
H9 -0.934 -2.777 0.000

Atom - Atom Distances (Å)
  O1 H2 O3 C4 C5 H6 C7 H8 H9
O10.96632.25251.35302.36693.32082.76402.45773.8483
H20.96632.27981.86853.17824.01763.71983.40584.8031
O32.25252.27981.21122.37842.59973.59223.94914.4860
C41.35301.86851.21121.47982.16352.48852.74293.4733
C52.36693.17822.37841.47981.08481.33922.11482.1118
H63.32084.01762.59972.16351.08482.12313.08812.4697
C72.76403.71983.59222.48851.33922.12311.08331.0843
H82.45773.40583.94912.74292.11483.08811.08331.8567
H93.84834.80314.48603.47332.11182.46971.08431.8567

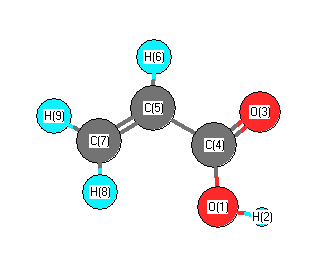
picture of 2-Propenoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C4 O3 122.813 O1 C4 C5 113.271
H2 O1 C4 106.145 O3 C4 C5 123.916
C4 C5 H6 114.165 C4 C5 C7 123.883
C5 C7 H8 121.254 C5 C7 H9 120.876
H6 C5 C7 121.952 H8 C7 H9 117.870
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability