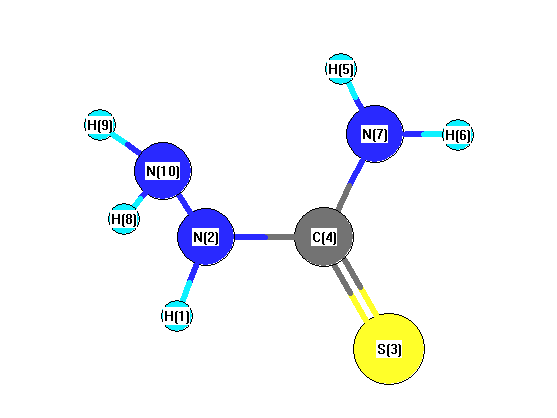
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3996 |
3614 |
86.15 |
|
|
|
| 2 |
A |
3888 |
3516 |
47.15 |
|
|
|
| 3 |
A |
3853 |
3484 |
97.62 |
|
|
|
| 4 |
A |
3835 |
3468 |
6.44 |
|
|
|
| 5 |
A |
3753 |
3394 |
1.97 |
|
|
|
| 6 |
A |
1889 |
1708 |
54.67 |
|
|
|
| 7 |
A |
1804 |
1631 |
230.28 |
|
|
|
| 8 |
A |
1713 |
1549 |
199.08 |
|
|
|
| 9 |
A |
1586 |
1434 |
113.33 |
|
|
|
| 10 |
A |
1458 |
1319 |
1.40 |
|
|
|
| 11 |
A |
1417 |
1282 |
291.85 |
|
|
|
| 12 |
A |
1302 |
1178 |
31.36 |
|
|
|
| 13 |
A |
1121 |
1014 |
93.35 |
|
|
|
| 14 |
A |
1052 |
952 |
114.79 |
|
|
|
| 15 |
A |
879 |
795 |
67.40 |
|
|
|
| 16 |
A |
750 |
678 |
0.96 |
|
|
|
| 17 |
A |
668 |
604 |
7.17 |
|
|
|
| 18 |
A |
569 |
515 |
76.67 |
|
|
|
| 19 |
A |
544 |
492 |
3.65 |
|
|
|
| 20 |
A |
455 |
411 |
373.23 |
|
|
|
| 21 |
A |
428 |
387 |
1.05 |
|
|
|
| 22 |
A |
327 |
296 |
10.74 |
|
|
|
| 23 |
A |
295 |
267 |
12.03 |
|
|
|
| 24 |
A |
105 |
95 |
15.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18842.8 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 17041.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.417 |
0.152 |
|
0.151 |
| 2 |
N |
-0.524 |
0.129 |
|
0.235 |
| 3 |
S |
-0.376 |
-0.527 |
|
-0.467 |
| 4 |
C |
0.473 |
0.154 |
|
-0.096 |
| 5 |
H |
0.407 |
0.261 |
|
0.259 |
| 6 |
H |
0.426 |
0.284 |
|
0.300 |
| 7 |
N |
-0.874 |
-0.401 |
|
-0.329 |
| 8 |
H |
0.371 |
0.343 |
|
0.343 |
| 9 |
H |
0.371 |
0.343 |
|
0.343 |
| 10 |
N |
-0.691 |
-0.737 |
|
-0.740 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
6.689 |
0.659 |
0.001 |
6.721 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
6.721 |
0.670 |
0.001 |
6.755 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.130 |
-5.451 |
0.002 |
| y |
-5.451 |
-31.179 |
-0.002 |
| z |
0.002 |
-0.002 |
-39.700 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.690 |
-5.451 |
0.002 |
| y |
-5.451 |
7.736 |
-0.002 |
| z |
0.002 |
-0.002 |
-5.046 |
|
| Polar |
|---|
| 3z2-r2 | -10.092 |
|---|
| x2-y2 | -6.951 |
|---|
| xy | -5.451 |
|---|
| xz | 0.002 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.985 |
0.667 |
0.000 |
| y |
0.667 |
6.453 |
0.000 |
| z |
0.000 |
0.000 |
3.823 |
<r2> (average value of r
2) Å
2
| <r2> |
160.429 |
| (<r2>)1/2 |
12.666 |