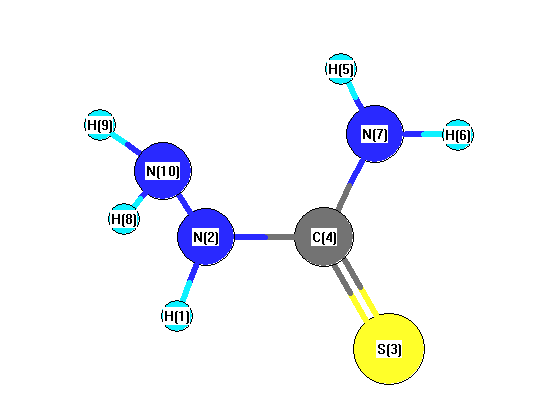
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3787 |
3614 |
70.84 |
|
|
|
| 2 |
A |
3640 |
3474 |
25.80 |
|
|
|
| 3 |
A |
3619 |
3454 |
59.80 |
|
|
|
| 4 |
A |
3606 |
3441 |
2.80 |
|
|
|
| 5 |
A |
3524 |
3363 |
3.18 |
|
|
|
| 6 |
A |
1753 |
1673 |
45.35 |
|
|
|
| 7 |
A |
1650 |
1575 |
253.50 |
|
|
|
| 8 |
A |
1562 |
1491 |
199.69 |
|
|
|
| 9 |
A |
1489 |
1421 |
26.53 |
|
|
|
| 10 |
A |
1345 |
1283 |
0.40 |
|
|
|
| 11 |
A |
1323 |
1263 |
236.34 |
|
|
|
| 12 |
A |
1214 |
1159 |
25.58 |
|
|
|
| 13 |
A |
1046 |
999 |
41.93 |
|
|
|
| 14 |
A |
932 |
889 |
109.72 |
|
|
|
| 15 |
A |
841 |
803 |
57.32 |
|
|
|
| 16 |
A |
695 |
663 |
0.01 |
|
|
|
| 17 |
A |
620 |
591 |
0.28 |
|
|
|
| 18 |
A |
526 |
502 |
0.68 |
|
|
|
| 19 |
A |
510 |
487 |
32.47 |
|
|
|
| 20 |
A |
413 |
394 |
317.45 |
|
|
|
| 21 |
A |
397 |
379 |
1.32 |
|
|
|
| 22 |
A |
307 |
293 |
10.83 |
|
|
|
| 23 |
A |
206 |
197 |
45.92 |
|
|
|
| 24 |
A |
64 |
61 |
21.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17533.6 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 16734.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.389 |
|
|
|
| 2 |
N |
-0.478 |
|
|
|
| 3 |
S |
-0.295 |
|
|
|
| 4 |
C |
0.349 |
|
|
|
| 5 |
H |
0.378 |
|
|
|
| 6 |
H |
0.400 |
|
|
|
| 7 |
N |
-0.797 |
|
|
|
| 8 |
H |
0.361 |
|
|
|
| 9 |
H |
0.361 |
|
|
|
| 10 |
N |
-0.669 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-6.189 |
0.472 |
0.006 |
6.207 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.654 |
5.169 |
-0.005 |
| y |
5.169 |
-30.838 |
-0.004 |
| z |
-0.005 |
-0.004 |
-39.085 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.692 |
5.169 |
-0.005 |
| y |
5.169 |
7.032 |
-0.004 |
| z |
-0.005 |
-0.004 |
-5.339 |
|
| Polar |
|---|
| 3z2-r2 | -10.679 |
|---|
| x2-y2 | -5.816 |
|---|
| xy | 5.169 |
|---|
| xz | -0.005 |
|---|
| yz | -0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
160.375 |
| (<r2>)1/2 |
12.664 |