return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COOCH3 (methyl acetate)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-267.782388
Energy at 298.15K-267.789231
HF Energy-266.895663
Nuclear repulsion energy181.173366
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3236 3065 14.71      
2 A' 3229 3058 4.76      
3 A' 3107 2943 28.51      
4 A' 3106 2941 0.74      
5 A' 1831 1734 199.77      
6 A' 1545 1463 9.22      
7 A' 1515 1435 21.29      
8 A' 1512 1432 12.33      
9 A' 1435 1359 74.90      
10 A' 1306 1237 331.81      
11 A' 1237 1172 0.99      
12 A' 1108 1050 67.04      
13 A' 1010 957 2.04      
14 A' 880 833 23.13      
15 A' 658 623 6.59      
16 A' 432 409 5.83      
17 A' 297 281 11.54      
18 A" 3199 3030 21.16      
19 A" 3191 3022 4.55      
20 A" 1530 1449 9.53      
21 A" 1514 1434 10.80      
22 A" 1208 1144 0.94      
23 A" 1085 1028 5.99      
24 A" 611 579 4.90      
25 A" 186 176 5.12      
26 A" 164 156 2.34      
27 A" 50 48 0.63      

Unscaled Zero Point Vibrational Energy (zpe) 20091.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19028.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.33965 0.14034 0.10313

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.126 1.488 0.000
C2 0.000 0.490 0.000
O3 -1.181 0.749 0.000
O4 0.486 -0.771 0.000
C5 -0.523 -1.788 0.000
H6 0.716 2.495 0.000
H7 1.753 1.339 0.880
H8 1.753 1.339 -0.880
H9 0.018 -2.730 0.000
H10 -1.151 -1.704 0.887
H11 -1.151 -1.704 -0.887

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 H6 H7 H8 H9 H10 H11
C11.50462.42222.34773.66771.08751.09111.09114.36154.01994.0199
C21.50461.20891.35142.33732.12902.13772.13773.22032.63142.6314
O32.42221.20892.25612.62112.57763.11963.11963.68022.60872.6087
O42.34771.35142.25611.43303.27412.61382.61382.01462.08272.0827
C53.66772.33732.62111.43304.45873.96683.96681.08651.08991.0899
H61.08752.12902.57763.27414.45871.78561.78565.27184.68004.6800
H71.09112.13773.11962.61383.96681.78561.76034.51064.20644.5625
H81.09112.13773.11962.61383.96681.78561.76034.51064.56254.2064
H94.36153.22033.68022.01461.08655.27184.51064.51061.79061.7906
H104.01992.63142.60872.08271.08994.68004.20644.56251.79061.7738
H114.01992.63142.60872.08271.08994.68004.56254.20641.79061.7738

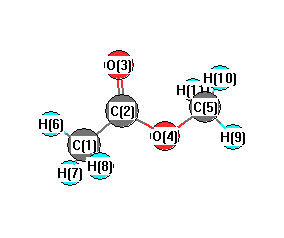
picture of methyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 126.064 C1 C2 O4 110.461
C2 C1 H6 109.380 C2 C1 H7 109.859
C2 C1 H8 109.859 C2 O4 C5 114.125
O3 C2 O4 123.475 O4 C5 H9 105.355
O4 C5 H10 110.548 O4 C5 H11 110.548
H6 C1 H7 110.092 H6 C1 H8 110.092
H7 C1 H8 107.540 H9 C5 H10 110.716
H9 C5 H11 110.716 H10 C5 H11 108.934
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability