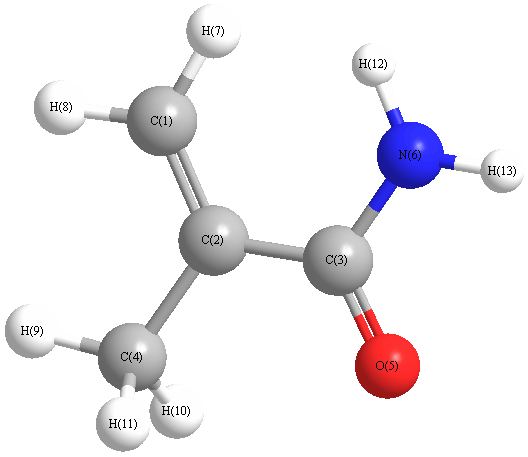
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3971 |
3591 |
33.41 |
|
|
|
| 2 |
A |
3844 |
3477 |
39.39 |
|
|
|
| 3 |
A |
3388 |
3065 |
16.47 |
|
|
|
| 4 |
A |
3309 |
2993 |
10.57 |
|
|
|
| 5 |
A |
3276 |
2963 |
19.97 |
|
|
|
| 6 |
A |
3251 |
2940 |
31.25 |
|
|
|
| 7 |
A |
3193 |
2888 |
29.77 |
|
|
|
| 8 |
A |
1956 |
1769 |
426.30 |
|
|
|
| 9 |
A |
1848 |
1671 |
51.85 |
|
|
|
| 10 |
A |
1810 |
1637 |
110.80 |
|
|
|
| 11 |
A |
1629 |
1473 |
17.28 |
|
|
|
| 12 |
A |
1611 |
1457 |
8.25 |
|
|
|
| 13 |
A |
1577 |
1427 |
10.92 |
|
|
|
| 14 |
A |
1555 |
1406 |
11.39 |
|
|
|
| 15 |
A |
1502 |
1358 |
110.19 |
|
|
|
| 16 |
A |
1340 |
1212 |
65.54 |
|
|
|
| 17 |
A |
1215 |
1098 |
12.99 |
|
|
|
| 18 |
A |
1174 |
1062 |
0.44 |
|
|
|
| 19 |
A |
1116 |
1009 |
7.13 |
|
|
|
| 20 |
A |
1081 |
978 |
43.65 |
|
|
|
| 21 |
A |
1023 |
925 |
3.33 |
|
|
|
| 22 |
A |
910 |
823 |
34.21 |
|
|
|
| 23 |
A |
803 |
726 |
6.11 |
|
|
|
| 24 |
A |
753 |
681 |
18.83 |
|
|
|
| 25 |
A |
628 |
568 |
21.23 |
|
|
|
| 26 |
A |
598 |
541 |
29.63 |
|
|
|
| 27 |
A |
553 |
500 |
5.21 |
|
|
|
| 28 |
A |
420 |
380 |
178.81 |
|
|
|
| 29 |
A |
398 |
360 |
56.83 |
|
|
|
| 30 |
A |
352 |
318 |
5.43 |
|
|
|
| 31 |
A |
284 |
257 |
11.97 |
|
|
|
| 32 |
A |
187 |
169 |
0.16 |
|
|
|
| 33 |
A |
77 |
70 |
3.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25314.5 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 22894.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.898 |
-1.024 |
|
-1.029 |
| 2 |
H |
0.377 |
0.433 |
|
0.438 |
| 3 |
H |
0.374 |
0.391 |
|
0.401 |
| 4 |
C |
-0.634 |
-0.142 |
|
-0.305 |
| 5 |
H |
0.223 |
0.041 |
|
0.084 |
| 6 |
H |
0.248 |
0.068 |
|
0.112 |
| 7 |
H |
0.213 |
0.062 |
|
0.107 |
| 8 |
C |
-0.444 |
-0.474 |
|
-0.578 |
| 9 |
H |
0.232 |
0.186 |
|
0.222 |
| 10 |
H |
0.232 |
0.199 |
|
0.235 |
| 11 |
C |
0.638 |
0.878 |
|
0.835 |
| 12 |
O |
-0.470 |
-0.630 |
|
-0.609 |
| 13 |
C |
-0.091 |
0.012 |
|
0.086 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.502 |
3.170 |
1.713 |
3.638 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.564 |
3.145 |
1.723 |
3.630 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.712 |
-3.919 |
-2.896 |
| y |
-3.919 |
-37.978 |
-0.266 |
| z |
-2.896 |
-0.266 |
-37.418 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.986 |
-3.919 |
-2.896 |
| y |
-3.919 |
-3.413 |
-0.266 |
| z |
-2.896 |
-0.266 |
-2.573 |
|
| Polar |
|---|
| 3z2-r2 | -5.146 |
|---|
| x2-y2 | 6.266 |
|---|
| xy | -3.919 |
|---|
| xz | -2.896 |
|---|
| yz | -0.266 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.616 |
0.535 |
-0.159 |
| y |
0.535 |
9.345 |
-0.268 |
| z |
-0.159 |
-0.268 |
4.822 |
<r2> (average value of r
2) Å
2
| <r2> |
162.436 |
| (<r2>)1/2 |
12.745 |