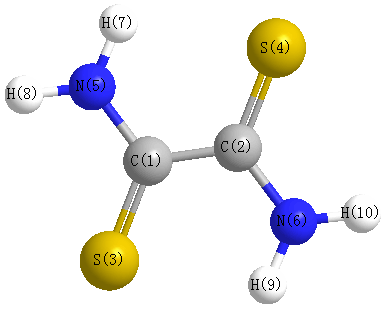
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3733 |
3563 |
0.00 |
|
|
|
| 2 |
Ag |
3526 |
3365 |
0.00 |
|
|
|
| 3 |
Ag |
1674 |
1597 |
0.00 |
|
|
|
| 4 |
Ag |
1468 |
1401 |
0.00 |
|
|
|
| 5 |
Ag |
1351 |
1289 |
0.00 |
|
|
|
| 6 |
Ag |
962 |
918 |
0.00 |
|
|
|
| 7 |
Ag |
689 |
657 |
0.00 |
|
|
|
| 8 |
Ag |
417 |
398 |
0.00 |
|
|
|
| 9 |
Ag |
338 |
322 |
0.00 |
|
|
|
| 10 |
Au |
675 |
645 |
17.30 |
|
|
|
| 11 |
Au |
553 |
528 |
388.38 |
|
|
|
| 12 |
Au |
392 |
374 |
49.37 |
|
|
|
| 13 |
Au |
44 |
42 |
9.71 |
|
|
|
| 14 |
Bg |
727 |
694 |
0.00 |
|
|
|
| 15 |
Bg |
664 |
634 |
0.00 |
|
|
|
| 16 |
Bg |
527 |
503 |
0.00 |
|
|
|
| 17 |
Bu |
3735 |
3564 |
175.71 |
|
|
|
| 18 |
Bu |
3531 |
3370 |
210.34 |
|
|
|
| 19 |
Bu |
1639 |
1564 |
510.15 |
|
|
|
| 20 |
Bu |
1457 |
1390 |
301.84 |
|
|
|
| 21 |
Bu |
1256 |
1199 |
158.30 |
|
|
|
| 22 |
Bu |
885 |
845 |
65.34 |
|
|
|
| 23 |
Bu |
463 |
441 |
1.51 |
|
|
|
| 24 |
Bu |
281 |
269 |
28.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15492.6 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 14786.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.097 |
|
|
|
| 2 |
C |
0.097 |
|
|
|
| 3 |
S |
-0.160 |
|
|
|
| 4 |
S |
-0.160 |
|
|
|
| 5 |
N |
-0.748 |
|
|
|
| 6 |
N |
-0.748 |
|
|
|
| 7 |
H |
0.410 |
|
|
|
| 8 |
H |
0.401 |
|
|
|
| 9 |
H |
0.410 |
|
|
|
| 10 |
H |
0.401 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.816 |
-12.581 |
0.000 |
| y |
-12.581 |
-51.899 |
0.000 |
| z |
0.000 |
0.000 |
-53.323 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.795 |
-12.581 |
0.000 |
| y |
-12.581 |
-2.829 |
0.000 |
| z |
0.000 |
0.000 |
-4.966 |
|
| Polar |
|---|
| 3z2-r2 | -9.932 |
|---|
| x2-y2 | 7.083 |
|---|
| xy | -12.581 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.631 |
3.382 |
0.000 |
| y |
3.382 |
13.394 |
0.000 |
| z |
0.000 |
0.000 |
4.495 |
<r2> (average value of r
2) Å
2
| <r2> |
255.434 |
| (<r2>)1/2 |
15.982 |