return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H12N2 (2-Methyl-1,2-propanediamine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-268.418639
Energy at 298.15K-268.433516
HF Energy-267.393771
Nuclear repulsion energy269.591765
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3564 3375 0.28      
2 A 3508 3322 1.98      
3 A 3181 3013 36.18      
4 A 3156 2989 41.00      
5 A 3071 2908 0.36      
6 A 3055 2894 44.09      
7 A 1738 1646 27.09      
8 A 1730 1639 42.36      
9 A 1548 1466 15.34      
10 A 1541 1460 8.14      
11 A 1510 1430 0.40      
12 A 1451 1374 15.75      
13 A 1396 1322 4.89      
14 A 1307 1237 6.41      
15 A 1261 1194 19.43      
16 A 1145 1084 9.94      
17 A 1052 997 1.39      
18 A 968 917 118.34      
19 A 932 883 163.42      
20 A 899 851 168.33      
21 A 755 716 14.75      
22 A 522 495 12.94      
23 A 433 410 12.21      
24 A 374 354 0.03      
25 A 280 265 0.36      
26 A 263 249 6.10      
27 A 3665 3471 0.53      
28 A 3605 3414 0.17      
29 A 3177 3009 0.19      
30 A 3154 2987 15.29      
31 A 3112 2947 35.36      
32 A 3067 2904 37.43      
33 A 1531 1450 1.61      
34 A 1519 1438 0.03      
35 A 1450 1374 5.47      
36 A 1427 1352 17.21      
37 A 1382 1309 0.23      
38 A 1214 1150 0.33      
39 A 1091 1033 0.94      
40 A 1021 967 0.42      
41 A 971 920 0.37      
42 A 874 827 0.15      
43 A 461 437 1.74      
44 A 362 343 13.34      
45 A 319 302 10.24      
46 A 280 265 81.44      
47 A 227 215 0.13      
48 A 114 108 2.80      

Unscaled Zero Point Vibrational Energy (zpe) 37329.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 35354.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14843 0.08590 0.08513

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.431 0.000
H2 1.473 -0.865 0.878
H3 1.473 -0.865 -0.878
N4 0.106 -2.154 0.000
H5 -0.482 -2.236 0.819
H6 -0.482 -2.236 -0.819
N7 0.896 1.595 0.000
H8 1.502 1.553 -0.815
H9 1.502 1.553 0.815
C10 0.814 -0.880 0.000
C11 -0.866 0.528 -1.250
C12 -0.866 0.528 1.250
H13 -0.251 0.459 -2.155
H14 -0.251 0.459 2.155
H15 -1.613 -0.268 1.286
H16 -1.613 -0.268 -1.286
H17 -1.384 1.489 -1.266
H18 -1.384 1.489 1.266

Atom - Atom Distances (Å)
  C1 H2 H3 N4 H5 H6 N7 H8 H9 C10 C11 C12 H13 H14 H15 H16 H17 H18
C12.14932.14932.58782.83182.83181.46852.04382.04381.54351.52371.52372.16942.16942.17852.17852.15372.1537
H22.14931.75632.07372.38862.92932.67432.95172.41861.09763.45502.74713.73102.52043.16953.81614.27793.7220
H32.14931.75632.07372.92932.38862.67432.41862.95171.09762.74713.45502.52043.73103.81613.16953.72204.2779
N42.58782.07372.07371.01161.01163.83124.04394.04391.45763.11503.11503.40553.40552.85832.85834.13504.1350
H52.83182.38862.92931.01161.63734.15294.57844.27712.04703.47412.82424.01943.01672.31793.09574.36323.8590
H62.83182.92932.38861.01161.63734.15294.27714.57842.04702.82423.47413.01674.01943.09572.31793.85904.3632
N71.46852.67432.67433.83124.15294.15291.01621.01622.47602.40932.40932.69262.69263.37953.37952.61082.6108
H82.04382.95172.41864.04394.57844.27711.01621.63012.65622.61613.30442.46273.61794.17533.63882.92193.5588
H92.04382.41862.95174.04394.27714.57841.01621.63012.65623.30442.61613.61792.46273.63884.17533.55882.9219
C101.54351.09761.09761.45762.04702.04702.47602.65622.65622.52352.52352.75132.75132.81462.81463.47123.4712
C111.52373.45502.74713.11503.47412.82422.40932.61613.30442.52352.50011.09583.46042.76111.09271.09192.7428
C121.52372.74713.45503.11502.82423.47412.40933.30442.61612.52352.50013.46041.09581.09272.76112.74281.0919
H132.16943.73102.52043.40554.01943.01672.69262.46273.61792.75131.09583.46044.30933.77101.77141.77073.7480
H142.16942.52043.73103.40553.01674.01942.69263.61792.46272.75133.46041.09584.30931.77143.77103.74801.7707
H152.17853.16953.81612.85832.31793.09573.37954.17533.63882.81462.76111.09273.77101.77142.57183.10671.7718
H162.17853.81613.16952.85833.09572.31793.37953.63884.17532.81461.09272.76111.77143.77102.57181.77183.1067
H172.15374.27793.72204.13504.36323.85902.61082.92193.55883.47121.09192.74281.77073.74803.10671.77182.5322
H182.15373.72204.27794.13503.85904.36322.61083.55882.92193.47122.74281.09193.74801.77071.77183.10672.5322

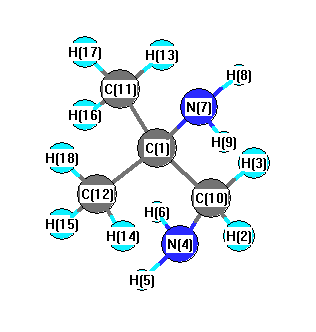
picture of 2-Methyl-1,2-propanediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N7 H8 109.330 C1 N7 H9 109.330
C1 C10 H2 107.733 C1 C10 H3 107.733
C1 C10 N4 119.113 C1 C11 H13 110.768
C1 C11 H16 111.678 C1 C11 H17 109.752
C1 C12 H14 110.768 C1 C12 H15 111.678
C1 C12 H18 109.752 H2 C10 H3 106.268
H2 C10 N4 107.661 H3 C10 N4 107.661
H5 N4 H6 108.044 H5 N4 C10 110.698
H6 N4 C10 110.698 N7 C1 C10 110.551
N7 C1 C11 107.242 N7 C1 C12 107.242
H8 N7 H9 106.663 C10 C1 C11 110.718
C10 C1 C12 110.718 C11 C1 C12 110.253
H13 C11 H16 108.076 H13 C11 H17 108.069
H14 C12 H15 108.076 H14 C12 H18 108.069
H15 C12 H18 108.392 H16 C11 H17 108.392
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability