Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -216.646382 |
| Energy at 298.15K | |
| HF Energy | -215.968753 |
| Nuclear repulsion energy | 116.466577 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3305 |
3130 |
4.84 |
49.56 |
0.73 |
0.85 |
| 2 |
A' |
3208 |
3038 |
5.29 |
149.05 |
0.15 |
0.26 |
| 3 |
A' |
3192 |
3023 |
8.30 |
29.92 |
0.68 |
0.81 |
| 4 |
A' |
3087 |
2924 |
36.72 |
146.60 |
0.11 |
0.20 |
| 5 |
A' |
1721 |
1630 |
2.48 |
5.46 |
0.06 |
0.12 |
| 6 |
A' |
1549 |
1467 |
2.68 |
14.05 |
0.64 |
0.78 |
| 7 |
A' |
1473 |
1395 |
9.91 |
7.90 |
0.57 |
0.73 |
| 8 |
A' |
1455 |
1378 |
11.28 |
5.18 |
0.75 |
0.85 |
| 9 |
A' |
1329 |
1259 |
0.08 |
12.55 |
0.32 |
0.48 |
| 10 |
A' |
1161 |
1099 |
47.88 |
2.36 |
0.75 |
0.85 |
| 11 |
A' |
1032 |
978 |
36.95 |
4.24 |
0.74 |
0.85 |
| 12 |
A' |
934 |
885 |
2.41 |
3.94 |
0.13 |
0.23 |
| 13 |
A' |
619 |
586 |
5.52 |
1.49 |
0.75 |
0.86 |
| 14 |
A' |
278 |
263 |
2.85 |
0.90 |
0.51 |
0.68 |
| 15 |
A" |
3140 |
2973 |
31.18 |
83.51 |
0.75 |
0.86 |
| 16 |
A" |
1296 |
1227 |
0.06 |
8.80 |
0.75 |
0.86 |
| 17 |
A" |
1067 |
1010 |
12.98 |
0.35 |
0.75 |
0.86 |
| 18 |
A" |
1023 |
969 |
20.78 |
0.11 |
0.75 |
0.86 |
| 19 |
A" |
919 |
871 |
46.39 |
0.51 |
0.75 |
0.86 |
| 20 |
A" |
559 |
529 |
11.04 |
7.72 |
0.75 |
0.86 |
| 21 |
A" |
168 |
159 |
3.11 |
3.80 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16256.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15396.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.946 |
-0.201 |
0.000 |
| C2 |
0.000 |
0.956 |
0.000 |
| C3 |
1.330 |
0.835 |
0.000 |
| F4 |
-0.271 |
-1.410 |
0.000 |
| H5 |
1.971 |
1.709 |
0.000 |
| H6 |
1.803 |
-0.140 |
0.000 |
| H7 |
-0.467 |
1.940 |
0.000 |
| H8 |
-1.586 |
-0.176 |
0.888 |
| H9 |
-1.586 |
-0.176 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4951 | 2.5009 | 1.3845 | 3.4865 | 2.7500 | 2.1947 | 1.0948 | 1.0948 |
C2 | 1.4951 | | 1.3357 | 2.3821 | 2.1095 | 2.1101 | 1.0889 | 2.1416 | 2.1416 | C3 | 2.5009 | 1.3357 | | 2.7576 | 1.0835 | 1.0830 | 2.1096 | 3.2115 | 3.2115 | F4 | 1.3845 | 2.3821 | 2.7576 | | 3.8411 | 2.4328 | 3.3563 | 2.0098 | 2.0098 | H5 | 3.4865 | 2.1095 | 1.0835 | 3.8411 | | 1.8557 | 2.4483 | 4.1220 | 4.1220 | H6 | 2.7500 | 2.1101 | 1.0830 | 2.4328 | 1.8557 | | 3.0785 | 3.5037 | 3.5037 | H7 | 2.1947 | 1.0889 | 2.1096 | 3.3563 | 2.4483 | 3.0785 | | 2.5538 | 2.5538 | H8 | 1.0948 | 2.1416 | 3.2115 | 2.0098 | 4.1220 | 3.5037 | 2.5538 | | 1.7761 | H9 | 1.0948 | 2.1416 | 3.2115 | 2.0098 | 4.1220 | 3.5037 | 2.5538 | 1.7761 | |
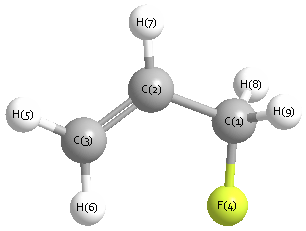 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.029 |
|
C1 |
C2 |
H7 |
115.378 |
| C2 |
C1 |
F4 |
111.575 |
|
C2 |
C1 |
H8 |
110.615 |
| C2 |
C1 |
H9 |
110.615 |
|
C2 |
C3 |
H5 |
121.019 |
| C2 |
C3 |
H6 |
121.126 |
|
C3 |
C2 |
H7 |
120.593 |
| F4 |
C1 |
H8 |
107.744 |
|
F4 |
C1 |
H9 |
107.744 |
| H5 |
C3 |
H6 |
117.855 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -216.645435 |
| Energy at 298.15K | |
| HF Energy | -215.967886 |
| Nuclear repulsion energy | 114.098899 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3285 |
3111 |
11.14 |
56.60 |
0.64 |
0.78 |
| 2 |
A |
3210 |
3040 |
6.84 |
104.09 |
0.25 |
0.40 |
| 3 |
A |
3185 |
3016 |
8.27 |
69.33 |
0.16 |
0.28 |
| 4 |
A |
3161 |
2994 |
25.95 |
68.06 |
0.75 |
0.86 |
| 5 |
A |
3098 |
2934 |
36.15 |
110.00 |
0.09 |
0.16 |
| 6 |
A |
1716 |
1625 |
0.44 |
5.54 |
0.08 |
0.15 |
| 7 |
A |
1552 |
1470 |
2.07 |
7.08 |
0.73 |
0.85 |
| 8 |
A |
1492 |
1413 |
25.62 |
7.69 |
0.66 |
0.79 |
| 9 |
A |
1430 |
1354 |
11.08 |
3.76 |
0.51 |
0.68 |
| 10 |
A |
1327 |
1257 |
0.18 |
9.45 |
0.38 |
0.56 |
| 11 |
A |
1301 |
1233 |
3.52 |
12.34 |
0.72 |
0.84 |
| 12 |
A |
1207 |
1143 |
2.40 |
1.38 |
0.71 |
0.83 |
| 13 |
A |
1082 |
1025 |
107.84 |
5.15 |
0.58 |
0.74 |
| 14 |
A |
1025 |
971 |
41.79 |
0.87 |
0.59 |
0.74 |
| 15 |
A |
1008 |
955 |
2.87 |
1.07 |
0.17 |
0.29 |
| 16 |
A |
948 |
898 |
10.10 |
2.29 |
0.13 |
0.23 |
| 17 |
A |
933 |
884 |
37.43 |
0.49 |
0.68 |
0.81 |
| 18 |
A |
655 |
621 |
7.90 |
4.11 |
0.63 |
0.78 |
| 19 |
A |
441 |
418 |
2.80 |
3.75 |
0.51 |
0.67 |
| 20 |
A |
336 |
318 |
6.85 |
3.30 |
0.75 |
0.86 |
| 21 |
A |
112 |
107 |
1.18 |
5.05 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16252.2 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15392.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.606 |
0.433 |
0.314 |
| C2 |
0.641 |
-0.383 |
0.244 |
| C3 |
1.805 |
0.092 |
-0.212 |
| F4 |
-1.645 |
-0.225 |
-0.341 |
| H5 |
2.702 |
-0.516 |
-0.223 |
| H6 |
1.895 |
1.108 |
-0.585 |
| H7 |
0.565 |
-1.407 |
0.604 |
| H8 |
-0.926 |
0.579 |
1.351 |
| H9 |
-0.467 |
1.407 |
-0.162 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4924 | 2.4916 | 1.3933 | 3.4835 | 2.7424 | 2.2002 | 1.0939 | 1.0929 |
C2 | 1.4924 | | 1.3373 | 2.3654 | 2.1171 | 2.1172 | 1.0878 | 2.1457 | 2.1440 | C3 | 2.4916 | 1.3373 | | 3.4672 | 1.0840 | 1.0859 | 2.1095 | 3.1833 | 2.6255 | F4 | 1.3933 | 2.3654 | 3.4672 | | 4.3584 | 3.7907 | 2.6787 | 2.0060 | 2.0205 | H5 | 3.4835 | 2.1171 | 1.0840 | 4.3584 | | 1.8498 | 2.4580 | 4.1030 | 3.7075 | H6 | 2.7424 | 2.1172 | 1.0859 | 3.7907 | 1.8498 | | 3.0832 | 3.4612 | 2.4182 | H7 | 2.2002 | 1.0878 | 2.1095 | 2.6787 | 2.4580 | 3.0832 | | 2.5927 | 3.0932 | H8 | 1.0939 | 2.1457 | 3.1833 | 2.0060 | 4.1030 | 3.4612 | 2.5927 | | 1.7838 | H9 | 1.0929 | 2.1440 | 2.6255 | 2.0205 | 3.7075 | 2.4182 | 3.0932 | 1.7838 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.314 |
|
C1 |
C2 |
H7 |
116.139 |
| C2 |
C1 |
F4 |
110.061 |
|
C2 |
C1 |
H8 |
111.192 |
| C2 |
C1 |
H9 |
111.116 |
|
C2 |
C3 |
H5 |
121.595 |
| C2 |
C3 |
H6 |
121.441 |
|
C3 |
C2 |
H7 |
120.540 |
| F4 |
C1 |
H8 |
106.906 |
|
F4 |
C1 |
H9 |
108.113 |
| H5 |
C3 |
H6 |
116.964 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
