return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((E)-hexa-1,3,5-triene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-232.754988
Energy at 298.15K-232.762005
HF Energy-231.848849
Nuclear repulsion energy193.587158
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3284 3110 0.00      
2 Ag 3187 3018 0.00      
3 Ag 3178 3010 0.00      
4 Ag 3169 3001 0.00      
5 Ag 1709 1619 0.00      
6 Ag 1634 1547 0.00      
7 Ag 1449 1372 0.00      
8 Ag 1346 1275 0.00      
9 Ag 1324 1254 0.00      
10 Ag 1236 1171 0.00      
11 Ag 959 908 0.00      
12 Ag 449 425 0.00      
13 Ag 353 334 0.00      
14 Au 1034 979 89.92      
15 Au 946 896 3.06      
16 Au 880 833 90.95      
17 Au 695 658 11.93      
18 Au 245 232 2.56      
19 Au 93 88 0.51      
20 Bg 1006 952 0.00      
21 Bg 879 833 0.00      
22 Bg 854 809 0.00      
23 Bg 601 569 0.00      
24 Bg 196 186 0.00      
25 Bu 3284 3110 22.61      
26 Bu 3187 3018 15.12      
27 Bu 3180 3012 38.53      
28 Bu 3171 3003 1.58      
29 Bu 1690 1600 10.42      
30 Bu 1485 1406 5.90      
31 Bu 1336 1265 4.14      
32 Bu 1298 1229 0.47      
33 Bu 1174 1112 6.87      
34 Bu 991 939 2.86      
35 Bu 548 519 3.35      
36 Bu 148 140 1.55      

Unscaled Zero Point Vibrational Energy (zpe) 26096.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24715.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.87839 0.04450 0.04235

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.677 0.000
C2 0.000 -0.677 0.000
C3 1.206 1.486 0.000
C4 -1.206 -1.486 0.000
C5 1.204 2.833 0.000
C6 -1.204 -2.833 0.000
H7 0.954 -1.206 0.000
H8 -0.954 1.206 0.000
H9 2.158 0.956 0.000
H10 -2.158 -0.956 0.000
H11 2.127 3.401 0.000
H12 -2.127 -3.401 0.000
H13 -0.275 -3.395 0.000
H14 0.275 3.395 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.35381.45202.47622.46903.71032.11121.09102.17582.70583.45584.59894.08152.7323
C21.35382.47621.45203.71032.46901.09102.11122.70582.17584.59893.45582.73234.0815
C31.45202.47623.82721.34664.94532.70422.17751.09014.15632.12475.91485.10092.1241
C42.47621.45203.82724.94531.34662.17752.70424.15631.09015.91482.12472.12415.1009
C52.46903.71031.34664.94536.15564.04682.70182.10575.06471.08407.06736.40111.0860
C63.71032.46904.94531.34666.15562.70184.04685.06472.10577.06731.08401.08606.4011
H72.11121.09102.70422.17754.04682.70183.07602.47463.12194.75403.78242.51024.6516
H81.09102.11122.17752.70422.70184.04683.07603.12192.47463.78244.75404.65162.5102
H92.17582.70581.09014.15632.10575.06472.47463.12194.72002.44526.11044.98483.0819
H102.70582.17584.15631.09015.06472.10573.12192.47464.72006.11042.44523.08194.9848
H113.45584.59892.12475.91481.08407.06734.75403.78242.44526.11048.02207.20781.8522
H124.59893.45585.91482.12477.06731.08403.78244.75406.11042.44528.02201.85227.2078
H134.08152.73235.10092.12416.40111.08602.51024.65164.98483.08197.20781.85226.8129
H142.73234.08152.12415.10091.08606.40114.65162.51023.08194.98481.85227.20786.8129

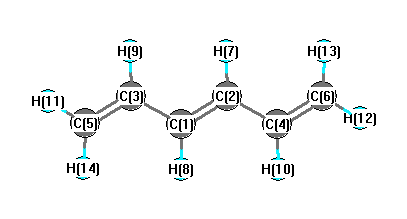
picture of (E)-hexa-1,3,5-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 123.866 C1 C2 H7 119.036
C1 C3 C5 123.787 C1 C3 H9 117.014
C2 C1 C3 123.866 C2 C1 H8 119.036
C2 C4 C6 123.787 C2 C4 H10 117.014
C3 C1 H8 117.098 C3 C5 H11 121.515
C3 C5 H14 121.285 C4 C2 H7 117.098
C4 C6 H12 121.515 C4 C6 H13 121.285
C5 C3 H9 119.199 C6 C4 H10 119.199
H11 C5 H14 117.201 H12 C6 H13 117.201
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability