return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O (allenol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-191.412724
Energy at 298.15K-191.416169
HF Energy-190.750389
Nuclear repulsion energy101.867061
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3819 3617 34.56      
2 A' 3244 3073 4.46      
3 A' 3151 2984 13.47      
4 A' 2051 1942 46.04      
5 A' 1523 1442 49.06      
6 A' 1441 1365 59.76      
7 A' 1316 1246 3.32      
8 A' 1219 1154 151.15      
9 A' 999 946 104.97      
10 A' 916 868 40.33      
11 A' 619 587 24.96      
12 A' 208 197 0.58      
13 A" 3235 3064 6.04      
14 A" 1052 996 0.70      
15 A" 911 863 32.84      
16 A" 636 602 0.09      
17 A" 425 403 138.31      
18 A" 253 239 5.88      

Unscaled Zero Point Vibrational Energy (zpe) 13508.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12793.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.46687 0.14442 0.13512

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.657 -0.495 0.000
C2 0.000 0.644 0.000
C3 -0.645 1.790 0.000
O4 0.110 -1.751 0.000
H5 1.741 -0.538 0.000
H6 -0.922 2.288 0.926
H7 -0.922 2.288 -0.926
H8 -0.849 -1.663 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8
C11.31552.63001.36971.08433.33163.33161.9061
C21.31551.31462.39732.10462.10002.10002.4581
C32.63001.31463.61993.33341.08751.08753.4584
O41.36972.39733.61992.03244.26994.26990.9631
H51.08432.10463.33342.03243.99233.99232.8238
H63.33162.10001.08754.26993.99231.85214.0581
H73.33162.10001.08754.26993.99231.85214.0581
H81.90612.45813.45840.96312.82384.05814.0581

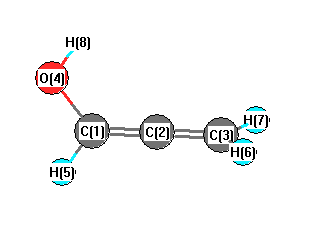
picture of allenol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.383 C1 O4 H8 108.322
C2 C1 O4 126.442 C2 C1 H5 122.261
C2 C3 H6 121.625 C2 C3 H7 121.625
O4 C1 H5 111.297 H6 C3 H7 116.749
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.050      
2 C -0.092      
3 C -0.492      
4 O -0.620      
5 H 0.248      
6 H 0.239      
7 H 0.239      
8 H 0.429      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -21.142 0.638 0.000
y 0.638 -25.162 0.000
z 0.000 0.000 -24.880
Traceless
 xyz
x 3.880 0.638 0.000
y 0.638 -2.151 0.000
z 0.000 0.000 -1.728
Polar
3z2-r2-3.457
x2-y24.021
xy0.638
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000