return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propenalol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-266.568748
Energy at 298.15K-266.573802
HF Energy-265.689834
Nuclear repulsion energy164.112076
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3463 3279 105.46      
2 A' 3270 3097 2.99      
3 A' 3228 3057 16.36      
4 A' 3030 2869 114.42      
5 A' 1726 1635 176.25      
6 A' 1671 1582 289.97      
7 A' 1492 1413 46.85      
8 A' 1443 1366 64.24      
9 A' 1427 1352 11.49      
10 A' 1306 1237 172.18      
11 A' 1129 1069 18.25      
12 A' 999 946 63.34      
13 A' 900 852 8.89      
14 A' 506 479 19.70      
15 A' 289 274 4.73      
16 A" 1042 987 10.91      
17 A" 989 937 24.62      
18 A" 849 804 86.00      
19 A" 777 736 62.21      
20 A" 376 356 4.12      
21 A" 263 249 5.21      

Unscaled Zero Point Vibrational Energy (zpe) 15086.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 14288.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.33003 0.16611 0.11050

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.094 0.000
C2 1.249 0.363 0.000
C3 -1.188 0.426 0.000
O4 1.327 -0.871 0.000
O5 -1.325 -0.891 0.000
H6 -0.421 -1.273 0.000
H7 0.002 2.177 0.000
H8 2.180 0.955 0.000
H9 -2.144 0.943 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.44731.36312.37152.38662.40441.08232.18482.1493
C21.44732.43771.23672.86282.33772.20081.10393.4420
C31.36312.43772.82971.32381.86382.11713.40971.0867
O42.37151.23672.82972.65161.79353.32332.01613.9162
O52.38662.86281.32382.65160.98113.34223.96152.0082
H62.40442.33771.86381.79350.98113.47543.42522.8065
H71.08232.20082.11713.32333.34223.47542.49722.4757
H82.18481.10393.40972.01613.96153.42522.49724.3243
H92.14933.44201.08673.91622.00822.80652.47574.3243

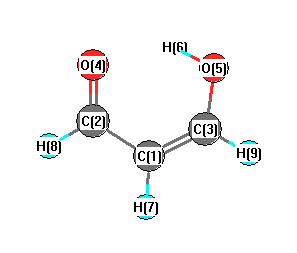
picture of propenalol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 123.969 C1 C2 H8 117.196
C1 C3 O5 125.295 C1 C3 H9 122.239
C2 C1 C3 120.280 C2 C1 H7 120.234
C3 C1 H7 119.486 C3 O5 H6 106.988
O4 C2 H8 118.835 O5 C3 H9 112.466
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.560      
2 C 0.273      
3 C 0.243      
4 O -0.517      
5 O -0.612      
6 H 0.485      
7 H 0.228      
8 H 0.205      
9 H 0.256      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000