return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4N2 (2H-Imidazole)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-225.590733
Energy at 298.15K-225.596718
HF Energy-224.768519
Nuclear repulsion energy166.518710
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3277 3103 11.58      
2 A1 3123 2958 0.70      
3 A1 1675 1586 18.75      
4 A1 1446 1369 13.46      
5 A1 1404 1330 14.23      
6 A1 1272 1204 6.32      
7 A1 1054 998 0.94      
8 A1 935 885 13.90      
9 A2 1159 1097 0.00      
10 A2 877 831 0.00      
11 A2 538 510 0.00      
12 B1 3180 3011 0.55      
13 B1 1020 966 21.71      
14 B1 819 776 9.89      
15 B1 351 333 37.67      
16 B2 3263 3090 4.22      
17 B2 1742 1650 0.01      
18 B2 1405 1331 33.18      
19 B2 1251 1185 1.93      
20 B2 1096 1038 34.67      
21 B2 940 891 68.74      

Unscaled Zero Point Vibrational Energy (zpe) 15912.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15070.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.36237 0.30273 0.17035

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.204
N2 0.000 0.997 0.284
N3 0.000 -0.997 0.284
C4 0.000 0.726 -0.950
C5 0.000 -0.726 -0.950
H6 -0.897 0.000 1.828
H7 0.897 0.000 1.828
H8 0.000 1.479 -1.728
H9 0.000 -1.479 -1.728

Atom - Atom Distances (Å)
  C1 N2 N3 C4 C5 H6 H7 H8 H9
C11.35611.35612.27292.27291.09281.09283.28393.2839
N21.35611.99351.26352.11962.04462.04462.06953.1905
N31.35611.99352.11961.26352.04462.04463.19052.0695
C42.27291.26352.11961.45283.00793.00791.08262.3387
C52.27292.11961.26351.45283.00793.00792.33871.0826
H61.09282.04462.04463.00793.00791.79443.95433.9543
H71.09282.04462.04463.00793.00791.79443.95433.9543
H83.28392.06953.19051.08262.33873.95433.95432.9579
H93.28393.19052.06952.33871.08263.95433.95432.9579

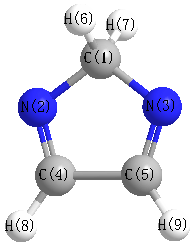
picture of 2H-Imidazole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 C4 120.337 C1 N3 C5 120.337
N2 C1 N3 94.616 N2 C1 H6 112.773
N2 C1 H7 112.773 N2 C4 C5 102.355
N2 C4 H8 123.609 N3 C1 H6 112.773
N3 C1 H7 112.773 N3 C5 C4 102.355
N3 C5 H9 123.609 C4 C5 H9 134.036
C5 C4 H8 134.036 H6 C1 H7 110.378
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability