Jump to
S1C2
S1C3
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.829663 |
| Energy at 298.15K | -169.833694 |
| HF Energy | -169.829663 |
| Nuclear repulsion energy | 71.029622 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3902 |
3902 |
62.87 |
|
|
|
| 2 |
A' |
3572 |
3572 |
6.29 |
|
|
|
| 3 |
A' |
3084 |
3084 |
60.56 |
|
|
|
| 4 |
A' |
1815 |
1815 |
210.68 |
|
|
|
| 5 |
A' |
1465 |
1465 |
17.06 |
|
|
|
| 6 |
A' |
1364 |
1364 |
197.53 |
|
|
|
| 7 |
A' |
1224 |
1224 |
50.02 |
|
|
|
| 8 |
A' |
1082 |
1082 |
209.58 |
|
|
|
| 9 |
A' |
631 |
631 |
1.23 |
|
|
|
| 10 |
A" |
1072 |
1072 |
7.49 |
|
|
|
| 11 |
A" |
847 |
847 |
71.32 |
|
|
|
| 12 |
A" |
413 |
413 |
88.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10236.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10236.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.372 |
0.000 |
| O2 |
-0.998 |
-0.536 |
0.000 |
| N3 |
1.195 |
-0.008 |
0.000 |
| H4 |
-0.343 |
1.414 |
0.000 |
| H5 |
-1.850 |
-0.094 |
0.000 |
| H6 |
1.815 |
0.796 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3497 | 1.2538 | 1.0969 | 1.9078 | 1.8636 |
O2 | 1.3497 | | 2.2553 | 2.0573 | 0.9603 | 3.1125 | N3 | 1.2538 | 2.2553 | | 2.0949 | 3.0459 | 1.0159 | H4 | 1.0969 | 2.0573 | 2.0949 | | 2.1315 | 2.2447 | H5 | 1.9078 | 0.9603 | 3.0459 | 2.1315 | | 3.7713 | H6 | 1.8636 | 3.1125 | 1.0159 | 2.2447 | 3.7713 | |
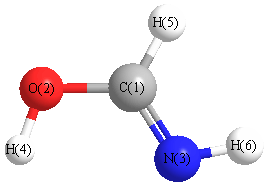 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
110.224 |
|
C1 |
N3 |
H6 |
109.945 |
| O2 |
C1 |
N3 |
120.011 |
|
O2 |
C1 |
H4 |
114.075 |
| N3 |
C1 |
H4 |
125.915 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.123 |
|
|
|
| 2 |
O |
-0.491 |
|
|
|
| 3 |
N |
-0.508 |
|
|
|
| 4 |
H |
0.178 |
|
|
|
| 5 |
H |
0.408 |
|
|
|
| 6 |
H |
0.290 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.128 |
3.453 |
0.000 |
4.056 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.679 |
1.251 |
0.000 |
| y |
1.251 |
-17.928 |
0.000 |
| z |
0.000 |
0.000 |
-18.843 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.706 |
1.251 |
0.000 |
| y |
1.251 |
-1.667 |
0.000 |
| z |
0.000 |
0.000 |
-3.039 |
|
| Polar |
|---|
| 3z2-r2 | -6.078 |
|---|
| x2-y2 | 4.249 |
|---|
| xy | 1.251 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.769 |
0.004 |
0.000 |
| y |
0.004 |
2.831 |
0.000 |
| z |
0.000 |
0.000 |
1.698 |
<r2> (average value of r
2) Å
2
| <r2> |
41.071 |
| (<r2>)1/2 |
6.409 |
Jump to
S1C1
S1C3
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.841009 |
| Energy at 298.15K | -169.845194 |
| HF Energy | -169.841009 |
| Nuclear repulsion energy | 71.406627 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3791 |
3791 |
45.08 |
|
|
|
| 2 |
A' |
3556 |
3556 |
7.33 |
|
|
|
| 3 |
A' |
3180 |
3180 |
32.32 |
|
|
|
| 4 |
A' |
1782 |
1782 |
297.78 |
|
|
|
| 5 |
A' |
1433 |
1433 |
28.17 |
|
|
|
| 6 |
A' |
1397 |
1397 |
1.83 |
|
|
|
| 7 |
A' |
1230 |
1230 |
128.59 |
|
|
|
| 8 |
A' |
1098 |
1098 |
178.86 |
|
|
|
| 9 |
A' |
604 |
604 |
55.03 |
|
|
|
| 10 |
A" |
1087 |
1087 |
0.00 |
|
|
|
| 11 |
A" |
850 |
850 |
33.21 |
|
|
|
| 12 |
A" |
640 |
640 |
224.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10323.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10323.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| O2 |
-1.106 |
-0.339 |
0.000 |
| N3 |
1.152 |
-0.089 |
0.000 |
| H4 |
-0.266 |
1.475 |
0.000 |
| H5 |
-0.818 |
-1.263 |
0.000 |
| H6 |
1.870 |
0.629 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3396 | 1.2583 | 1.0913 | 1.8681 | 1.8820 |
O2 | 1.3396 | | 2.2722 | 1.9986 | 0.9679 | 3.1296 | N3 | 1.2583 | 2.2722 | | 2.1115 | 2.2936 | 1.0153 | H4 | 1.0913 | 1.9986 | 2.1115 | | 2.7927 | 2.2977 | H5 | 1.8681 | 0.9679 | 2.2936 | 2.7927 | | 3.2872 | H6 | 1.8820 | 3.1296 | 1.0153 | 2.2977 | 3.2872 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
107.014 |
|
C1 |
N3 |
H6 |
111.293 |
| O2 |
C1 |
N3 |
121.980 |
|
O2 |
C1 |
H4 |
110.193 |
| N3 |
C1 |
H4 |
127.826 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.157 |
|
|
|
| 2 |
O |
-0.514 |
|
|
|
| 3 |
N |
-0.566 |
|
|
|
| 4 |
H |
0.214 |
|
|
|
| 5 |
H |
0.409 |
|
|
|
| 6 |
H |
0.300 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.208 |
1.053 |
0.000 |
1.074 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.697 |
3.446 |
0.000 |
| y |
3.446 |
-13.865 |
0.000 |
| z |
0.000 |
0.000 |
-18.835 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.347 |
3.446 |
0.000 |
| y |
3.446 |
5.401 |
0.000 |
| z |
0.000 |
0.000 |
-2.053 |
|
| Polar |
|---|
| 3z2-r2 | -4.107 |
|---|
| x2-y2 | -5.832 |
|---|
| xy | 3.446 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.275 |
0.052 |
0.000 |
| y |
0.052 |
3.246 |
0.000 |
| z |
0.000 |
0.000 |
1.686 |
<r2> (average value of r
2) Å
2
| <r2> |
40.410 |
| (<r2>)1/2 |
6.357 |
Jump to
S1C1
S1C2
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.834825 |
| Energy at 298.15K | -169.838932 |
| HF Energy | -169.834825 |
| Nuclear repulsion energy | 70.990736 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3821 |
3821 |
26.97 |
|
|
|
| 2 |
A' |
3474 |
3474 |
6.32 |
|
|
|
| 3 |
A' |
3228 |
3228 |
13.13 |
|
|
|
| 4 |
A' |
1788 |
1788 |
302.25 |
|
|
|
| 5 |
A' |
1428 |
1428 |
1.47 |
|
|
|
| 6 |
A' |
1372 |
1372 |
23.72 |
|
|
|
| 7 |
A' |
1152 |
1152 |
295.03 |
|
|
|
| 8 |
A' |
1100 |
1100 |
35.50 |
|
|
|
| 9 |
A' |
607 |
607 |
40.84 |
|
|
|
| 10 |
A" |
1102 |
1102 |
74.37 |
|
|
|
| 11 |
A" |
870 |
870 |
70.10 |
|
|
|
| 12 |
A" |
515 |
515 |
96.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10227.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10227.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.439 |
0.000 |
| O2 |
-1.094 |
-0.354 |
0.000 |
| N3 |
1.218 |
0.127 |
0.000 |
| H4 |
-0.305 |
1.482 |
0.000 |
| H5 |
-0.835 |
-1.284 |
0.000 |
| H6 |
1.366 |
-0.885 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3512 | 1.2574 | 1.0872 | 1.9146 | 1.9025 |
O2 | 1.3512 | | 2.3615 | 1.9989 | 0.9652 | 2.5170 | N3 | 1.2574 | 2.3615 | | 2.0390 | 2.4909 | 1.0223 | H4 | 1.0872 | 1.9989 | 2.0390 | | 2.8167 | 2.8978 | H5 | 1.9146 | 0.9652 | 2.4909 | 2.8167 | | 2.2373 | H6 | 1.9025 | 2.5170 | 1.0223 | 2.8978 | 2.2373 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
110.374 |
|
C1 |
N3 |
H6 |
112.726 |
| O2 |
C1 |
N3 |
129.683 |
|
O2 |
C1 |
H4 |
109.647 |
| N3 |
C1 |
H4 |
120.670 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.150 |
|
|
|
| 2 |
O |
-0.514 |
|
|
|
| 3 |
N |
-0.537 |
|
|
|
| 4 |
H |
0.242 |
|
|
|
| 5 |
H |
0.393 |
|
|
|
| 6 |
H |
0.267 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.905 |
-2.013 |
0.000 |
2.207 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.362 |
-1.994 |
0.000 |
| y |
-1.994 |
-12.804 |
0.000 |
| z |
0.000 |
0.000 |
-18.815 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.553 |
-1.994 |
0.000 |
| y |
-1.994 |
8.285 |
0.000 |
| z |
0.000 |
0.000 |
-0.732 |
|
| Polar |
|---|
| 3z2-r2 | -1.463 |
|---|
| x2-y2 | -10.558 |
|---|
| xy | -1.994 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.086 |
-0.174 |
0.000 |
| y |
-0.174 |
3.422 |
0.000 |
| z |
0.000 |
0.000 |
1.706 |
<r2> (average value of r
2) Å
2
| <r2> |
40.964 |
| (<r2>)1/2 |
6.400 |
