Jump to
S1C2
S1C3
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.757083 |
| Energy at 298.15K | -169.761087 |
| HF Energy | -169.575839 |
| Nuclear repulsion energy | 70.795794 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3846 |
3846 |
49.47 |
|
|
|
| 2 |
A' |
3552 |
3552 |
2.50 |
|
|
|
| 3 |
A' |
3056 |
3056 |
76.50 |
|
|
|
| 4 |
A' |
1766 |
1766 |
172.52 |
|
|
|
| 5 |
A' |
1457 |
1457 |
14.38 |
|
|
|
| 6 |
A' |
1359 |
1359 |
171.59 |
|
|
|
| 7 |
A' |
1217 |
1217 |
75.59 |
|
|
|
| 8 |
A' |
1073 |
1073 |
208.53 |
|
|
|
| 9 |
A' |
625 |
625 |
1.06 |
|
|
|
| 10 |
A" |
1049 |
1049 |
6.57 |
|
|
|
| 11 |
A" |
842 |
842 |
71.15 |
|
|
|
| 12 |
A" |
397 |
397 |
82.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10118.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10118.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.375 |
0.000 |
| O2 |
-1.003 |
-0.535 |
0.000 |
| N3 |
1.200 |
-0.014 |
0.000 |
| H4 |
-0.343 |
1.417 |
0.000 |
| H5 |
-1.849 |
-0.081 |
0.000 |
| H6 |
1.821 |
0.789 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3546 | 1.2611 | 1.0966 | 1.9048 | 1.8677 |
O2 | 1.3546 | | 2.2638 | 2.0603 | 0.9605 | 3.1194 | N3 | 1.2611 | 2.2638 | | 2.1039 | 3.0499 | 1.0151 | H4 | 1.0966 | 2.0603 | 2.1039 | | 2.1239 | 2.2538 | H5 | 1.9048 | 0.9605 | 3.0499 | 2.1239 | | 3.7724 | H6 | 1.8677 | 3.1194 | 1.0151 | 2.2538 | 3.7724 | |
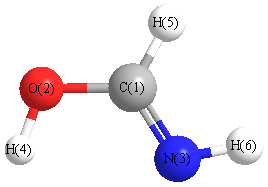 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.552 |
|
C1 |
N3 |
H6 |
109.810 |
| O2 |
C1 |
N3 |
119.827 |
|
O2 |
C1 |
H4 |
113.977 |
| N3 |
C1 |
H4 |
126.196 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.757083 |
| Energy at 298.15K | -169.761087 |
| HF Energy | -169.575839 |
| Nuclear repulsion energy | 70.795794 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -169.762488 |
| Energy at 298.15K | -169.766597 |
| HF Energy | -169.580909 |
| Nuclear repulsion energy | 70.684364 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3772 |
3772 |
19.85 |
|
|
|
| 2 |
A' |
3462 |
3462 |
9.52 |
|
|
|
| 3 |
A' |
3213 |
3213 |
14.64 |
|
|
|
| 4 |
A' |
1728 |
1728 |
258.98 |
|
|
|
| 5 |
A' |
1434 |
1434 |
0.36 |
|
|
|
| 6 |
A' |
1372 |
1372 |
28.78 |
|
|
|
| 7 |
A' |
1146 |
1146 |
273.79 |
|
|
|
| 8 |
A' |
1092 |
1092 |
55.32 |
|
|
|
| 9 |
A' |
594 |
594 |
37.25 |
|
|
|
| 10 |
A" |
1089 |
1089 |
82.13 |
|
|
|
| 11 |
A" |
854 |
854 |
56.26 |
|
|
|
| 12 |
A" |
542 |
542 |
90.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10148.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10148.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.437 |
0.000 |
| O2 |
-1.102 |
-0.355 |
0.000 |
| N3 |
1.226 |
0.129 |
0.000 |
| H4 |
-0.306 |
1.479 |
0.000 |
| H5 |
-0.840 |
-1.284 |
0.000 |
| H6 |
1.380 |
-0.881 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3575 | 1.2647 | 1.0856 | 1.9150 | 1.9088 |
O2 | 1.3575 | | 2.3786 | 1.9992 | 0.9654 | 2.5379 | N3 | 1.2647 | 2.3786 | | 2.0424 | 2.5029 | 1.0217 | H4 | 1.0856 | 1.9992 | 2.0424 | | 2.8137 | 2.9007 | H5 | 1.9150 | 0.9654 | 2.5029 | 2.8137 | | 2.2561 | H6 | 1.9088 | 2.5379 | 1.0217 | 2.9007 | 2.2561 | |
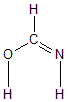 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.905 |
|
C1 |
N3 |
H6 |
112.774 |
| O2 |
C1 |
N3 |
130.182 |
|
O2 |
C1 |
H4 |
109.322 |
| N3 |
C1 |
H4 |
120.496 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
