return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 (3-Methylenecyclopentene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-232.783929
Energy at 298.15K-232.792281
HF Energy-231.869573
Nuclear repulsion energy216.875157
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3275 3102 10.95      
2 A 3240 3068 16.61      
3 A 3217 3046 2.46      
4 A 3179 3011 5.03      
5 A 3155 2989 20.15      
6 A 3127 2962 19.79      
7 A 3091 2927 15.00      
8 A 3072 2910 33.11      
9 A 1711 1621 11.30      
10 A 1631 1544 0.08      
11 A 1532 1451 5.13      
12 A 1506 1427 3.86      
13 A 1468 1390 1.30      
14 A 1380 1307 1.56      
15 A 1351 1279 3.46      
16 A 1312 1242 0.37      
17 A 1286 1218 0.48      
18 A 1251 1185 2.07      
19 A 1191 1128 1.59      
20 A 1127 1068 1.26      
21 A 1045 990 5.28      
22 A 1031 977 2.18      
23 A 976 924 2.41      
24 A 941 891 5.59      
25 A 924 875 1.07      
26 A 889 842 1.09      
27 A 843 798 0.36      
28 A 822 778 44.48      
29 A 800 757 23.22      
30 A 752 712 5.04      
31 A 668 633 18.15      
32 A 637 603 2.36      
33 A 491 465 12.78      
34 A 356 338 0.50      
35 A 289 274 1.69      
36 A 102 97 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 26832.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 25413.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.24266 0.11583 0.08154

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.102 -1.180 0.153
H2 -0.182 -2.001 -0.510
H3 0.066 -1.556 1.181
C4 1.509 -0.615 -0.145
H5 1.827 -0.854 -1.167
H6 2.275 -1.015 0.527
C7 -2.167 -0.018 -0.100
H8 -2.747 0.897 -0.161
H9 -2.712 -0.955 -0.148
C10 -0.830 0.013 0.032
C11 1.321 0.874 0.006
H12 2.145 1.582 -0.004
C13 0.016 1.207 0.078
H14 -0.375 2.219 0.137

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 H9 C10 C11 H12 C13 H14
C11.09271.09521.54462.19652.21092.56213.54042.83891.51862.39343.43932.39013.4325
H21.09271.76622.21662.40552.84352.83573.88632.76122.18383.28544.30263.26794.2739
H31.09521.76622.17333.01812.36642.99883.96653.13722.14052.97683.94622.97513.9408
C41.54462.21662.17331.09721.09433.72414.51684.23412.42781.50902.29212.36623.4146
H52.19652.40553.01811.09721.75984.21765.00024.65303.04102.14892.71793.01303.9988
H62.21092.84352.36641.09431.75984.59535.41775.03243.30752.17962.65383.19994.1986
C72.56212.83572.99883.72414.21764.59531.08531.08521.34393.60154.60012.50952.8759
H83.54043.88633.96654.51685.00025.41771.08531.85282.12054.07154.94242.79102.7321
H92.83892.76123.13724.23414.65305.03241.08521.85282.12374.43075.48133.48823.9518
C101.51862.18382.14052.42783.04103.30751.34392.12052.12372.31703.36371.46452.2552
C112.39343.28542.97681.50902.14892.17963.60154.07154.43072.31701.08641.34842.1679
H123.43934.30263.94622.29212.71792.65384.60014.94245.48133.36371.08642.16312.6027
C132.39013.26792.97512.36623.01303.19992.50952.79103.48821.46451.34842.16311.0862
H143.43254.27393.94083.41463.99884.19862.87592.73213.95182.25522.16792.60271.0862

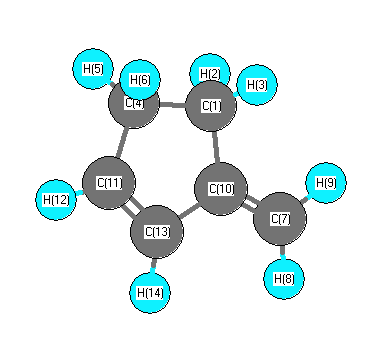
picture of 3-Methylenecyclopentene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 111.363 C1 C4 H6 112.698
C1 C4 C11 103.210 C1 C10 C7 126.922
C1 C10 C13 106.478 H2 C1 H3 107.651
H2 C1 C4 113.265 H2 C1 C10 112.474
H3 C1 C4 109.651 H3 C1 C10 108.877
C4 C1 C10 104.848 C4 C11 H12 123.220
C4 C11 C13 111.685 H5 C4 H6 106.832
H5 C4 C11 110.079 H6 C4 C11 112.726
C7 C10 C13 126.599 H8 C7 H9 117.212
H8 C7 C10 121.234 H9 C7 C10 121.553
C10 C13 C11 110.848 C10 C13 H14 123.614
C11 C13 H14 125.520 H12 C11 C13 125.005
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability