return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H8 (Indene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-346.882302
Energy at 298.15K-346.890267
HF Energy-345.519569
Nuclear repulsion energy394.081976
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3249 3077 15.49      
2 A' 3226 3055 16.39      
3 A' 3226 3055 9.95      
4 A' 3213 3043 24.98      
5 A' 3201 3032 1.33      
6 A' 3195 3026 6.12      
7 A' 3070 2908 8.83      
8 A' 1666 1577 1.17      
9 A' 1640 1553 0.49      
10 A' 1598 1514 0.41      
11 A' 1510 1430 8.53      
12 A' 1497 1418 7.30      
13 A' 1478 1399 3.39      
14 A' 1458 1381 12.31      
15 A' 1382 1309 6.58      
16 A' 1330 1259 0.07      
17 A' 1268 1201 1.66      
18 A' 1247 1181 3.64      
19 A' 1197 1134 1.15      
20 A' 1189 1126 0.08      
21 A' 1141 1081 0.40      
22 A' 1094 1036 1.22      
23 A' 1047 992 3.53      
24 A' 984 932 8.14      
25 A' 872 826 2.71      
26 A' 849 804 1.05      
27 A' 743 703 2.29      
28 A' 600 569 1.46      
29 A' 541 513 0.19      
30 A' 382 362 1.39      
31 A" 3115 2950 6.48      
32 A" 1153 1092 1.52      
33 A" 927 878 2.48      
34 A" 879 832 0.40      
35 A" 845 800 0.93      
36 A" 833 789 0.00      
37 A" 812 769 0.24      
38 A" 725 686 108.20      
39 A" 687 651 7.72      
40 A" 524 496 7.97      
41 A" 398 377 4.02      
42 A" 378 358 1.30      
43 A" 348 330 4.98      
44 A" 200 189 4.38      
45 A" 182 173 0.99      

Unscaled Zero Point Vibrational Energy (zpe) 30549.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 28933.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.12580 0.05265 0.03738

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.285 -0.018 0.000
C2 -1.854 -1.353 0.000
C3 -0.491 -1.665 0.000
C4 -1.359 1.032 0.000
C5 0.000 0.726 0.000
C6 0.435 -0.616 0.000
C7 1.899 -0.622 0.000
C8 2.355 0.653 0.000
C9 1.206 1.628 0.000
H10 -3.349 0.202 0.000
H11 -2.590 -2.153 0.000
H12 -0.163 -2.702 0.000
H13 -1.701 2.065 0.000
H14 2.514 -1.516 0.000
H15 3.398 0.953 0.000
H16 1.228 2.281 0.881
H17 1.228 2.281 -0.881

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.40252.43471.40032.40302.78434.22684.68813.85961.08692.15623.42112.16375.02735.76514.29044.2904
C21.40251.39762.43582.78522.40393.82304.66224.27162.15701.08692.16303.42144.37085.73544.84594.8459
C32.43471.39762.83302.44061.39882.60773.67063.70433.41332.15421.08773.92123.00914.68814.39384.3938
C41.40032.43582.83301.39292.43583.65333.73312.63332.15683.41453.92071.08824.63614.75733.00513.0051
C52.40302.78522.44061.39291.41072.32842.35611.50613.39003.87203.43152.16493.36873.40542.16912.1691
C62.78432.40391.39882.43581.41071.46422.30192.37293.87113.39212.16943.42782.26603.35313.13093.1309
C74.22683.82302.60773.65332.32841.46421.35382.35365.31214.74222.92854.49191.08582.17423.10713.1071
C84.68814.66223.67063.73312.35612.30191.35381.50665.72215.68514.19444.29482.17501.08522.16752.1675
C93.85964.27163.70432.63331.50612.37292.35361.50664.77355.35724.54072.93993.40522.29311.09761.0976
H101.08692.15703.41332.15683.39003.87115.31215.72214.77352.47394.31072.48806.10986.78885.10465.1046
H112.15621.08692.15423.41453.87203.39214.74225.68515.35722.47392.48794.31065.14316.74495.91735.9173
H123.42112.16301.08773.92073.43152.16942.92854.19444.54074.31072.48795.00892.92765.10255.24815.2481
H132.16373.42143.92121.08822.16493.42784.49194.29482.93992.48804.31065.00895.53115.21873.06673.0667
H145.02734.37083.00914.63613.36872.26601.08582.17503.40526.10985.14312.92765.53112.62274.10514.1051
H155.76515.73544.68814.75733.40543.35312.17421.08522.29316.78886.74495.10255.21872.62272.69222.6922
H164.29044.84594.39383.00512.16913.13093.10712.16751.09765.10465.91735.24813.06674.10512.69221.7629
H174.29044.84594.39383.00512.16913.13093.10712.16751.09765.10465.91735.24813.06674.10512.69221.7629

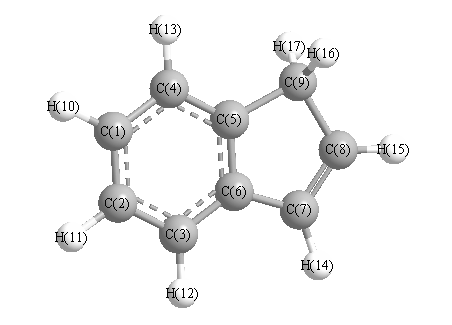
picture of Indene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.799 C1 C2 H11 119.493
C1 C4 C5 118.699 C1 C4 H13 120.275
C2 C1 C4 120.698 C2 C1 H10 119.571
C2 C3 C6 118.548 C2 C3 H12 120.473
C3 C2 H11 119.708 C3 C6 C5 120.614
C3 C6 C7 131.227 C4 C1 H10 119.731
C4 C5 C6 120.641 C4 C5 C9 130.520
C5 C4 H13 121.026 C5 C6 C7 108.159
C5 C9 C8 102.902 C5 C9 H16 111.884
C5 C9 H17 111.884 C6 C3 H12 120.979
C6 C5 C9 108.839 C6 C7 C8 109.479
C6 C7 H14 124.741 C7 C8 C9 110.621
C7 C8 H15 125.751 C8 C7 H14 125.780
C8 C9 H16 111.712 C8 C9 H17 111.712
C9 C8 H15 123.628 H16 C9 H17 106.849
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability