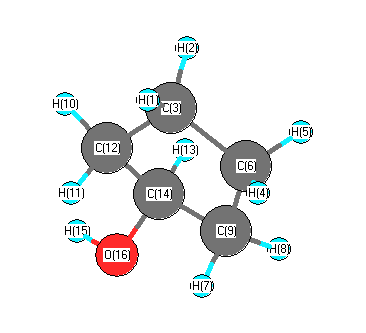
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3817 |
3675 |
11.10 |
|
|
|
| 2 |
A |
3117 |
3001 |
85.30 |
|
|
|
| 3 |
A |
3103 |
2988 |
19.59 |
|
|
|
| 4 |
A |
3089 |
2974 |
8.40 |
|
|
|
| 5 |
A |
3083 |
2968 |
48.93 |
|
|
|
| 6 |
A |
3066 |
2952 |
37.97 |
|
|
|
| 7 |
A |
3055 |
2941 |
35.27 |
|
|
|
| 8 |
A |
3045 |
2931 |
24.54 |
|
|
|
| 9 |
A |
3038 |
2925 |
36.80 |
|
|
|
| 10 |
A |
2966 |
2855 |
57.08 |
|
|
|
| 11 |
A |
1532 |
1475 |
3.80 |
|
|
|
| 12 |
A |
1506 |
1450 |
7.35 |
|
|
|
| 13 |
A |
1503 |
1447 |
3.69 |
|
|
|
| 14 |
A |
1493 |
1437 |
1.58 |
|
|
|
| 15 |
A |
1445 |
1391 |
2.25 |
|
|
|
| 16 |
A |
1385 |
1334 |
35.91 |
|
|
|
| 17 |
A |
1345 |
1295 |
0.33 |
|
|
|
| 18 |
A |
1335 |
1285 |
6.94 |
|
|
|
| 19 |
A |
1326 |
1276 |
7.91 |
|
|
|
| 20 |
A |
1306 |
1258 |
5.54 |
|
|
|
| 21 |
A |
1284 |
1236 |
18.13 |
|
|
|
| 22 |
A |
1272 |
1225 |
4.98 |
|
|
|
| 23 |
A |
1218 |
1172 |
25.08 |
|
|
|
| 24 |
A |
1208 |
1163 |
0.34 |
|
|
|
| 25 |
A |
1198 |
1153 |
3.53 |
|
|
|
| 26 |
A |
1117 |
1076 |
64.60 |
|
|
|
| 27 |
A |
1089 |
1049 |
33.95 |
|
|
|
| 28 |
A |
1060 |
1021 |
4.55 |
|
|
|
| 29 |
A |
995 |
958 |
1.18 |
|
|
|
| 30 |
A |
978 |
941 |
7.93 |
|
|
|
| 31 |
A |
962 |
926 |
6.48 |
|
|
|
| 32 |
A |
906 |
873 |
0.14 |
|
|
|
| 33 |
A |
890 |
856 |
0.09 |
|
|
|
| 34 |
A |
820 |
790 |
1.30 |
|
|
|
| 35 |
A |
770 |
741 |
2.01 |
|
|
|
| 36 |
A |
623 |
600 |
0.61 |
|
|
|
| 37 |
A |
542 |
521 |
6.43 |
|
|
|
| 38 |
A |
469 |
451 |
8.32 |
|
|
|
| 39 |
A |
360 |
347 |
24.66 |
|
|
|
| 40 |
A |
302 |
291 |
119.92 |
|
|
|
| 41 |
A |
180 |
173 |
0.90 |
|
|
|
| 42 |
A |
28 |
27 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31912.4 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 30722.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.227 |
|
|
|
| 2 |
H |
0.220 |
|
|
|
| 3 |
C |
-0.457 |
|
|
|
| 4 |
H |
0.225 |
|
|
|
| 5 |
H |
0.222 |
|
|
|
| 6 |
C |
-0.432 |
|
|
|
| 7 |
H |
0.234 |
|
|
|
| 8 |
H |
0.227 |
|
|
|
| 9 |
C |
-0.454 |
|
|
|
| 10 |
H |
0.211 |
|
|
|
| 11 |
H |
0.223 |
|
|
|
| 12 |
C |
-0.454 |
|
|
|
| 13 |
H |
0.188 |
|
|
|
| 14 |
C |
0.005 |
|
|
|
| 15 |
H |
0.386 |
|
|
|
| 16 |
O |
-0.571 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.494 |
1.256 |
0.879 |
1.610 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.678 |
3.932 |
2.228 |
| y |
3.932 |
-37.748 |
0.278 |
| z |
2.228 |
0.278 |
-37.599 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.004 |
3.932 |
2.228 |
| y |
3.932 |
0.390 |
0.278 |
| z |
2.228 |
0.278 |
0.614 |
|
| Polar |
|---|
| 3z2-r2 | 1.228 |
|---|
| x2-y2 | -0.930 |
|---|
| xy | 3.932 |
|---|
| xz | 2.228 |
|---|
| yz | 0.278 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
165.780 |
| (<r2>)1/2 |
12.876 |