Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -342.605360 |
| Energy at 298.15K | -342.610124 |
| HF Energy | -341.423938 |
| Nuclear repulsion energy | 271.380201 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3316 |
3141 |
0.06 |
140.98 |
0.15 |
0.26 |
| 2 |
A' |
3295 |
3121 |
0.07 |
57.55 |
0.40 |
0.57 |
| 3 |
A' |
3282 |
3108 |
1.22 |
71.66 |
0.63 |
0.77 |
| 4 |
A' |
2974 |
2817 |
109.90 |
142.47 |
0.29 |
0.45 |
| 5 |
A' |
1761 |
1668 |
190.20 |
121.09 |
0.31 |
0.48 |
| 6 |
A' |
1595 |
1510 |
6.01 |
0.65 |
0.70 |
0.82 |
| 7 |
A' |
1520 |
1440 |
34.02 |
79.76 |
0.32 |
0.48 |
| 8 |
A' |
1453 |
1376 |
23.90 |
39.37 |
0.35 |
0.52 |
| 9 |
A' |
1410 |
1335 |
1.25 |
14.25 |
0.11 |
0.20 |
| 10 |
A' |
1346 |
1274 |
22.55 |
7.54 |
0.35 |
0.52 |
| 11 |
A' |
1262 |
1195 |
13.16 |
5.60 |
0.43 |
0.60 |
| 12 |
A' |
1222 |
1157 |
19.19 |
5.06 |
0.10 |
0.18 |
| 13 |
A' |
1117 |
1058 |
8.20 |
8.14 |
0.36 |
0.52 |
| 14 |
A' |
1059 |
1003 |
36.07 |
4.18 |
0.27 |
0.43 |
| 15 |
A' |
958 |
907 |
15.77 |
3.95 |
0.13 |
0.23 |
| 16 |
A' |
898 |
850 |
7.75 |
6.60 |
0.72 |
0.84 |
| 17 |
A' |
774 |
733 |
64.46 |
1.96 |
0.71 |
0.83 |
| 18 |
A' |
500 |
474 |
1.22 |
6.23 |
0.30 |
0.46 |
| 19 |
A' |
201 |
191 |
4.90 |
1.02 |
0.67 |
0.80 |
| 20 |
A" |
1011 |
958 |
0.05 |
3.80 |
0.75 |
0.86 |
| 21 |
A" |
803 |
760 |
0.96 |
0.44 |
0.75 |
0.86 |
| 22 |
A" |
789 |
747 |
20.05 |
0.09 |
0.75 |
0.86 |
| 23 |
A" |
746 |
707 |
67.52 |
0.19 |
0.75 |
0.86 |
| 24 |
A" |
651 |
616 |
4.72 |
1.81 |
0.75 |
0.86 |
| 25 |
A" |
579 |
549 |
3.01 |
0.04 |
0.75 |
0.86 |
| 26 |
A" |
284 |
269 |
12.85 |
0.61 |
0.75 |
0.86 |
| 27 |
A" |
131 |
124 |
1.22 |
1.54 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17468.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16544.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.504 |
-0.893 |
0.000 |
| C2 |
-0.553 |
-1.736 |
0.000 |
| C3 |
-1.738 |
-1.043 |
0.000 |
| C4 |
0.000 |
0.367 |
0.000 |
| C5 |
-1.380 |
0.331 |
0.000 |
| C6 |
0.925 |
1.498 |
0.000 |
| O7 |
2.138 |
1.404 |
0.000 |
| H8 |
-0.309 |
-2.788 |
0.000 |
| H9 |
-2.730 |
-1.471 |
0.000 |
| H10 |
-2.039 |
1.189 |
0.000 |
| H11 |
0.418 |
2.483 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3521 | 2.2466 | 1.3571 | 2.2460 | 2.4272 | 2.8188 | 2.0621 | 3.2848 | 3.2855 | 3.3773 |
C2 | 1.3521 | | 1.3731 | 2.1754 | 2.2265 | 3.5558 | 4.1355 | 1.0797 | 2.1930 | 3.2807 | 4.3301 | C3 | 2.2466 | 1.3731 | | 2.2383 | 1.4196 | 3.6805 | 4.5837 | 2.2558 | 1.0803 | 2.2517 | 4.1330 | C4 | 1.3571 | 2.1754 | 2.2383 | | 1.3803 | 1.4605 | 2.3760 | 3.1707 | 3.2912 | 2.1977 | 2.1568 | C5 | 2.2460 | 2.2265 | 1.4196 | 1.3803 | | 2.5834 | 3.6780 | 3.2978 | 2.2514 | 1.0815 | 2.8044 | C6 | 2.4272 | 3.5558 | 3.6805 | 1.4605 | 2.5834 | | 1.2167 | 4.4598 | 4.7086 | 2.9797 | 1.1088 | O7 | 2.8188 | 4.1355 | 4.5837 | 2.3760 | 3.6780 | 1.2167 | | 4.8537 | 5.6534 | 4.1823 | 2.0314 | H8 | 2.0621 | 1.0797 | 2.2558 | 3.1707 | 3.2978 | 4.4598 | 4.8537 | | 2.7563 | 4.3367 | 5.3214 | H9 | 3.2848 | 2.1930 | 1.0803 | 3.2912 | 2.2514 | 4.7086 | 5.6534 | 2.7563 | | 2.7480 | 5.0541 | H10 | 3.2855 | 3.2807 | 2.2517 | 2.1977 | 1.0815 | 2.9797 | 4.1823 | 4.3367 | 2.7480 | | 2.7765 | H11 | 3.3773 | 4.3301 | 4.1330 | 2.1568 | 2.8044 | 1.1088 | 2.0314 | 5.3214 | 5.0541 | 2.7765 | |
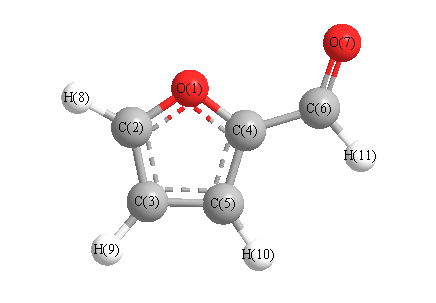 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.052 |
|
O1 |
C2 |
H8 |
115.532 |
| O1 |
C4 |
C5 |
110.260 |
|
O1 |
C4 |
C6 |
118.915 |
| C2 |
O1 |
C4 |
106.826 |
|
C2 |
C3 |
C5 |
105.727 |
| C2 |
C3 |
H9 |
126.318 |
|
C3 |
C2 |
H8 |
133.416 |
| C3 |
C5 |
C4 |
106.134 |
|
C3 |
C5 |
H10 |
127.869 |
| C4 |
C5 |
H10 |
125.997 |
|
C4 |
C6 |
O7 |
124.875 |
| C4 |
C6 |
H11 |
113.457 |
|
C5 |
C3 |
H9 |
127.955 |
| C5 |
C4 |
C6 |
130.825 |
|
O7 |
C6 |
H11 |
121.668 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -342.606885 |
| Energy at 298.15K | -342.611625 |
| HF Energy | -341.425932 |
| Nuclear repulsion energy | 270.542216 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3317 |
3141 |
0.11 |
146.95 |
0.15 |
0.27 |
| 2 |
A' |
3300 |
3126 |
0.65 |
37.55 |
0.24 |
0.39 |
| 3 |
A' |
3288 |
3114 |
0.59 |
79.16 |
0.75 |
0.85 |
| 4 |
A' |
2996 |
2838 |
99.12 |
150.80 |
0.29 |
0.45 |
| 5 |
A' |
1756 |
1663 |
162.13 |
106.87 |
0.34 |
0.51 |
| 6 |
A' |
1606 |
1521 |
40.77 |
18.80 |
0.31 |
0.47 |
| 7 |
A' |
1515 |
1435 |
60.00 |
150.27 |
0.29 |
0.45 |
| 8 |
A' |
1462 |
1385 |
2.13 |
5.30 |
0.74 |
0.85 |
| 9 |
A' |
1411 |
1337 |
12.35 |
23.46 |
0.38 |
0.55 |
| 10 |
A' |
1294 |
1226 |
9.47 |
7.48 |
0.15 |
0.27 |
| 11 |
A' |
1276 |
1208 |
27.97 |
2.21 |
0.75 |
0.86 |
| 12 |
A' |
1204 |
1141 |
11.62 |
5.74 |
0.13 |
0.23 |
| 13 |
A' |
1122 |
1062 |
15.58 |
10.57 |
0.37 |
0.54 |
| 14 |
A' |
1053 |
997 |
36.33 |
2.31 |
0.21 |
0.34 |
| 15 |
A' |
970 |
919 |
7.00 |
3.59 |
0.16 |
0.27 |
| 16 |
A' |
898 |
850 |
5.33 |
5.43 |
0.75 |
0.86 |
| 17 |
A' |
761 |
721 |
66.08 |
2.51 |
0.75 |
0.86 |
| 18 |
A' |
500 |
473 |
0.76 |
5.52 |
0.37 |
0.54 |
| 19 |
A' |
203 |
193 |
7.01 |
0.40 |
0.41 |
0.58 |
| 20 |
A" |
1014 |
961 |
0.00 |
4.98 |
0.75 |
0.86 |
| 21 |
A" |
811 |
768 |
0.02 |
0.43 |
0.75 |
0.86 |
| 22 |
A" |
796 |
754 |
29.14 |
0.12 |
0.75 |
0.86 |
| 23 |
A" |
735 |
696 |
58.65 |
0.13 |
0.75 |
0.86 |
| 24 |
A" |
640 |
606 |
5.92 |
1.04 |
0.75 |
0.86 |
| 25 |
A" |
574 |
544 |
2.86 |
0.05 |
0.75 |
0.86 |
| 26 |
A" |
246 |
233 |
13.75 |
1.84 |
0.75 |
0.86 |
| 27 |
A" |
154 |
145 |
2.39 |
1.10 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17450.8 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16527.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.255 |
-0.242 |
0.000 |
| C2 |
1.115 |
-1.588 |
0.000 |
| C3 |
-0.213 |
-1.939 |
0.000 |
| C4 |
0.000 |
0.285 |
0.000 |
| C5 |
-0.942 |
-0.721 |
0.000 |
| C6 |
-0.111 |
1.742 |
0.000 |
| O7 |
-1.177 |
2.331 |
0.000 |
| H8 |
2.034 |
-2.154 |
0.000 |
| H9 |
-0.605 |
-2.945 |
0.000 |
| H10 |
-2.012 |
-0.573 |
0.000 |
| H11 |
0.856 |
2.281 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3531 | 2.2441 | 1.3616 | 2.2487 | 2.4087 | 3.5405 | 2.0648 | 3.2821 | 3.2841 | 2.5543 |
C2 | 1.3531 | | 1.3740 | 2.1799 | 2.2320 | 3.5481 | 4.5397 | 1.0795 | 2.1920 | 3.2880 | 3.8774 | C3 | 2.2441 | 1.3740 | | 2.2342 | 1.4188 | 3.6821 | 4.3770 | 2.2577 | 1.0805 | 2.2589 | 4.3533 | C4 | 1.3616 | 2.1799 | 2.2342 | | 1.3783 | 1.4608 | 2.3600 | 3.1763 | 3.2869 | 2.1872 | 2.1718 | C5 | 2.2487 | 2.2320 | 1.4188 | 1.3783 | | 2.5995 | 3.0610 | 3.3029 | 2.2494 | 1.0807 | 3.4995 | C6 | 2.4087 | 3.5481 | 3.6821 | 1.4608 | 2.5995 | | 1.2181 | 4.4475 | 4.7134 | 2.9952 | 1.1073 | O7 | 3.5405 | 4.5397 | 4.3770 | 2.3600 | 3.0610 | 1.2181 | | 5.5160 | 5.3070 | 3.0209 | 2.0341 | H8 | 2.0648 | 1.0795 | 2.2577 | 3.1763 | 3.3029 | 4.4475 | 5.5160 | | 2.7559 | 4.3444 | 4.5890 | H9 | 3.2821 | 2.1920 | 1.0805 | 3.2869 | 2.2494 | 4.7134 | 5.3070 | 2.7559 | | 2.7584 | 5.4271 | H10 | 3.2841 | 3.2880 | 2.2589 | 2.1872 | 1.0807 | 2.9952 | 3.0209 | 4.3444 | 2.7584 | | 4.0461 | H11 | 2.5543 | 3.8774 | 4.3533 | 2.1718 | 3.4995 | 1.1073 | 2.0341 | 4.5890 | 5.4271 | 4.0461 | |
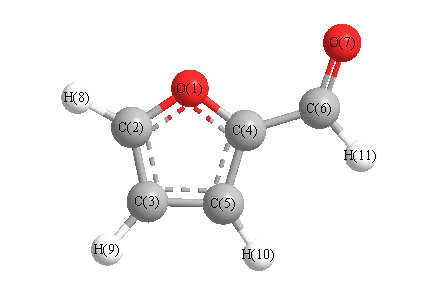 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.745 |
|
O1 |
C2 |
H8 |
115.707 |
| O1 |
C4 |
C5 |
110.316 |
|
O1 |
C4 |
C6 |
117.125 |
| C2 |
O1 |
C4 |
106.836 |
|
C2 |
C3 |
C5 |
106.091 |
| C2 |
C3 |
H9 |
126.104 |
|
C3 |
C2 |
H8 |
133.548 |
| C3 |
C5 |
C4 |
106.013 |
|
C3 |
C5 |
H10 |
128.805 |
| C4 |
C5 |
H10 |
125.182 |
|
C4 |
C6 |
O7 |
123.256 |
| C4 |
C6 |
H11 |
114.794 |
|
C5 |
C3 |
H9 |
127.806 |
| C5 |
C4 |
C6 |
132.560 |
|
O7 |
C6 |
H11 |
121.950 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
