Jump to
S1C2
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -343.039380 |
| Energy at 298.15K | -343.044261 |
| HF Energy | -343.039380 |
| Nuclear repulsion energy | 272.638166 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3303 |
3166 |
0.64 |
142.14 |
0.15 |
0.26 |
| 2 |
A' |
3281 |
3145 |
0.34 |
54.91 |
0.36 |
0.53 |
| 3 |
A' |
3269 |
3134 |
1.75 |
79.85 |
0.71 |
0.83 |
| 4 |
A' |
2928 |
2806 |
122.26 |
143.40 |
0.32 |
0.49 |
| 5 |
A' |
1820 |
1744 |
343.59 |
128.22 |
0.34 |
0.51 |
| 6 |
A' |
1633 |
1565 |
8.41 |
10.05 |
0.44 |
0.61 |
| 7 |
A' |
1534 |
1470 |
51.08 |
98.10 |
0.31 |
0.47 |
| 8 |
A' |
1456 |
1395 |
40.84 |
40.61 |
0.41 |
0.58 |
| 9 |
A' |
1409 |
1350 |
0.27 |
14.14 |
0.11 |
0.20 |
| 10 |
A' |
1325 |
1270 |
48.10 |
6.89 |
0.40 |
0.58 |
| 11 |
A' |
1261 |
1209 |
2.20 |
9.81 |
0.43 |
0.60 |
| 12 |
A' |
1217 |
1166 |
8.24 |
3.51 |
0.12 |
0.22 |
| 13 |
A' |
1127 |
1080 |
12.59 |
11.17 |
0.30 |
0.46 |
| 14 |
A' |
1051 |
1008 |
44.84 |
4.22 |
0.34 |
0.51 |
| 15 |
A' |
965 |
925 |
18.01 |
5.53 |
0.11 |
0.20 |
| 16 |
A' |
907 |
870 |
10.45 |
7.34 |
0.73 |
0.85 |
| 17 |
A' |
776 |
743 |
67.92 |
1.44 |
0.73 |
0.84 |
| 18 |
A' |
505 |
484 |
1.09 |
7.65 |
0.32 |
0.48 |
| 19 |
A' |
199 |
191 |
5.57 |
0.97 |
0.68 |
0.81 |
| 20 |
A" |
1016 |
974 |
0.01 |
4.11 |
0.75 |
0.86 |
| 21 |
A" |
894 |
857 |
1.44 |
1.06 |
0.75 |
0.86 |
| 22 |
A" |
844 |
809 |
6.26 |
0.08 |
0.75 |
0.86 |
| 23 |
A" |
780 |
747 |
70.41 |
0.08 |
0.75 |
0.86 |
| 24 |
A" |
666 |
639 |
0.98 |
2.30 |
0.75 |
0.86 |
| 25 |
A" |
613 |
588 |
8.99 |
0.17 |
0.75 |
0.86 |
| 26 |
A" |
298 |
286 |
13.44 |
0.57 |
0.75 |
0.86 |
| 27 |
A" |
137 |
131 |
1.15 |
1.60 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17606.0 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 16875.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.509 |
-0.894 |
0.000 |
| C2 |
-0.533 |
-1.737 |
0.000 |
| C3 |
-1.716 |
-1.061 |
0.000 |
| C4 |
0.000 |
0.364 |
0.000 |
| C5 |
-1.368 |
0.313 |
0.000 |
| C6 |
0.904 |
1.501 |
0.000 |
| O7 |
2.108 |
1.439 |
0.000 |
| H8 |
-0.284 |
-2.787 |
0.000 |
| H9 |
-2.705 |
-1.494 |
0.000 |
| H10 |
-2.035 |
1.163 |
0.000 |
| H11 |
0.370 |
2.476 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3402 | 2.2315 | 1.3568 | 2.2312 | 2.4269 | 2.8280 | 2.0522 | 3.2698 | 3.2716 | 3.3725 |
C2 | 1.3402 | | 1.3624 | 2.1671 | 2.2130 | 3.5418 | 4.1301 | 1.0792 | 2.1856 | 3.2654 | 4.3082 | C3 | 2.2315 | 1.3624 | | 2.2307 | 1.4176 | 3.6643 | 4.5687 | 2.2426 | 1.0796 | 2.2466 | 4.1066 | C4 | 1.3568 | 2.1671 | 2.2307 | | 1.3687 | 1.4522 | 2.3660 | 3.1633 | 3.2819 | 2.1866 | 2.1441 | C5 | 2.2312 | 2.2130 | 1.4176 | 1.3687 | | 2.5632 | 3.6533 | 3.2836 | 2.2483 | 1.0807 | 2.7747 | C6 | 2.4269 | 3.5418 | 3.6643 | 1.4522 | 2.5632 | | 1.2056 | 4.4488 | 4.6897 | 2.9585 | 1.1113 | O7 | 2.8280 | 4.1301 | 4.5687 | 2.3660 | 3.6533 | 1.2056 | | 4.8554 | 5.6363 | 4.1524 | 2.0231 | H8 | 2.0522 | 1.0792 | 2.2426 | 3.1633 | 3.2836 | 4.4488 | 4.8554 | | 2.7446 | 4.3203 | 5.3030 | H9 | 3.2698 | 2.1856 | 1.0796 | 3.2819 | 2.2483 | 4.6897 | 5.6363 | 2.7446 | | 2.7400 | 5.0220 | H10 | 3.2716 | 3.2654 | 2.2466 | 2.1866 | 1.0807 | 2.9585 | 4.1524 | 4.3203 | 2.7400 | | 2.7410 | H11 | 3.3725 | 4.3082 | 4.1066 | 2.1441 | 2.7747 | 1.1113 | 2.0231 | 5.3030 | 5.0220 | 2.7410 | |
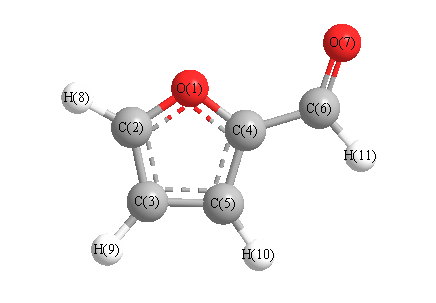 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.313 |
|
O1 |
C2 |
H8 |
115.618 |
| O1 |
C4 |
C5 |
109.898 |
|
O1 |
C4 |
C6 |
119.487 |
| C2 |
O1 |
C4 |
106.934 |
|
C2 |
C3 |
C5 |
105.488 |
| C2 |
C3 |
H9 |
126.628 |
|
C3 |
C2 |
H8 |
133.069 |
| C3 |
C5 |
C4 |
106.367 |
|
C3 |
C5 |
H10 |
127.604 |
| C4 |
C5 |
H10 |
126.030 |
|
C4 |
C6 |
O7 |
125.542 |
| C4 |
C6 |
H11 |
112.846 |
|
C5 |
C3 |
H9 |
127.884 |
| C5 |
C4 |
C6 |
130.615 |
|
O7 |
C6 |
H11 |
121.612 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.216 |
|
|
|
| 2 |
C |
-0.020 |
|
|
|
| 3 |
C |
-0.281 |
|
|
|
| 4 |
C |
0.142 |
|
|
|
| 5 |
C |
-0.255 |
|
|
|
| 6 |
C |
0.067 |
|
|
|
| 7 |
O |
-0.294 |
|
|
|
| 8 |
H |
0.235 |
|
|
|
| 9 |
H |
0.225 |
|
|
|
| 10 |
H |
0.230 |
|
|
|
| 11 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.938 |
-1.185 |
0.000 |
4.112 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.454 |
-2.927 |
0.000 |
| y |
-2.927 |
-33.460 |
0.000 |
| z |
0.000 |
0.000 |
-41.208 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.120 |
-2.927 |
0.000 |
| y |
-2.927 |
8.872 |
0.000 |
| z |
0.000 |
0.000 |
-2.751 |
|
| Polar |
|---|
| 3z2-r2 | -5.503 |
|---|
| x2-y2 | -9.995 |
|---|
| xy | -2.927 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.682 |
1.892 |
0.000 |
| y |
1.892 |
10.546 |
0.000 |
| z |
0.000 |
0.000 |
3.986 |
<r2> (average value of r
2) Å
2
| <r2> |
189.566 |
| (<r2>)1/2 |
13.768 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -343.041067 |
| Energy at 298.15K | -343.045940 |
| HF Energy | -343.041067 |
| Nuclear repulsion energy | 271.833603 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3303 |
3166 |
0.77 |
151.12 |
0.15 |
0.26 |
| 2 |
A' |
3290 |
3154 |
0.06 |
30.41 |
0.34 |
0.51 |
| 3 |
A' |
3274 |
3138 |
1.27 |
86.87 |
0.71 |
0.83 |
| 4 |
A' |
2948 |
2825 |
114.18 |
155.01 |
0.33 |
0.49 |
| 5 |
A' |
1814 |
1738 |
287.61 |
97.77 |
0.36 |
0.53 |
| 6 |
A' |
1640 |
1572 |
57.65 |
26.55 |
0.37 |
0.54 |
| 7 |
A' |
1525 |
1461 |
116.19 |
168.08 |
0.29 |
0.45 |
| 8 |
A' |
1466 |
1405 |
3.24 |
9.79 |
0.72 |
0.84 |
| 9 |
A' |
1414 |
1356 |
14.18 |
23.17 |
0.38 |
0.55 |
| 10 |
A' |
1282 |
1229 |
29.69 |
4.80 |
0.34 |
0.51 |
| 11 |
A' |
1261 |
1208 |
0.73 |
3.04 |
0.48 |
0.65 |
| 12 |
A' |
1203 |
1153 |
15.05 |
4.82 |
0.16 |
0.27 |
| 13 |
A' |
1132 |
1085 |
26.13 |
14.42 |
0.29 |
0.45 |
| 14 |
A' |
1045 |
1002 |
42.33 |
2.81 |
0.30 |
0.46 |
| 15 |
A' |
978 |
938 |
6.15 |
5.48 |
0.15 |
0.26 |
| 16 |
A' |
907 |
869 |
7.07 |
6.44 |
0.75 |
0.86 |
| 17 |
A' |
764 |
732 |
74.89 |
2.62 |
0.65 |
0.79 |
| 18 |
A' |
503 |
482 |
1.06 |
6.18 |
0.38 |
0.55 |
| 19 |
A' |
204 |
195 |
6.98 |
0.34 |
0.41 |
0.58 |
| 20 |
A" |
1025 |
982 |
0.04 |
5.54 |
0.75 |
0.86 |
| 21 |
A" |
904 |
866 |
2.16 |
1.26 |
0.75 |
0.86 |
| 22 |
A" |
851 |
816 |
7.78 |
0.00 |
0.75 |
0.86 |
| 23 |
A" |
778 |
746 |
65.17 |
0.21 |
0.75 |
0.86 |
| 24 |
A" |
656 |
629 |
0.67 |
1.01 |
0.75 |
0.86 |
| 25 |
A" |
615 |
590 |
12.58 |
0.37 |
0.75 |
0.86 |
| 26 |
A" |
258 |
247 |
14.30 |
1.75 |
0.75 |
0.86 |
| 27 |
A" |
162 |
155 |
2.15 |
1.19 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17599.6 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 16869.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.248 |
-0.261 |
0.000 |
| C2 |
1.090 |
-1.593 |
0.000 |
| C3 |
-0.231 |
-1.929 |
0.000 |
| C4 |
0.000 |
0.286 |
0.000 |
| C5 |
-0.942 |
-0.704 |
0.000 |
| C6 |
-0.096 |
1.735 |
0.000 |
| O7 |
-1.142 |
2.339 |
0.000 |
| H8 |
2.000 |
-2.173 |
0.000 |
| H9 |
-0.638 |
-2.930 |
0.000 |
| H10 |
-2.011 |
-0.547 |
0.000 |
| H11 |
0.883 |
2.258 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3410 | 2.2295 | 1.3622 | 2.2342 | 2.4063 | 3.5312 | 2.0542 | 3.2676 | 3.2707 | 2.5455 |
C2 | 1.3410 | | 1.3632 | 2.1718 | 2.2178 | 3.5328 | 4.5209 | 1.0791 | 2.1846 | 3.2718 | 3.8565 | C3 | 2.2295 | 1.3632 | | 2.2274 | 1.4168 | 3.6670 | 4.3645 | 2.2442 | 1.0798 | 2.2530 | 4.3333 | C4 | 1.3622 | 2.1718 | 2.2274 | | 1.3667 | 1.4524 | 2.3493 | 3.1691 | 3.2783 | 2.1764 | 2.1608 | C5 | 2.2342 | 2.2178 | 1.4168 | 1.3667 | | 2.5817 | 3.0495 | 3.2882 | 2.2464 | 1.0797 | 3.4793 | C6 | 2.4063 | 3.5328 | 3.6670 | 1.4524 | 2.5817 | | 1.2076 | 4.4343 | 4.6962 | 2.9790 | 1.1100 | O7 | 3.5312 | 4.5209 | 4.3645 | 2.3493 | 3.0495 | 1.2076 | | 5.4976 | 5.2927 | 3.0142 | 2.0263 | H8 | 2.0542 | 1.0791 | 2.2442 | 3.1691 | 3.2882 | 4.4343 | 5.4976 | | 2.7442 | 4.3271 | 4.5694 | H9 | 3.2676 | 2.1846 | 1.0798 | 3.2783 | 2.2464 | 4.6962 | 5.2927 | 2.7442 | | 2.7495 | 5.4062 | H10 | 3.2707 | 3.2718 | 2.2530 | 2.1764 | 1.0797 | 2.9790 | 3.0142 | 4.3271 | 2.7495 | | 4.0302 | H11 | 2.5455 | 3.8565 | 4.3333 | 2.1608 | 3.4793 | 1.1100 | 2.0263 | 4.5694 | 5.4062 | 4.0302 | |
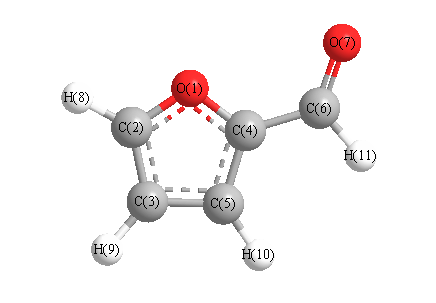 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.063 |
|
O1 |
C2 |
H8 |
115.741 |
| O1 |
C4 |
C5 |
109.916 |
|
O1 |
C4 |
C6 |
117.468 |
| C2 |
O1 |
C4 |
106.909 |
|
C2 |
C3 |
C5 |
105.822 |
| C2 |
C3 |
H9 |
126.426 |
|
C3 |
C2 |
H8 |
133.196 |
| C3 |
C5 |
C4 |
106.290 |
|
C3 |
C5 |
H10 |
128.466 |
| C4 |
C5 |
H10 |
125.244 |
|
C4 |
C6 |
O7 |
123.800 |
| C4 |
C6 |
H11 |
114.321 |
|
C5 |
C3 |
H9 |
127.752 |
| C5 |
C4 |
C6 |
132.616 |
|
O7 |
C6 |
H11 |
121.879 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.239 |
|
|
|
| 2 |
C |
-0.023 |
|
|
|
| 3 |
C |
-0.278 |
|
|
|
| 4 |
C |
0.135 |
|
|
|
| 5 |
C |
-0.219 |
|
|
|
| 6 |
C |
0.053 |
|
|
|
| 7 |
O |
-0.301 |
|
|
|
| 8 |
H |
0.234 |
|
|
|
| 9 |
H |
0.225 |
|
|
|
| 10 |
H |
0.236 |
|
|
|
| 11 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.527 |
-3.218 |
0.000 |
3.562 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.270 |
3.831 |
0.000 |
| y |
3.831 |
-39.494 |
0.000 |
| z |
0.000 |
0.000 |
-41.177 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.065 |
3.831 |
0.000 |
| y |
3.831 |
0.230 |
0.000 |
| z |
0.000 |
0.000 |
-2.295 |
|
| Polar |
|---|
| 3z2-r2 | -4.590 |
|---|
| x2-y2 | 1.224 |
|---|
| xy | 3.831 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.902 |
-1.100 |
0.000 |
| y |
-1.100 |
12.319 |
0.000 |
| z |
0.000 |
0.000 |
3.963 |
<r2> (average value of r
2) Å
2
| <r2> |
191.801 |
| (<r2>)1/2 |
13.849 |
