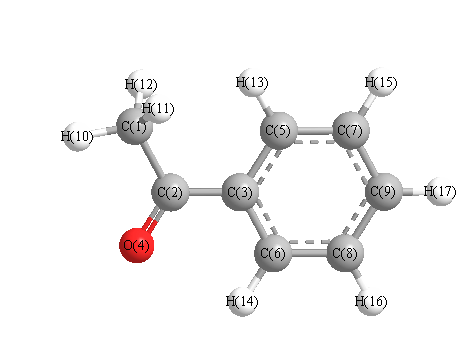
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3214 |
3094 |
7.33 |
|
|
|
| 2 |
A' |
3204 |
3085 |
16.29 |
|
|
|
| 3 |
A' |
3195 |
3076 |
26.04 |
|
|
|
| 4 |
A' |
3185 |
3067 |
8.69 |
|
|
|
| 5 |
A' |
3175 |
3056 |
0.07 |
|
|
|
| 6 |
A' |
3163 |
3045 |
13.84 |
|
|
|
| 7 |
A' |
3047 |
2933 |
2.89 |
|
|
|
| 8 |
A' |
1782 |
1715 |
181.23 |
|
|
|
| 9 |
A' |
1660 |
1598 |
21.31 |
|
|
|
| 10 |
A' |
1640 |
1579 |
8.35 |
|
|
|
| 11 |
A' |
1531 |
1473 |
0.79 |
|
|
|
| 12 |
A' |
1487 |
1432 |
20.04 |
|
|
|
| 13 |
A' |
1480 |
1425 |
12.47 |
|
|
|
| 14 |
A' |
1393 |
1341 |
69.12 |
|
|
|
| 15 |
A' |
1373 |
1322 |
3.22 |
|
|
|
| 16 |
A' |
1343 |
1293 |
2.80 |
|
|
|
| 17 |
A' |
1288 |
1240 |
180.66 |
|
|
|
| 18 |
A' |
1204 |
1159 |
13.74 |
|
|
|
| 19 |
A' |
1186 |
1142 |
0.52 |
|
|
|
| 20 |
A' |
1112 |
1070 |
7.77 |
|
|
|
| 21 |
A' |
1099 |
1058 |
1.19 |
|
|
|
| 22 |
A' |
1052 |
1012 |
5.98 |
|
|
|
| 23 |
A' |
1019 |
981 |
0.81 |
|
|
|
| 24 |
A' |
963 |
927 |
28.33 |
|
|
|
| 25 |
A' |
745 |
717 |
0.82 |
|
|
|
| 26 |
A' |
629 |
606 |
1.03 |
|
|
|
| 27 |
A' |
597 |
575 |
25.94 |
|
|
|
| 28 |
A' |
465 |
448 |
0.66 |
|
|
|
| 29 |
A' |
365 |
351 |
0.75 |
|
|
|
| 30 |
A' |
215 |
207 |
4.89 |
|
|
|
| 31 |
A" |
3111 |
2995 |
10.02 |
|
|
|
| 32 |
A" |
1490 |
1434 |
12.90 |
|
|
|
| 33 |
A" |
1051 |
1012 |
1.25 |
|
|
|
| 34 |
A" |
1007 |
970 |
0.45 |
|
|
|
| 35 |
A" |
984 |
947 |
0.39 |
|
|
|
| 36 |
A" |
940 |
905 |
2.55 |
|
|
|
| 37 |
A" |
860 |
828 |
0.24 |
|
|
|
| 38 |
A" |
776 |
747 |
37.01 |
|
|
|
| 39 |
A" |
706 |
680 |
46.98 |
|
|
|
| 40 |
A" |
604 |
582 |
12.18 |
|
|
|
| 41 |
A" |
428 |
412 |
0.47 |
|
|
|
| 42 |
A" |
413 |
397 |
0.01 |
|
|
|
| 43 |
A" |
169 |
163 |
0.28 |
|
|
|
| 44 |
A" |
153 |
147 |
0.01 |
|
|
|
| 45 |
A" |
62 |
59 |
3.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30281.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 29151.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.780 |
|
|
|
| 2 |
C |
0.333 |
|
|
|
| 3 |
C |
-0.123 |
|
|
|
| 4 |
O |
-0.307 |
|
|
|
| 5 |
C |
-0.199 |
|
|
|
| 6 |
C |
-0.161 |
|
|
|
| 7 |
C |
-0.200 |
|
|
|
| 8 |
C |
-0.201 |
|
|
|
| 9 |
C |
-0.183 |
|
|
|
| 10 |
H |
0.256 |
|
|
|
| 11 |
H |
0.249 |
|
|
|
| 12 |
H |
0.249 |
|
|
|
| 13 |
H |
0.214 |
|
|
|
| 14 |
H |
0.229 |
|
|
|
| 15 |
H |
0.206 |
|
|
|
| 16 |
H |
0.209 |
|
|
|
| 17 |
H |
0.208 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.854 |
-2.407 |
0.000 |
3.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.980 |
5.421 |
0.000 |
| y |
5.421 |
-53.159 |
0.000 |
| z |
0.000 |
0.000 |
-55.348 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.274 |
5.421 |
0.000 |
| y |
5.421 |
-1.996 |
0.000 |
| z |
0.000 |
0.000 |
-5.278 |
|
| Polar |
|---|
| 3z2-r2 | -10.556 |
|---|
| x2-y2 | 6.180 |
|---|
| xy | 5.421 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.241 |
0.732 |
0.000 |
| y |
0.732 |
17.334 |
0.000 |
| z |
0.000 |
0.000 |
6.268 |
<r2> (average value of r
2) Å
2
| <r2> |
344.478 |
| (<r2>)1/2 |
18.560 |