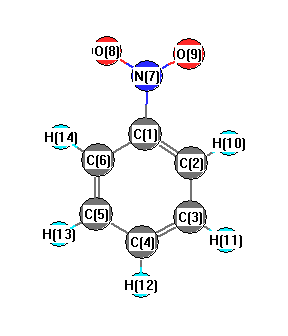
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3417 |
3090 |
2.16 |
|
|
|
| 2 |
A1 |
3375 |
3052 |
20.54 |
|
|
|
| 3 |
A1 |
3349 |
3029 |
1.71 |
|
|
|
| 4 |
A1 |
1785 |
1614 |
5.95 |
|
|
|
| 5 |
A1 |
1652 |
1494 |
12.46 |
|
|
|
| 6 |
A1 |
1632 |
1476 |
319.07 |
|
|
|
| 7 |
A1 |
1291 |
1168 |
4.06 |
|
|
|
| 8 |
A1 |
1234 |
1116 |
18.68 |
|
|
|
| 9 |
A1 |
1116 |
1010 |
3.43 |
|
|
|
| 10 |
A1 |
1088 |
984 |
0.12 |
|
|
|
| 11 |
A1 |
972 |
879 |
45.10 |
|
|
|
| 12 |
A1 |
755 |
683 |
17.66 |
|
|
|
| 13 |
A1 |
433 |
392 |
3.61 |
|
|
|
| 14 |
A2 |
1109 |
1003 |
0.00 |
|
|
|
| 15 |
A2 |
949 |
859 |
0.00 |
|
|
|
| 16 |
A2 |
455 |
412 |
0.00 |
|
|
|
| 17 |
A2 |
50 |
45 |
0.00 |
|
|
|
| 18 |
B1 |
1125 |
1018 |
0.05 |
|
|
|
| 19 |
B1 |
1074 |
972 |
4.08 |
|
|
|
| 20 |
B1 |
890 |
805 |
5.75 |
|
|
|
| 21 |
B1 |
803 |
727 |
111.40 |
|
|
|
| 22 |
B1 |
741 |
670 |
18.32 |
|
|
|
| 23 |
B1 |
486 |
440 |
0.55 |
|
|
|
| 24 |
B1 |
191 |
173 |
1.37 |
|
|
|
| 25 |
B2 |
3417 |
3090 |
0.01 |
|
|
|
| 26 |
B2 |
3364 |
3043 |
18.86 |
|
|
|
| 27 |
B2 |
1852 |
1675 |
406.53 |
|
|
|
| 28 |
B2 |
1773 |
1603 |
145.92 |
|
|
|
| 29 |
B2 |
1613 |
1459 |
0.37 |
|
|
|
| 30 |
B2 |
1460 |
1320 |
0.02 |
|
|
|
| 31 |
B2 |
1349 |
1220 |
15.83 |
|
|
|
| 32 |
B2 |
1219 |
1102 |
22.42 |
|
|
|
| 33 |
B2 |
1172 |
1060 |
0.36 |
|
|
|
| 34 |
B2 |
670 |
606 |
0.16 |
|
|
|
| 35 |
B2 |
572 |
517 |
2.48 |
|
|
|
| 36 |
B2 |
284 |
256 |
1.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24358.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 22029.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.134 |
|
|
|
| 2 |
C |
-0.183 |
|
|
|
| 3 |
C |
-0.227 |
|
|
|
| 4 |
C |
-0.181 |
|
|
|
| 5 |
C |
-0.227 |
|
|
|
| 6 |
C |
-0.183 |
|
|
|
| 7 |
N |
0.366 |
|
|
|
| 8 |
O |
-0.382 |
|
|
|
| 9 |
O |
-0.382 |
|
|
|
| 10 |
H |
0.281 |
|
|
|
| 11 |
H |
0.236 |
|
|
|
| 12 |
H |
0.234 |
|
|
|
| 13 |
H |
0.236 |
|
|
|
| 14 |
H |
0.281 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-5.034 |
5.034 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-53.666 |
0.000 |
0.000 |
| y |
0.000 |
-47.638 |
0.000 |
| z |
0.000 |
0.000 |
-55.594 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.050 |
0.000 |
0.000 |
| y |
0.000 |
6.992 |
0.000 |
| z |
0.000 |
0.000 |
-4.942 |
|
| Polar |
|---|
| 3z2-r2 | -9.884 |
|---|
| x2-y2 | -6.028 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |