return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Dihydro-3-(2H)-thiophenone)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-624.813740
Energy at 298.15K-624.821182
HF Energy-624.206248
Nuclear repulsion energy285.659373
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3184 3029 4.42      
2 A 3167 3013 3.61      
3 A 3166 3012 1.11      
4 A 3118 2966 16.04      
5 A 3106 2955 9.15      
6 A 3099 2948 2.72      
7 A 1662 1581 68.48      
8 A 1577 1500 3.43      
9 A 1545 1469 7.95      
10 A 1541 1466 10.03      
11 A 1375 1308 4.61      
12 A 1359 1293 12.07      
13 A 1297 1233 6.85      
14 A 1256 1195 2.35      
15 A 1219 1160 0.45      
16 A 1172 1115 3.04      
17 A 1089 1036 77.66      
18 A 1028 978 2.08      
19 A 991 943 11.07      
20 A 918 873 9.00      
21 A 865 823 2.30      
22 A 785 747 1.80      
23 A 711 676 4.99      
24 A 685 651 0.75      
25 A 568 541 2.53      
26 A 490 466 2.93      
27 A 451 429 0.13      
28 A 427 406 6.59      
29 A 201 191 2.39      
30 A 52 49 9.38      

Unscaled Zero Point Vibrational Energy (zpe) 21050.7 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20025.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.18601 0.07991 0.05921

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.187 -0.085 -0.019
C2 -0.512 1.285 -0.239
H3 -1.113 2.068 0.233
H4 -0.467 1.470 -1.321
C5 0.917 1.188 0.330
H6 0.913 1.292 1.419
H7 1.590 1.931 -0.103
C8 -0.124 -1.218 0.143
S9 1.522 -0.489 -0.144
H10 -0.327 -2.007 -0.587
O11 -2.413 -0.288 0.027
H12 -0.210 -1.640 1.150

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 H6 H7 C8 S9 H10 O11 H12
C11.54292.16882.15172.48352.89333.43211.56192.74122.18101.24362.1767
C21.54291.09371.09781.54192.18652.20332.56232.70113.31592.48113.2524
H32.16881.09371.78572.21482.47212.72693.43303.69084.23042.69813.9247
H42.15171.09781.78572.17243.07222.43373.07993.02943.55632.94813.9801
C52.48351.54192.21482.17241.09371.09202.62861.84463.54933.65473.1528
H62.89332.18652.47213.07221.09371.78413.00112.44644.05543.93603.1512
H73.43212.20332.72692.43371.09201.78413.59402.42174.40654.57814.1908
C81.56192.56233.43303.07992.62863.00113.59401.82281.09362.47341.0956
S92.74122.70113.69083.02941.84462.44642.42171.82282.43283.94332.4491
H102.18103.31594.23043.55633.54934.05544.40651.09362.43282.77211.7798
O111.24362.48112.69812.94813.65473.93604.57812.47343.94332.77212.8180
H122.17673.25243.92473.98013.15283.15124.19081.09562.44911.77982.8180

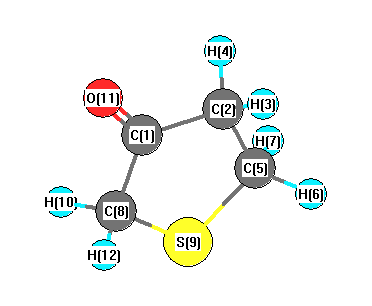
picture of Dihydro-3-(2H)-thiophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H3 109.506 C1 C2 H4 107.950
C1 C2 C5 107.236 C1 C8 S9 107.918
C1 C8 H10 109.153 C1 C8 H12 108.713
C2 C1 C8 111.225 C2 C1 O11 125.503
C2 C5 H6 110.966 C2 C5 H7 112.425
C2 C5 S9 105.455 H3 C2 H4 109.140
H3 C2 C5 113.254 H4 C2 C5 109.611
C5 S9 C8 91.574 H6 C5 H7 109.427
H6 C5 S9 110.096 H7 C5 S9 108.369
C8 C1 O11 123.273 S9 C8 H10 110.556
S9 C8 H12 111.675 H10 C8 H12 108.775
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability