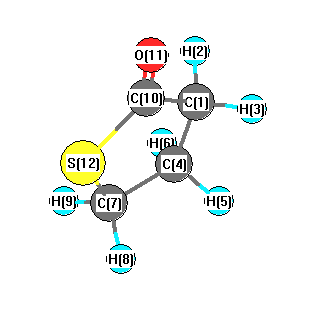
Vibrational Frequencies calculated at B3PW91/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3156 |
3028 |
8.35 |
|
|
|
| 2 |
A |
3149 |
3022 |
2.72 |
|
|
|
| 3 |
A |
3139 |
3011 |
7.24 |
|
|
|
| 4 |
A |
3088 |
2962 |
13.14 |
|
|
|
| 5 |
A |
3087 |
2962 |
15.57 |
|
|
|
| 6 |
A |
3060 |
2936 |
3.77 |
|
|
|
| 7 |
A |
1754 |
1683 |
246.94 |
|
|
|
| 8 |
A |
1537 |
1475 |
2.09 |
|
|
|
| 9 |
A |
1529 |
1467 |
12.39 |
|
|
|
| 10 |
A |
1501 |
1440 |
13.08 |
|
|
|
| 11 |
A |
1388 |
1332 |
0.18 |
|
|
|
| 12 |
A |
1349 |
1294 |
13.18 |
|
|
|
| 13 |
A |
1330 |
1276 |
1.15 |
|
|
|
| 14 |
A |
1285 |
1233 |
2.47 |
|
|
|
| 15 |
A |
1212 |
1163 |
2.76 |
|
|
|
| 16 |
A |
1163 |
1116 |
2.99 |
|
|
|
| 17 |
A |
1065 |
1022 |
9.41 |
|
|
|
| 18 |
A |
1049 |
1007 |
0.89 |
|
|
|
| 19 |
A |
1021 |
980 |
80.08 |
|
|
|
| 20 |
A |
917 |
879 |
13.58 |
|
|
|
| 21 |
A |
885 |
850 |
12.06 |
|
|
|
| 22 |
A |
843 |
808 |
31.74 |
|
|
|
| 23 |
A |
698 |
670 |
2.18 |
|
|
|
| 24 |
A |
610 |
585 |
10.92 |
|
|
|
| 25 |
A |
600 |
576 |
34.60 |
|
|
|
| 26 |
A |
491 |
471 |
2.28 |
|
|
|
| 27 |
A |
480 |
460 |
2.15 |
|
|
|
| 28 |
A |
390 |
374 |
2.90 |
|
|
|
| 29 |
A |
231 |
222 |
1.40 |
|
|
|
| 30 |
A |
129 |
123 |
0.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21068.3 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20212.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/3-21G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.516 |
|
|
|
| 2 |
H |
0.266 |
|
|
|
| 3 |
H |
0.262 |
|
|
|
| 4 |
C |
-0.455 |
|
|
|
| 5 |
H |
0.240 |
|
|
|
| 6 |
H |
0.247 |
|
|
|
| 7 |
C |
-0.618 |
|
|
|
| 8 |
H |
0.252 |
|
|
|
| 9 |
H |
0.258 |
|
|
|
| 10 |
C |
0.263 |
|
|
|
| 11 |
O |
-0.446 |
|
|
|
| 12 |
S |
0.246 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.593 |
1.891 |
0.408 |
4.081 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.629 |
-0.382 |
-0.084 |
| y |
-0.382 |
-42.537 |
0.189 |
| z |
-0.084 |
0.189 |
-42.278 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.222 |
-0.382 |
-0.084 |
| y |
-0.382 |
1.917 |
0.189 |
| z |
-0.084 |
0.189 |
2.305 |
|
| Polar |
|---|
| 3z2-r2 | 4.610 |
|---|
| x2-y2 | -4.092 |
|---|
| xy | -0.382 |
|---|
| xz | -0.084 |
|---|
| yz | 0.189 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
179.965 |
| (<r2>)1/2 |
13.415 |