return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Dihydro-2-(3H)-thiophenone)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-624.825654
Energy at 298.15K-624.833274
HF Energy-624.213742
Nuclear repulsion energy288.364343
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3183 3028 9.82      
2 A 3172 3017 4.63      
3 A 3163 3009 5.97      
4 A 3117 2965 13.95      
5 A 3111 2960 12.72      
6 A 3092 2941 4.13      
7 A 1650 1570 160.18      
8 A 1583 1505 0.77      
9 A 1576 1499 8.04      
10 A 1552 1476 8.33      
11 A 1419 1350 0.06      
12 A 1374 1307 15.25      
13 A 1363 1296 1.17      
14 A 1300 1236 3.14      
15 A 1229 1169 2.96      
16 A 1179 1122 2.56      
17 A 1086 1033 4.67      
18 A 1041 990 4.28      
19 A 1023 973 70.97      
20 A 908 864 0.69      
21 A 903 859 24.60      
22 A 846 805 40.29      
23 A 702 667 2.49      
24 A 607 578 2.46      
25 A 585 557 49.39      
26 A 494 469 3.53      
27 A 476 453 2.16      
28 A 390 371 4.42      
29 A 242 230 1.39      
30 A 118 112 0.20      

Unscaled Zero Point Vibrational Energy (zpe) 21239.4 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20205.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14481 0.10178 0.06326

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.115 1.435 0.216
H2 -0.581 2.276 -0.303
H3 -0.073 1.656 1.292
C4 1.286 1.078 -0.315
H5 2.051 1.776 0.038
H6 1.274 1.081 -1.410
C7 1.587 -0.365 0.186
H8 1.866 -0.355 1.243
H9 2.380 -0.839 -0.397
C10 -1.008 0.186 0.024
O11 -2.241 0.166 -0.069
S12 0.032 -1.308 -0.036

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 H9 C10 O11 S12
C11.09341.09851.53972.19972.16742.47772.86073.43101.54772.49312.7580
H21.09341.78462.21872.70092.46923.45273.91244.29952.15832.69563.6459
H31.09851.78462.18192.46963.07282.84032.79473.88552.15492.96203.2494
C41.53972.21872.18191.09401.09511.55722.19502.20902.48483.65182.7095
H52.19972.70092.46961.09401.78352.19642.45592.67163.44814.58603.6868
H62.16742.46923.07281.09511.78352.17603.07412.43612.84003.87253.0223
C72.47773.45272.84031.55722.19642.17601.09361.09222.65883.87391.8320
H82.86073.91242.79472.19502.45593.07411.09361.78573.16944.34362.4308
H93.43104.29953.88552.20902.67162.43611.09221.78573.56554.74122.4217
C101.54772.15832.15492.48483.44812.84002.65883.16943.56551.23661.8215
O112.49312.69562.96203.65184.58603.87253.87394.34364.74121.23662.7094
S122.75803.64593.24942.70953.68683.02231.83202.43082.42171.82152.7094

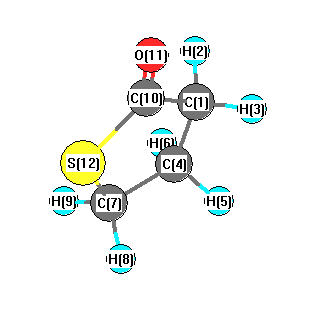
picture of Dihydro-2-(3H)-thiophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 112.166 C1 C4 H6 109.538
C1 C4 C7 106.267 C1 C10 O11 126.761
C1 C10 S12 109.625 H2 C1 H3 109.014
H2 C1 C4 113.754 H2 C1 C10 108.385
H3 C1 C4 110.472 H3 C1 C10 107.835
C4 C1 C10 107.186 C4 C7 H8 110.572
C4 C7 H9 111.776 C4 C7 S12 105.871
H5 C4 H6 109.123 H5 C4 C7 110.665
H6 C4 C7 109.010 C7 S12 C10 93.396
H8 C7 H9 109.563 H8 C7 S12 109.789
H9 C7 S12 109.195 O11 C10 S12 123.612
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability