return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2CH3 (Ethylbenzene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-307.764387
Energy at 298.15K-307.774805
HF Energy-307.055589
Nuclear repulsion energy337.823070
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3216 3059 14.01      
2 A' 3190 3034 2.92      
3 A' 3171 3016 7.97      
4 A' 3157 3003 34.52      
5 A' 3082 2932 21.88      
6 A' 3079 2929 21.24      
7 A' 1608 1530 3.74      
8 A' 1602 1524 2.23      
9 A' 1580 1503 2.44      
10 A' 1539 1464 12.00      
11 A' 1488 1416 2.81      
12 A' 1397 1329 1.06      
13 A' 1250 1189 0.01      
14 A' 1232 1172 0.53      
15 A' 1121 1066 12.52      
16 A' 1059 1008 1.59      
17 A' 1025 975 0.45      
18 A' 965 918 0.52      
19 A' 924 879 0.09      
20 A' 877 834 0.40      
21 A' 783 744 1.23      
22 A' 743 707 57.95      
23 A' 684 651 19.87      
24 A' 572 544 7.07      
25 A' 503 479 6.91      
26 A' 314 299 0.25      
27 A' 133 126 0.89      
28 A" 3200 3044 26.10      
29 A" 3171 3017 11.50      
30 A" 3165 3011 17.40      
31 A" 3122 2970 5.95      
32 A" 1600 1522 8.24      
33 A" 1590 1513 0.83      
34 A" 1503 1430 8.46      
35 A" 1411 1342 0.92      
36 A" 1350 1285 0.08      
37 A" 1308 1244 0.19      
38 A" 1240 1179 0.11      
39 A" 1155 1099 1.02      
40 A" 1074 1021 2.83      
41 A" 919 874 0.00      
42 A" 841 800 0.05      
43 A" 813 773 0.87      
44 A" 658 626 0.05      
45 A" 413 393 0.00      
46 A" 366 348 0.11      
47 A" 235 223 0.02      
48 A" 48 46 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 34736.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 33044.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.14760 0.04852 0.04027

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.252 0.505 0.000
C2 -0.247 -0.214 1.213
C3 -0.247 -1.620 1.216
C4 -0.246 -2.327 0.000
C5 -0.247 -1.620 -1.216
C6 -0.247 -0.214 -1.213
C7 -0.186 2.030 0.000
C8 1.291 2.518 0.000
H9 -0.254 0.330 2.156
H10 -0.250 -2.160 2.160
H11 -0.248 -3.414 0.000
H12 -0.250 -2.160 -2.160
H13 -0.254 0.330 -2.156
H14 -0.704 2.419 -0.886
H15 -0.704 2.419 0.886
H16 1.807 2.138 0.888
H17 1.807 2.138 -0.888
H18 1.344 3.612 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.40982.44832.83212.44831.40981.52662.53662.16353.43023.91903.43022.16352.15652.15652.77412.77413.4935
C21.40981.40592.43632.80622.42532.55163.36171.08932.16423.42203.89343.41273.39752.69193.13953.76344.3179
C32.44831.40591.40672.43202.80623.84784.57902.16471.08722.16733.41853.89554.57564.07774.29514.77145.6026
C42.83212.43631.40671.40672.43634.35765.08293.42192.16601.08692.16603.42194.84944.84944.99364.99366.1486
C52.44832.80622.43201.40671.40593.84784.57903.89553.41852.16731.08722.16474.07774.57564.77144.29515.6026
C61.40982.42532.80622.43631.40592.55163.36173.41273.89343.42202.16421.08932.69193.39753.76343.13954.3179
C71.52662.55163.84784.35763.84782.55161.55532.74704.71445.44454.71442.74701.09751.09752.18422.18422.2008
C82.53663.36174.57905.08294.57903.36171.55533.43875.37806.12825.37803.43872.18532.18531.09501.09501.0958
H92.16351.08932.16473.42193.89553.41272.74703.43872.48994.32054.98274.31293.71772.48603.02074.09674.2401
H103.43022.16421.08722.16603.41853.89344.71445.37802.48992.49724.31924.98275.51774.77424.93165.65606.3661
H113.91903.42202.16731.08692.16733.42205.44456.12824.32052.49722.49724.32055.91715.91715.98595.98597.2045
H123.43023.89343.41852.16601.08722.16424.71445.37804.98274.31922.49722.48994.77425.51775.65604.93166.3661
H132.16353.41273.89553.42192.16471.08932.74703.43874.31294.98274.32052.48992.48603.71774.09673.02074.2401
H142.15653.39754.57564.84944.07772.69191.09752.18533.71775.51775.91714.77422.48601.77163.08722.52692.5307
H152.15652.69194.07774.84944.57563.39751.09752.18532.48604.77425.91715.51773.71771.77162.52693.08722.5307
H162.77413.13954.29514.99364.77143.76342.18421.09503.02074.93165.98595.65604.09673.08722.52691.77561.7826
H172.77413.76344.77144.99364.29513.13952.18421.09504.09675.65605.98594.93163.02072.52693.08721.77561.7826
H183.49354.31795.60266.14865.60264.31792.20081.09584.24016.36617.20456.36614.24012.53072.53071.78261.7826

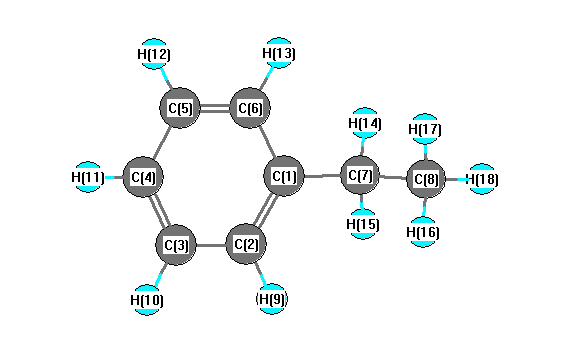
picture of Ethylbenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.803 C1 C2 H9 119.378
C1 C6 C5 120.803 C1 C6 H13 119.378
C1 C7 C8 110.785 C1 C7 H14 109.451
C1 C7 H15 109.451 C2 C1 C6 118.667
C2 C1 C7 120.623 C2 C3 C4 120.043
C2 C3 H10 119.925 C3 C2 H9 119.816
C3 C4 C5 119.640 C3 C4 H11 120.180
C4 C3 H10 120.033 C4 C5 C6 120.043
C4 C5 H12 120.033 C5 C4 H11 120.180
C5 C6 H13 119.816 C6 C1 C7 120.623
C6 C5 H12 119.925 C7 C8 H16 109.785
C7 C8 H17 109.785 C7 C8 H18 111.038
C8 C7 H14 109.730 C8 C7 H15 109.730
H14 C7 H15 107.635 H16 C8 H17 108.352
H16 C8 H18 108.913 H17 C8 H18 108.913
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability