return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-1143.848401
Energy at 298.15K-1143.852546
HF Energy-1143.057192
Nuclear repulsion energy445.267419
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3233 3076 0.00      
2 Ag 1583 1506 0.00      
3 Ag 1247 1186 0.00      
4 Ag 1097 1044 0.00      
5 Ag 764 727 0.00      
6 Ag 332 316 0.00      
7 Au 926 881 0.00      
8 Au 425 404 0.00      
9 B1g 824 784 0.00      
10 B1u 3216 3059 0.18      
11 B1u 1524 1450 77.61      
12 B1u 1123 1068 123.84      
13 B1u 1057 1006 17.80      
14 B1u 540 514 29.85      
15 B2g 896 853 0.00      
16 B2g 661 629 0.00      
17 B2g 299 285 0.00      
18 B2u 3230 3073 0.37      
19 B2u 1426 1357 9.36      
20 B2u 1303 1239 0.01      
21 B2u 1144 1088 2.96      
22 B2u 222 211 1.20      
23 B3g 3216 3059 0.00      
24 B3g 1569 1492 0.00      
25 B3g 1368 1301 0.00      
26 B3g 667 634 0.00      
27 B3g 361 343 0.00      
28 B3u 813 773 76.68      
29 B3u 488 464 21.02      
30 B3u 103 98 0.59      

Unscaled Zero Point Vibrational Energy (zpe) 17827.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 16959.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.18673 0.02184 0.01955

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.393
C2 0.000 0.000 -1.393
C3 0.000 1.221 0.702
C4 0.000 -1.221 0.702
C5 0.000 -1.221 -0.702
C6 0.000 1.221 -0.702
Cl7 0.000 0.000 3.153
Cl8 0.000 0.000 -3.153
H9 0.000 2.157 1.252
H10 0.000 -2.157 1.252
H11 0.000 -2.157 -1.252
H12 0.000 2.157 -1.252

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.78631.40261.40262.42512.42511.76014.54642.16152.16153.41303.4130
C22.78632.42512.42511.40261.40264.54641.76013.41303.41302.16152.1615
C31.40262.42512.44132.81661.40472.73804.04421.08563.42203.90222.1670
C41.40262.42512.44131.40472.81662.73804.04423.42201.08562.16703.9022
C52.42511.40262.81661.40472.44134.04422.73803.90222.16701.08563.4220
C62.42511.40261.40472.81662.44134.04422.73802.16703.90223.42201.0856
Cl71.76014.54642.73802.73804.04424.04426.30652.87532.87534.90494.9049
Cl84.54641.76014.04424.04422.73802.73806.30654.90494.90492.87532.8753
H92.16153.41301.08563.42203.90222.16702.87534.90494.31384.98782.5038
H102.16153.41303.42201.08562.16703.90222.87534.90494.31382.50384.9878
H113.41302.16153.90222.16701.08563.42204.90492.87534.98782.50384.3138
H123.41302.16152.16703.90223.42201.08564.90492.87532.50384.98784.3138

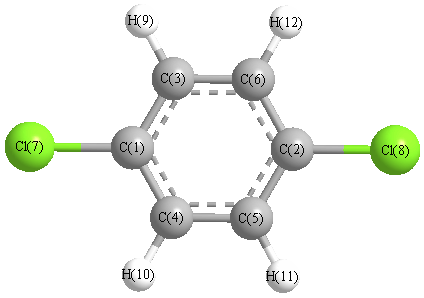
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.507 C1 C3 H9 120.080
C1 C4 C5 119.507 C1 C4 H10 120.080
C2 C5 C4 119.507 C2 C5 H11 120.080
C2 C6 C3 119.507 C2 C6 H12 120.080
C3 C1 C4 120.987 C3 C1 Cl7 119.507
C3 C6 H12 120.413 C4 C1 Cl7 119.507
C4 C5 H11 120.413 C5 C2 C6 120.987
C5 C2 Cl8 119.507 C5 C4 H10 120.413
C6 C2 Cl8 119.507 C6 C3 H9 120.413
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability