Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -5198.620910 |
| Energy at 298.15K | -5198.631163 |
| HF Energy | -5198.175117 |
| Nuclear repulsion energy | 415.233254 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3152 |
2999 |
0.00 |
|
|
|
| 2 |
Ag |
1573 |
1496 |
0.00 |
|
|
|
| 3 |
Ag |
1352 |
1286 |
0.00 |
|
|
|
| 4 |
Ag |
1034 |
983 |
0.00 |
|
|
|
| 5 |
Ag |
711 |
676 |
0.00 |
|
|
|
| 6 |
Ag |
198 |
188 |
0.00 |
|
|
|
| 7 |
Au |
3238 |
3081 |
0.53 |
|
|
|
| 8 |
Au |
1162 |
1105 |
3.14 |
|
|
|
| 9 |
Au |
795 |
756 |
5.73 |
|
|
|
| 10 |
Au |
111 |
105 |
4.59 |
|
|
|
| 11 |
Bg |
3216 |
3060 |
0.00 |
|
|
|
| 12 |
Bg |
1355 |
1289 |
0.00 |
|
|
|
| 13 |
Bg |
986 |
938 |
0.00 |
|
|
|
| 14 |
Bu |
3161 |
3007 |
4.57 |
|
|
|
| 15 |
Bu |
1575 |
1498 |
10.12 |
|
|
|
| 16 |
Bu |
1272 |
1210 |
47.58 |
|
|
|
| 17 |
Bu |
638 |
607 |
53.98 |
|
|
|
| 18 |
Bu |
186 |
177 |
7.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12856.1 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12230.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.504 |
0.575 |
0.000 |
| C2 |
-0.504 |
-0.575 |
0.000 |
| Br3 |
-0.504 |
2.270 |
0.000 |
| Br4 |
0.504 |
-2.270 |
0.000 |
| H5 |
1.123 |
0.567 |
0.897 |
| H6 |
1.123 |
0.567 |
-0.897 |
| H7 |
-1.123 |
-0.567 |
0.897 |
| H8 |
-1.123 |
-0.567 |
-0.897 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5288 | 1.9726 | 2.8450 | 1.0895 | 1.0895 | 2.1801 | 2.1801 |
C2 | 1.5288 | | 2.8450 | 1.9726 | 2.1801 | 2.1801 | 1.0895 | 1.0895 | Br3 | 1.9726 | 2.8450 | | 4.6512 | 2.5204 | 2.5204 | 3.0390 | 3.0390 | Br4 | 2.8450 | 1.9726 | 4.6512 | | 3.0390 | 3.0390 | 2.5204 | 2.5204 | H5 | 1.0895 | 2.1801 | 2.5204 | 3.0390 | | 1.7937 | 2.5150 | 3.0891 | H6 | 1.0895 | 2.1801 | 2.5204 | 3.0390 | 1.7937 | | 3.0891 | 2.5150 | H7 | 2.1801 | 1.0895 | 3.0390 | 2.5204 | 2.5150 | 3.0891 | | 1.7937 | H8 | 2.1801 | 1.0895 | 3.0390 | 2.5204 | 3.0891 | 2.5150 | 1.7937 | |
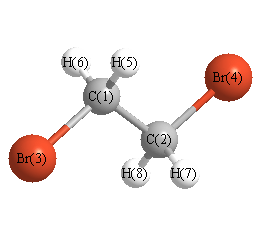 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.023 |
|
C1 |
C2 |
H7 |
111.641 |
| C1 |
C2 |
H8 |
111.641 |
|
C2 |
C1 |
Br3 |
108.023 |
| C2 |
C1 |
H5 |
111.641 |
|
C2 |
C1 |
H6 |
111.641 |
| Br3 |
C1 |
H5 |
107.246 |
|
Br3 |
C1 |
H6 |
107.246 |
| Br4 |
C2 |
H7 |
107.246 |
|
Br4 |
C2 |
H8 |
107.246 |
| H5 |
C1 |
H6 |
110.803 |
|
H7 |
C2 |
H8 |
110.803 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -5198.618076 |
| Energy at 298.15K | |
| HF Energy | -5198.171206 |
| Nuclear repulsion energy | 456.518704 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3198 |
3042 |
0.04 |
85.67 |
0.75 |
0.86 |
| 2 |
A |
3134 |
2981 |
11.10 |
156.02 |
0.02 |
0.05 |
| 3 |
A |
1557 |
1481 |
0.00 |
8.12 |
0.69 |
0.82 |
| 4 |
A |
1371 |
1304 |
20.38 |
4.56 |
0.71 |
0.83 |
| 5 |
A |
1234 |
1174 |
2.66 |
18.83 |
0.75 |
0.86 |
| 6 |
A |
1023 |
973 |
0.28 |
4.00 |
0.72 |
0.84 |
| 7 |
A |
953 |
907 |
8.60 |
6.37 |
0.55 |
0.71 |
| 8 |
A |
593 |
564 |
8.22 |
15.94 |
0.10 |
0.17 |
| 9 |
A |
231 |
220 |
0.99 |
0.82 |
0.38 |
0.55 |
| 10 |
A |
87 |
82 |
0.19 |
0.84 |
0.72 |
0.84 |
| 11 |
B |
3212 |
3055 |
1.11 |
25.49 |
0.75 |
0.86 |
| 12 |
B |
3127 |
2975 |
1.39 |
47.34 |
0.75 |
0.86 |
| 13 |
B |
1560 |
1484 |
12.59 |
20.97 |
0.75 |
0.86 |
| 14 |
B |
1359 |
1292 |
42.73 |
0.22 |
0.75 |
0.86 |
| 15 |
B |
1184 |
1126 |
2.37 |
7.00 |
0.75 |
0.86 |
| 16 |
B |
885 |
842 |
15.36 |
0.76 |
0.75 |
0.86 |
| 17 |
B |
629 |
599 |
7.58 |
9.13 |
0.75 |
0.86 |
| 18 |
B |
368 |
350 |
6.19 |
2.82 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12852.1 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12226.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.310 |
0.700 |
1.239 |
| C2 |
-0.310 |
-0.700 |
1.239 |
| Br3 |
-0.310 |
1.738 |
-0.307 |
| Br4 |
0.310 |
-1.738 |
-0.307 |
| H5 |
-0.000 |
1.251 |
2.130 |
| H6 |
1.397 |
0.652 |
1.176 |
| H7 |
0.000 |
-1.251 |
2.130 |
| H8 |
-1.397 |
-0.652 |
1.176 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5310 | 1.9623 | 2.8869 | 1.0930 | 1.0901 | 2.1670 | 2.1784 |
C2 | 1.5310 | | 2.8869 | 1.9623 | 2.1670 | 2.1784 | 1.0930 | 1.0901 | Br3 | 1.9623 | 2.8869 | | 3.5318 | 2.5045 | 2.5085 | 3.8692 | 3.0155 | Br4 | 2.8869 | 1.9623 | 3.5318 | | 3.8692 | 3.0155 | 2.5045 | 2.5085 | H5 | 1.0930 | 2.1670 | 2.5045 | 3.8692 | | 1.7952 | 2.5014 | 2.5460 | H6 | 1.0901 | 2.1784 | 2.5085 | 3.0155 | 1.7952 | | 2.5460 | 3.0835 | H7 | 2.1670 | 1.0930 | 3.8692 | 2.5045 | 2.5014 | 2.5460 | | 1.7952 | H8 | 2.1784 | 1.0901 | 3.0155 | 2.5085 | 2.5460 | 3.0835 | 1.7952 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
110.856 |
|
C1 |
C2 |
H7 |
110.232 |
| C1 |
C2 |
H8 |
111.311 |
|
C2 |
C1 |
Br3 |
110.856 |
| C2 |
C1 |
H5 |
110.232 |
|
C2 |
C1 |
H6 |
111.311 |
| Br3 |
C1 |
H5 |
106.625 |
|
Br3 |
C1 |
H6 |
107.036 |
| Br4 |
C2 |
H7 |
106.625 |
|
Br4 |
C2 |
H8 |
107.036 |
| H5 |
C1 |
H6 |
110.634 |
|
H7 |
C2 |
H8 |
110.634 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
