return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2Br (n-propyl bromide)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-2677.713009
Energy at 298.15K-2677.723531
HF Energy-2677.304577
Nuclear repulsion energy234.495012
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3160 3007 22.11      
2 A' 3133 2980 13.23      
3 A' 3108 2957 4.60      
4 A' 3083 2933 16.15      
5 A' 1605 1527 6.13      
6 A' 1592 1514 2.51      
7 A' 1574 1498 2.69      
8 A' 1494 1422 5.79      
9 A' 1418 1349 3.93      
10 A' 1315 1251 34.47      
11 A' 1146 1090 5.15      
12 A' 1022 972 2.09      
13 A' 907 863 6.99      
14 A' 692 659 17.44      
15 A' 323 307 1.52      
16 A' 219 208 1.51      
17 A" 3201 3045 11.95      
18 A" 3164 3010 23.58      
19 A" 3143 2990 1.45      
20 A" 1602 1524 8.60      
21 A" 1387 1320 0.02      
22 A" 1310 1246 0.19      
23 A" 1116 1062 2.35      
24 A" 905 861 0.01      
25 A" 776 739 5.43      
26 A" 247 235 0.04      
27 A" 121 115 0.75      

Unscaled Zero Point Vibrational Energy (zpe) 21382.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 20340.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.82474 0.05383 0.05203

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.854 0.000
C2 1.522 0.637 0.000
C3 2.267 1.999 0.000
Br4 -0.944 -0.881 0.000
H5 -0.332 1.383 0.895
H6 -0.332 1.383 -0.895
H7 1.799 0.055 0.885
H8 1.799 0.055 -0.885
H9 3.351 1.839 0.000
H10 2.004 2.582 -0.889
H11 2.004 2.582 0.889

Atom - Atom Distances (Å)
  C1 C2 C3 Br4 H5 H6 H7 H8 H9 H10 H11
C11.53782.53991.97461.09161.09162.15842.15843.49262.79142.7914
C21.53781.55172.89592.19012.19011.09491.09492.18792.19172.1917
C32.53991.55174.31292.81702.81702.18662.18661.09501.09561.0956
Br41.97462.89594.31292.51022.51023.03013.03015.08334.63334.6333
H51.09162.19012.81702.51021.79012.51163.07823.81713.17432.6253
H61.09162.19012.81702.51021.79013.07822.51163.81712.62533.1743
H72.15841.09492.18663.03012.51163.07821.76922.52483.09452.5355
H82.15841.09492.18663.03013.07822.51161.76922.52482.53553.0945
H93.49262.18791.09505.08333.81713.81712.52482.52481.77651.7765
H102.79142.19171.09564.63333.17432.62533.09452.53551.77651.7787
H112.79142.19171.09564.63332.62533.17432.53553.09451.77651.7787

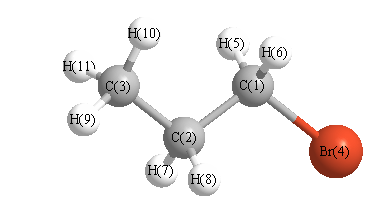
picture of n-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 110.592 C1 C2 H7 108.980
C1 C2 H8 108.980 C2 C1 Br4 110.460
C2 C1 H5 111.676 C2 C1 H6 111.676
C2 C3 H9 110.319 C2 C3 H10 110.579
C2 C3 H11 110.579 C3 C2 H7 110.220
C3 C2 H8 110.220 Br4 C1 H5 106.305
Br4 C1 H6 106.305 H5 C1 H6 110.146
H7 C2 H8 107.787 H9 C3 H10 108.382
H9 C3 H11 108.382 H10 C3 H11 108.529
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability