return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-155.605360
Energy at 298.15K-155.613387
HF Energy-155.241416
Nuclear repulsion energy116.393783
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3255 3097 18.79      
2 A 3174 3019 4.25      
3 A 3166 3012 39.00      
4 A 3161 3007 22.69      
5 A 3156 3002 15.90      
6 A 3117 2966 6.99      
7 A 3084 2934 20.93      
8 A 3076 2926 19.05      
9 A 1692 1610 3.58      
10 A 1605 1527 4.09      
11 A 1602 1524 6.03      
12 A 1579 1503 3.74      
13 A 1534 1460 1.10      
14 A 1488 1416 2.59      
15 A 1402 1333 1.21      
16 A 1380 1312 0.51      
17 A 1355 1289 0.02      
18 A 1245 1184 0.06      
19 A 1143 1088 4.22      
20 A 1057 1006 9.57      
21 A 1049 998 11.06      
22 A 1005 956 1.39      
23 A 960 914 43.44      
24 A 861 819 1.09      
25 A 827 787 3.75      
26 A 666 634 10.68      
27 A 444 422 0.56      
28 A 332 316 0.73      
29 A 242 230 0.02      
30 A 100 95 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 24379.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 23191.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.71304 0.13813 0.13526

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.715 -0.296 -0.288
C2 0.550 0.569 0.273
C3 -0.727 -0.245 0.402
C4 -1.848 -0.019 -0.301
H5 -0.686 -1.078 1.103
H6 1.452 -0.673 -1.282
H7 2.635 0.293 -0.366
H8 1.904 -1.153 0.368
H9 0.836 0.969 1.255
H10 0.369 1.422 -0.394
H11 -1.918 0.802 -1.010
H12 -2.732 -0.640 -0.191

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.55622.53813.57382.88371.09451.09531.09552.18102.18503.86394.4612
C21.55621.51982.53522.22122.18492.19842.19251.09791.09762.79213.5285
C32.53811.51981.34241.09042.78643.49032.78322.15462.14742.12372.1280
C43.57382.53521.34242.10793.50394.49463.97603.25582.64591.08761.0858
H52.88372.22121.09042.10793.22873.88242.69362.55603.09963.08582.4599
H61.09452.18492.78643.50393.22871.78091.77663.08402.51933.68864.3235
H71.09532.19843.49034.49463.88241.78091.77872.51482.53194.62715.4504
H81.09552.19252.78323.97602.69361.77661.77872.53603.09264.50894.6973
H92.18101.09792.15463.25582.55603.08402.51482.53601.77253.57014.1724
H102.18501.09762.14742.64593.09962.51932.53193.09261.77252.44893.7296
H113.86392.79212.12371.08763.08583.68864.62714.50893.57012.44891.8475
H124.46123.52852.12801.08582.45994.32355.45044.69734.17243.72961.8475

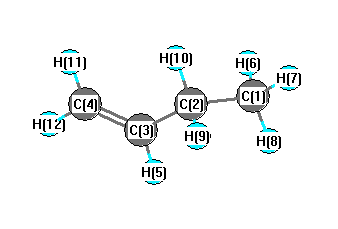
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.200 C1 C2 H9 109.311
C1 C2 H10 109.635 C2 C1 H6 109.804
C2 C1 H7 110.817 C2 C1 H8 110.347
C2 C3 C4 124.580 C2 C3 H5 115.655
C3 C2 H9 109.747 C3 C2 H10 109.199
C3 C4 H11 121.495 C3 C4 H12 122.064
C4 C3 H5 119.743 H6 C1 H7 108.824
H6 C1 H8 108.436 H7 C1 H8 108.557
H9 C2 H10 107.679 H11 C4 H12 116.441
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability