Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -190.079047 |
| Energy at 298.15K | |
| HF Energy | -189.685168 |
| Nuclear repulsion energy | 101.596175 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3266 |
3107 |
7.66 |
56.08 |
0.45 |
0.62 |
| 2 |
A' |
3216 |
3060 |
1.25 |
76.72 |
0.36 |
0.53 |
| 3 |
A' |
3168 |
3013 |
4.44 |
58.51 |
0.11 |
0.19 |
| 4 |
A' |
2926 |
2784 |
103.72 |
78.87 |
0.25 |
0.40 |
| 5 |
A' |
1678 |
1596 |
7.37 |
9.22 |
0.37 |
0.54 |
| 6 |
A' |
1615 |
1536 |
51.18 |
12.98 |
0.56 |
0.72 |
| 7 |
A' |
1509 |
1435 |
3.49 |
5.18 |
0.42 |
0.60 |
| 8 |
A' |
1447 |
1377 |
0.57 |
12.67 |
0.54 |
0.70 |
| 9 |
A' |
1340 |
1275 |
1.39 |
8.30 |
0.38 |
0.55 |
| 10 |
A' |
1174 |
1117 |
18.77 |
8.82 |
0.67 |
0.80 |
| 11 |
A' |
932 |
887 |
33.25 |
0.81 |
0.22 |
0.36 |
| 12 |
A' |
562 |
535 |
2.81 |
4.94 |
0.34 |
0.51 |
| 13 |
A' |
323 |
307 |
7.24 |
0.14 |
0.33 |
0.50 |
| 14 |
A" |
1053 |
1002 |
15.25 |
2.86 |
0.75 |
0.86 |
| 15 |
A" |
1036 |
986 |
13.82 |
4.13 |
0.75 |
0.86 |
| 16 |
A" |
1001 |
952 |
41.38 |
0.12 |
0.75 |
0.86 |
| 17 |
A" |
618 |
588 |
13.01 |
6.82 |
0.75 |
0.86 |
| 18 |
A" |
174 |
166 |
2.87 |
1.23 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13518.6 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12860.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.141 |
-0.747 |
0.000 |
| C2 |
0.000 |
0.735 |
0.000 |
| C3 |
1.217 |
1.306 |
0.000 |
| O4 |
-1.232 |
-1.361 |
0.000 |
| H5 |
0.832 |
-1.279 |
0.000 |
| H6 |
-0.915 |
1.321 |
0.000 |
| H7 |
1.356 |
2.382 |
0.000 |
| H8 |
2.122 |
0.701 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4887 | 2.4619 | 1.2518 | 1.1082 | 2.2081 | 3.4689 | 2.6869 |
C2 | 1.4887 | | 1.3447 | 2.4306 | 2.1784 | 1.0862 | 2.1338 | 2.1227 | C3 | 2.4619 | 1.3447 | | 3.6208 | 2.6132 | 2.1319 | 1.0847 | 1.0889 | O4 | 1.2518 | 2.4306 | 3.6208 | | 2.0657 | 2.6999 | 4.5503 | 3.9373 | H5 | 1.1082 | 2.1784 | 2.6132 | 2.0657 | | 3.1315 | 3.6978 | 2.3629 | H6 | 2.2081 | 1.0862 | 2.1319 | 2.6999 | 3.1315 | | 2.5067 | 3.0996 | H7 | 3.4689 | 2.1338 | 1.0847 | 4.5503 | 3.6978 | 2.5067 | | 1.8473 | H8 | 2.6869 | 2.1227 | 1.0889 | 3.9373 | 2.3629 | 3.0996 | 1.8473 | |
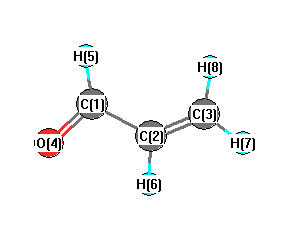 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.570 |
|
C1 |
C2 |
H6 |
117.223 |
| C2 |
C1 |
O4 |
124.750 |
|
C2 |
C1 |
H5 |
113.216 |
| C2 |
C3 |
H7 |
122.515 |
|
C2 |
C3 |
H8 |
121.084 |
| C3 |
C2 |
H6 |
122.207 |
|
O4 |
C1 |
H5 |
122.034 |
| H7 |
C3 |
H8 |
116.401 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -190.079041 |
| Energy at 298.15K | |
| HF Energy | -189.685340 |
| Nuclear repulsion energy | 103.486232 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3277 |
3117 |
4.24 |
48.96 |
0.52 |
0.68 |
| 2 |
A' |
3213 |
3057 |
6.48 |
88.41 |
0.29 |
0.45 |
| 3 |
A' |
3179 |
3024 |
3.07 |
49.15 |
0.11 |
0.20 |
| 4 |
A' |
2993 |
2847 |
134.05 |
127.17 |
0.31 |
0.47 |
| 5 |
A' |
1688 |
1605 |
2.02 |
5.22 |
0.20 |
0.34 |
| 6 |
A' |
1596 |
1519 |
33.96 |
14.99 |
0.40 |
0.57 |
| 7 |
A' |
1508 |
1434 |
24.29 |
8.98 |
0.24 |
0.39 |
| 8 |
A' |
1458 |
1387 |
7.55 |
10.98 |
0.63 |
0.78 |
| 9 |
A' |
1364 |
1297 |
1.46 |
10.41 |
0.47 |
0.64 |
| 10 |
A' |
1116 |
1062 |
7.24 |
5.12 |
0.74 |
0.85 |
| 11 |
A' |
915 |
871 |
56.46 |
3.16 |
0.17 |
0.29 |
| 12 |
A' |
702 |
668 |
3.09 |
1.64 |
0.49 |
0.66 |
| 13 |
A' |
297 |
282 |
8.96 |
1.89 |
0.52 |
0.69 |
| 14 |
A" |
1052 |
1001 |
1.31 |
9.44 |
0.75 |
0.86 |
| 15 |
A" |
1046 |
995 |
43.84 |
0.65 |
0.75 |
0.86 |
| 16 |
A" |
1012 |
963 |
16.72 |
0.04 |
0.75 |
0.86 |
| 17 |
A" |
591 |
562 |
14.53 |
8.64 |
0.75 |
0.86 |
| 18 |
A" |
222 |
211 |
5.88 |
1.45 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13613.5 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12950.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.902 |
-0.271 |
0.000 |
| C2 |
0.000 |
0.923 |
0.000 |
| C3 |
1.333 |
0.755 |
0.000 |
| O4 |
-0.490 |
-1.455 |
0.000 |
| H5 |
-1.979 |
-0.035 |
0.000 |
| H6 |
-0.464 |
1.906 |
0.000 |
| H7 |
2.034 |
1.582 |
0.000 |
| H8 |
1.742 |
-0.253 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4966 | 2.4592 | 1.2544 | 1.1022 | 2.2199 | 3.4717 | 2.6443 |
C2 | 1.4966 | | 1.3438 | 2.4287 | 2.1990 | 1.0864 | 2.1378 | 2.1023 | C3 | 2.4592 | 1.3438 | | 2.8648 | 3.4050 | 2.1345 | 1.0842 | 1.0876 | O4 | 1.2544 | 2.4287 | 2.8648 | | 2.0575 | 3.3611 | 3.9489 | 2.5351 | H5 | 1.1022 | 2.1990 | 3.4050 | 2.0575 | | 2.4622 | 4.3265 | 3.7274 | H6 | 2.2199 | 1.0864 | 2.1345 | 3.3611 | 2.4622 | | 2.5191 | 3.0869 | H7 | 3.4717 | 2.1378 | 1.0842 | 3.9489 | 4.3265 | 2.5191 | | 1.8582 | H8 | 2.6443 | 2.1023 | 1.0876 | 2.5351 | 3.7274 | 3.0869 | 1.8582 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
119.850 |
|
C1 |
C2 |
H6 |
117.629 |
| C2 |
C1 |
O4 |
123.737 |
|
C2 |
C1 |
H5 |
114.753 |
| C2 |
C3 |
H7 |
123.042 |
|
C2 |
C3 |
H8 |
119.307 |
| C3 |
C2 |
H6 |
122.521 |
|
O4 |
C1 |
H5 |
121.510 |
| H7 |
C3 |
H8 |
117.651 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
