Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -3095.753277 |
| Energy at 298.15K | |
| HF Energy | -3095.306001 |
| Nuclear repulsion energy | 286.627973 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3163 |
3009 |
6.11 |
3.40 |
0.12 |
0.21 |
| 2 |
A' |
3154 |
3000 |
0.11 |
119.56 |
0.10 |
0.18 |
| 3 |
A' |
1579 |
1503 |
9.56 |
2.78 |
0.75 |
0.86 |
| 4 |
A' |
1574 |
1498 |
1.09 |
21.01 |
0.75 |
0.86 |
| 5 |
A' |
1383 |
1316 |
1.46 |
10.10 |
0.52 |
0.68 |
| 6 |
A' |
1296 |
1232 |
34.92 |
1.26 |
0.45 |
0.62 |
| 7 |
A' |
1033 |
983 |
1.01 |
7.96 |
0.75 |
0.86 |
| 8 |
A' |
742 |
706 |
11.59 |
49.69 |
0.30 |
0.47 |
| 9 |
A' |
673 |
641 |
59.14 |
1.23 |
0.30 |
0.47 |
| 10 |
A' |
257 |
245 |
1.15 |
5.19 |
0.34 |
0.51 |
| 11 |
A' |
200 |
190 |
7.27 |
0.19 |
0.35 |
0.52 |
| 12 |
A" |
3238 |
3080 |
1.07 |
0.09 |
0.75 |
0.86 |
| 13 |
A" |
3217 |
3060 |
0.03 |
85.11 |
0.75 |
0.86 |
| 14 |
A" |
1363 |
1297 |
0.05 |
12.94 |
0.75 |
0.86 |
| 15 |
A" |
1191 |
1133 |
2.06 |
0.16 |
0.75 |
0.86 |
| 16 |
A" |
1018 |
968 |
0.11 |
10.30 |
0.75 |
0.86 |
| 17 |
A" |
804 |
765 |
4.97 |
0.03 |
0.75 |
0.86 |
| 18 |
A" |
117 |
111 |
6.33 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13000.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12367.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.667 |
0.000 |
| C2 |
1.226 |
-0.251 |
0.000 |
| Br3 |
-1.606 |
-0.472 |
0.000 |
| Cl4 |
2.731 |
0.776 |
0.000 |
| H5 |
-0.039 |
1.284 |
0.897 |
| H6 |
-0.039 |
1.284 |
-0.897 |
| H7 |
1.251 |
-0.872 |
0.894 |
| H8 |
1.251 |
-0.872 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5313 | 1.9685 | 2.7327 | 1.0896 | 1.0896 | 2.1755 | 2.1755 |
C2 | 1.5313 | | 2.8403 | 1.8212 | 2.1819 | 2.1819 | 1.0891 | 1.0891 | Br3 | 1.9685 | 2.8403 | | 4.5121 | 2.5185 | 2.5185 | 3.0199 | 3.0199 | Cl4 | 2.7327 | 1.8212 | 4.5121 | | 2.9554 | 2.9554 | 2.3881 | 2.3881 | H5 | 1.0896 | 2.1819 | 2.5185 | 2.9554 | | 1.7940 | 2.5127 | 3.0857 | H6 | 1.0896 | 2.1819 | 2.5185 | 2.9554 | 1.7940 | | 3.0857 | 2.5127 | H7 | 2.1755 | 1.0891 | 3.0199 | 2.3881 | 2.5127 | 3.0857 | | 1.7883 | H8 | 2.1755 | 1.0891 | 3.0199 | 2.3881 | 3.0857 | 2.5127 | 1.7883 | |
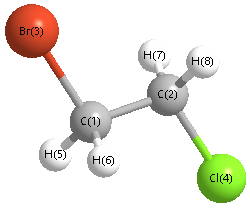 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.891 |
|
C1 |
C2 |
H7 |
111.115 |
| C1 |
C2 |
H8 |
111.115 |
|
C2 |
C1 |
Br3 |
107.839 |
| C2 |
C1 |
Cl4 |
39.091 |
|
C2 |
C1 |
H6 |
111.604 |
| Br3 |
C1 |
H5 |
107.368 |
|
Br3 |
C1 |
H6 |
107.368 |
| Cl4 |
C2 |
H7 |
107.605 |
|
Cl4 |
C2 |
H8 |
107.605 |
| H5 |
C1 |
H6 |
110.824 |
|
H7 |
C2 |
H8 |
110.369 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -3095.750422 |
| Energy at 298.15K | |
| HF Energy | -3095.302526 |
| Nuclear repulsion energy | 304.771267 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3213 |
3057 |
1.41 |
23.29 |
0.75 |
0.86 |
| 2 |
A |
3200 |
3044 |
0.24 |
86.40 |
0.75 |
0.86 |
| 3 |
A |
3138 |
2985 |
10.82 |
144.48 |
0.03 |
0.05 |
| 4 |
A |
3131 |
2979 |
2.51 |
49.29 |
0.61 |
0.75 |
| 5 |
A |
1565 |
1489 |
11.79 |
19.16 |
0.74 |
0.85 |
| 6 |
A |
1558 |
1482 |
2.22 |
9.97 |
0.72 |
0.84 |
| 7 |
A |
1400 |
1332 |
20.38 |
3.71 |
0.75 |
0.86 |
| 8 |
A |
1371 |
1305 |
32.04 |
2.69 |
0.72 |
0.84 |
| 9 |
A |
1260 |
1198 |
3.37 |
19.05 |
0.74 |
0.85 |
| 10 |
A |
1210 |
1151 |
1.40 |
7.35 |
0.74 |
0.85 |
| 11 |
A |
1037 |
986 |
0.61 |
4.61 |
0.66 |
0.80 |
| 12 |
A |
969 |
921 |
10.25 |
5.37 |
0.55 |
0.71 |
| 13 |
A |
908 |
864 |
17.48 |
1.26 |
0.72 |
0.84 |
| 14 |
A |
673 |
640 |
17.33 |
10.90 |
0.48 |
0.65 |
| 15 |
A |
608 |
578 |
9.46 |
14.28 |
0.17 |
0.29 |
| 16 |
A |
401 |
382 |
7.22 |
2.83 |
0.74 |
0.85 |
| 17 |
A |
248 |
236 |
1.22 |
0.88 |
0.36 |
0.53 |
| 18 |
A |
103 |
98 |
0.60 |
1.27 |
0.73 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 12995.7 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12362.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.000 |
1.142 |
-0.391 |
| C2 |
1.287 |
0.926 |
0.411 |
| Br3 |
-1.336 |
-0.231 |
0.037 |
| Cl4 |
2.132 |
-0.595 |
-0.090 |
| H5 |
-0.445 |
2.104 |
-0.128 |
| H6 |
0.186 |
1.086 |
-1.463 |
| H7 |
1.979 |
1.751 |
0.225 |
| H8 |
1.072 |
0.846 |
1.476 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5310 | 1.9624 | 2.7664 | 1.0925 | 1.0902 | 2.1598 | 2.1725 |
C2 | 1.5310 | | 2.8903 | 1.8105 | 2.1629 | 2.1793 | 1.0926 | 1.0897 | Br3 | 1.9624 | 2.8903 | | 3.4892 | 2.5045 | 2.5096 | 3.8661 | 3.0044 | Cl4 | 2.7664 | 1.8105 | 3.4892 | | 3.7323 | 2.9154 | 2.3717 | 2.3776 | H5 | 1.0925 | 2.1629 | 2.5045 | 3.7323 | | 1.7943 | 2.4748 | 2.5408 | H6 | 1.0902 | 2.1793 | 2.5096 | 2.9154 | 1.7943 | | 2.5510 | 3.0792 | H7 | 2.1598 | 1.0926 | 3.8661 | 2.3717 | 2.4748 | 2.5510 | | 1.7903 | H8 | 2.1725 | 1.0897 | 3.0044 | 2.3776 | 2.5408 | 3.0792 | 1.7903 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
111.491 |
|
C1 |
C2 |
H7 |
109.682 |
| C1 |
C2 |
H8 |
110.863 |
|
C2 |
C1 |
Br3 |
111.056 |
| C2 |
C1 |
Cl4 |
37.514 |
|
C2 |
C1 |
H6 |
111.378 |
| Br3 |
C1 |
H5 |
106.641 |
|
Br3 |
C1 |
H6 |
107.108 |
| Cl4 |
C2 |
H7 |
106.950 |
|
Cl4 |
C2 |
H8 |
107.517 |
| H5 |
C1 |
H6 |
110.587 |
|
H7 |
C2 |
H8 |
110.245 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
