Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -992.885776 |
| Energy at 298.15K | -992.890497 |
| HF Energy | -992.436961 |
| Nuclear repulsion energy | 192.554653 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3155 |
3001 |
0.00 |
|
|
|
| 2 |
Ag |
1578 |
1501 |
0.00 |
|
|
|
| 3 |
Ag |
1404 |
1335 |
0.00 |
|
|
|
| 4 |
Ag |
1034 |
984 |
0.00 |
|
|
|
| 5 |
Ag |
767 |
730 |
0.00 |
|
|
|
| 6 |
Ag |
306 |
291 |
0.00 |
|
|
|
| 7 |
Au |
3237 |
3080 |
1.78 |
|
|
|
| 8 |
Au |
1214 |
1155 |
1.40 |
|
|
|
| 9 |
Au |
816 |
776 |
4.16 |
|
|
|
| 10 |
Au |
122 |
116 |
8.13 |
|
|
|
| 11 |
Bg |
3216 |
3060 |
0.00 |
|
|
|
| 12 |
Bg |
1372 |
1305 |
0.00 |
|
|
|
| 13 |
Bg |
1056 |
1004 |
0.00 |
|
|
|
| 14 |
Bu |
3164 |
3010 |
8.08 |
|
|
|
| 15 |
Bu |
1583 |
1506 |
11.37 |
|
|
|
| 16 |
Bu |
1330 |
1265 |
27.08 |
|
|
|
| 17 |
Bu |
715 |
680 |
81.44 |
|
|
|
| 18 |
Bu |
223 |
213 |
9.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13145.9 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12505.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.766 |
0.000 |
| C2 |
0.000 |
-0.766 |
0.000 |
| Cl3 |
-1.721 |
1.349 |
0.000 |
| Cl4 |
1.721 |
-1.349 |
0.000 |
| H5 |
0.483 |
1.159 |
0.894 |
| H6 |
0.483 |
1.159 |
-0.894 |
| H7 |
-0.483 |
-1.159 |
0.894 |
| H8 |
-0.483 |
-1.159 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5330 | 1.8173 | 2.7275 | 1.0892 | 1.0892 | 2.1768 | 2.1768 |
C2 | 1.5330 | | 2.7275 | 1.8173 | 2.1768 | 2.1768 | 1.0892 | 1.0892 | Cl3 | 1.8173 | 2.7275 | | 4.3743 | 2.3861 | 2.3861 | 2.9365 | 2.9365 | Cl4 | 2.7275 | 1.8173 | 4.3743 | | 2.9365 | 2.9365 | 2.3861 | 2.3861 | H5 | 1.0892 | 2.1768 | 2.3861 | 2.9365 | | 1.7886 | 2.5101 | 3.0821 | H6 | 1.0892 | 2.1768 | 2.3861 | 2.9365 | 1.7886 | | 3.0821 | 2.5101 | H7 | 2.1768 | 1.0892 | 2.9365 | 2.3861 | 2.5101 | 3.0821 | | 1.7886 | H8 | 2.1768 | 1.0892 | 2.9365 | 2.3861 | 3.0821 | 2.5101 | 1.7886 | |
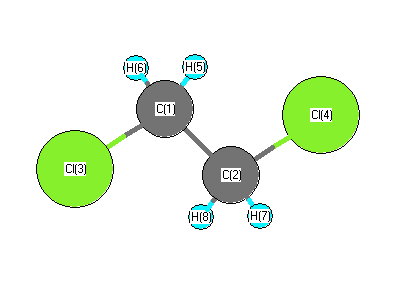 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.705 |
|
C1 |
C2 |
H7 |
111.097 |
| C1 |
C2 |
H8 |
111.097 |
|
C2 |
C1 |
Cl3 |
108.705 |
| C2 |
C1 |
H5 |
111.097 |
|
C2 |
C1 |
H6 |
111.097 |
| Cl3 |
C1 |
H5 |
107.715 |
|
Cl3 |
C1 |
H6 |
107.715 |
| Cl4 |
C2 |
H7 |
107.715 |
|
Cl4 |
C2 |
H8 |
107.715 |
| H5 |
C1 |
H6 |
110.377 |
|
H7 |
C2 |
H8 |
110.377 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -992.883281 |
| Energy at 298.15K | -992.888147 |
| HF Energy | -992.434006 |
| Nuclear repulsion energy | 200.823866 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3201 |
3045 |
0.22 |
|
|
|
| 2 |
A |
3140 |
2987 |
12.09 |
|
|
|
| 3 |
A |
1562 |
1486 |
0.16 |
|
|
|
| 4 |
A |
1407 |
1338 |
15.99 |
|
|
|
| 5 |
A |
1287 |
1224 |
1.20 |
|
|
|
| 6 |
A |
1055 |
1004 |
0.01 |
|
|
|
| 7 |
A |
980 |
933 |
13.80 |
|
|
|
| 8 |
A |
657 |
625 |
18.99 |
|
|
|
| 9 |
A |
270 |
257 |
0.94 |
|
|
|
| 10 |
A |
118 |
112 |
1.05 |
|
|
|
| 11 |
B |
3213 |
3056 |
2.01 |
|
|
|
| 12 |
B |
3134 |
2982 |
2.16 |
|
|
|
| 13 |
B |
1567 |
1491 |
14.96 |
|
|
|
| 14 |
B |
1402 |
1333 |
26.50 |
|
|
|
| 15 |
B |
1235 |
1175 |
1.04 |
|
|
|
| 16 |
B |
937 |
891 |
16.65 |
|
|
|
| 17 |
B |
685 |
652 |
21.25 |
|
|
|
| 18 |
B |
428 |
407 |
8.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13138.6 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 12498.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.301 |
0.704 |
0.915 |
| C2 |
-0.301 |
-0.704 |
0.915 |
| Cl3 |
-0.301 |
1.686 |
-0.480 |
| Cl4 |
0.301 |
-1.686 |
-0.480 |
| H5 |
0.003 |
1.226 |
1.827 |
| H6 |
1.388 |
0.662 |
0.845 |
| H7 |
-0.003 |
-1.226 |
1.827 |
| H8 |
-1.388 |
-0.662 |
0.845 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5308 | 1.8097 | 2.7674 | 1.0922 | 1.0899 | 2.1557 | 2.1729 |
C2 | 1.5308 | | 2.7674 | 1.8097 | 2.1557 | 2.1729 | 1.0922 | 1.0899 | Cl3 | 1.8097 | 2.7674 | | 3.4252 | 2.3727 | 2.3783 | 3.7272 | 2.9066 | Cl4 | 2.7674 | 1.8097 | 3.4252 | | 3.7272 | 2.9066 | 2.3727 | 2.3783 | H5 | 1.0922 | 2.1557 | 2.3727 | 3.7272 | | 1.7892 | 2.4515 | 2.5421 | H6 | 1.0899 | 2.1729 | 2.3783 | 2.9066 | 1.7892 | | 2.5421 | 3.0749 | H7 | 2.1557 | 1.0922 | 3.7272 | 2.3727 | 2.4515 | 2.5421 | | 1.7892 | H8 | 2.1729 | 1.0899 | 2.9066 | 2.3783 | 2.5421 | 3.0749 | 1.7892 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
111.605 |
|
C1 |
C2 |
H7 |
109.398 |
| C1 |
C2 |
H8 |
110.899 |
|
C2 |
C1 |
Cl3 |
111.605 |
| C2 |
C1 |
H5 |
109.398 |
|
C2 |
C1 |
H6 |
110.899 |
| Cl3 |
C1 |
H5 |
107.084 |
|
Cl3 |
C1 |
H6 |
107.612 |
| Cl4 |
C2 |
H7 |
107.084 |
|
Cl4 |
C2 |
H8 |
107.612 |
| H5 |
C1 |
H6 |
110.159 |
|
H7 |
C2 |
H8 |
110.159 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
