return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: MP2=FULL/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/3-21G*
 hartrees
Energy at 0K-172.713376
Energy at 298.15K-172.723863
HF Energy-172.322058
Nuclear repulsion energy129.669272
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3421 3243 1.96      
2 A' 3157 2993 28.21      
3 A' 3088 2927 48.89      
4 A' 3081 2921 13.28      
5 A' 3072 2913 9.74      
6 A' 1759 1668 17.70      
7 A' 1607 1523 4.63      
8 A' 1592 1509 1.11      
9 A' 1579 1497 0.43      
10 A' 1494 1416 5.03      
11 A' 1433 1359 0.89      
12 A' 1383 1311 2.87      
13 A' 1169 1108 3.31      
14 A' 1079 1023 4.18      
15 A' 1016 963 6.42      
16 A' 910 862 68.06      
17 A' 796 755 194.76      
18 A' 449 426 2.26      
19 A' 267 253 4.50      
20 A" 3522 3339 1.40      
21 A" 3156 2992 61.15      
22 A" 3138 2975 15.85      
23 A" 3109 2947 1.79      
24 A" 1604 1521 7.31      
25 A" 1430 1356 0.97      
26 A" 1387 1315 0.45      
27 A" 1310 1242 0.03      
28 A" 1081 1025 0.00      
29 A" 902 855 2.30      
30 A" 768 728 2.69      
31 A" 316 300 58.66      
32 A" 238 226 5.66      
33 A" 141 134 0.25      

Unscaled Zero Point Vibrational Energy (zpe) 27225.9 cm-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 25810.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/3-21G*
ABC
0.81000 0.12001 0.11314

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.453 1.300 0.000
C2 0.000 0.764 0.000
C3 -0.046 -0.787 0.000
N4 -1.416 -1.385 0.000
H5 1.471 2.394 0.000
H6 1.989 0.945 0.888
H7 1.989 0.945 -0.888
H8 -0.534 1.133 0.886
H9 -0.534 1.133 -0.886
H10 0.481 -1.164 -0.884
H11 0.481 -1.164 0.884
H12 -1.936 -1.094 -0.833
H13 -1.936 -1.094 0.833

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.54892.56993.92981.09491.09571.09572.18192.18192.79182.79184.23194.2319
C21.54891.55222.57402.19602.18542.18541.09791.09792.17472.17472.80952.8095
C32.56991.55221.49483.52512.81612.81612.17032.17031.09601.09602.08762.0876
N43.92982.57401.49484.75644.22054.22052.81172.81172.10512.10511.02401.0240
H51.09492.19603.52514.75641.77661.77662.52892.52893.79753.79754.94674.9467
H61.09572.18542.81614.22051.77661.77592.52943.08923.13982.59204.74554.4228
H71.09572.18542.81614.22051.77661.77593.08922.52942.59203.13984.42284.7455
H82.18191.09792.17032.81172.52892.52943.08921.77123.07202.51123.14312.6322
H92.18191.09792.17032.81172.52893.08922.52941.77122.51123.07202.63223.1431
H102.79182.17471.09602.10513.79753.13982.59203.07202.51121.76782.41872.9656
H112.79182.17471.09602.10513.79752.59203.13982.51123.07201.76782.96562.4187
H124.23192.80952.08761.02404.94674.74554.42283.14312.63222.41872.96561.6659
H134.23192.80952.08761.02404.94674.42284.74552.63223.14312.96562.41871.6659

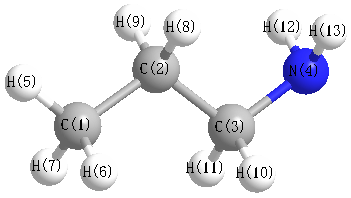
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.932 C1 C2 H8 109.875
C1 C2 H9 109.875 C2 C1 H5 111.165
C2 C1 H6 110.273 C2 C1 H7 110.273
C2 C3 N4 115.284 C2 C3 H10 109.199
C2 C3 H11 109.199 C3 C2 H8 108.754
C3 C2 H9 108.754 C3 N4 H12 110.557
C3 N4 H13 110.557 N4 C3 H10 107.691
N4 C3 H11 107.691 H5 C1 H6 108.391
H5 C1 H7 108.391 H6 C1 H7 108.266
H8 C2 H9 107.537 H10 C3 H11 107.502
H12 N4 H13 108.869
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability