Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Geometric Data calculated at MP2=FULL/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -206.031146 |
| Energy at 298.15K | -206.034594 |
| HF Energy | -205.604338 |
| Nuclear repulsion energy | 99.984787 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3511 |
3328 |
6.92 |
75.19 |
0.30 |
0.46 |
| 2 |
A |
3170 |
3006 |
4.74 |
72.53 |
0.43 |
0.61 |
| 3 |
A |
3079 |
2919 |
23.57 |
87.64 |
0.18 |
0.31 |
| 4 |
A |
2140 |
2029 |
15.17 |
15.94 |
0.27 |
0.42 |
| 5 |
A |
1592 |
1509 |
2.64 |
19.60 |
0.69 |
0.82 |
| 6 |
A |
1438 |
1363 |
17.61 |
11.39 |
0.74 |
0.85 |
| 7 |
A |
1419 |
1345 |
4.90 |
11.48 |
0.75 |
0.85 |
| 8 |
A |
1282 |
1216 |
21.98 |
5.26 |
0.70 |
0.82 |
| 9 |
A |
1027 |
974 |
0.31 |
5.06 |
0.40 |
0.57 |
| 10 |
A |
983 |
932 |
84.86 |
2.99 |
0.33 |
0.50 |
| 11 |
A |
878 |
832 |
24.82 |
2.80 |
0.11 |
0.19 |
| 12 |
A |
585 |
554 |
2.35 |
1.95 |
0.26 |
0.41 |
| 13 |
A |
441 |
418 |
67.81 |
1.63 |
0.73 |
0.84 |
| 14 |
A |
343 |
325 |
109.68 |
8.34 |
0.75 |
0.86 |
| 15 |
A |
224 |
213 |
6.11 |
3.81 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 11055.8 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 10480.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.564 |
0.616 |
0.037 |
| C2 |
0.836 |
0.129 |
0.003 |
| O3 |
-1.541 |
-0.470 |
-0.116 |
| H4 |
-0.716 |
1.193 |
0.958 |
| H5 |
-0.738 |
1.276 |
-0.815 |
| H6 |
-1.449 |
-1.063 |
0.676 |
| N7 |
1.943 |
-0.301 |
-0.019 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4829 | 1.4683 | 1.0978 | 1.0915 | 2.0020 | 2.6699 |
C2 | 1.4829 | | 2.4542 | 2.1104 | 2.1129 | 2.6634 | 1.1874 | O3 | 1.4683 | 2.4542 | | 2.1446 | 2.0444 | 0.9933 | 3.4890 | H4 | 1.0978 | 2.1104 | 2.1446 | | 1.7749 | 2.3886 | 3.2027 | H5 | 1.0915 | 2.1129 | 2.0444 | 1.7749 | | 2.8627 | 3.2108 | H6 | 2.0020 | 2.6634 | 0.9933 | 2.3886 | 2.8627 | | 3.5448 | N7 | 2.6699 | 1.1874 | 3.4890 | 3.2027 | 3.2108 | 3.5448 | |
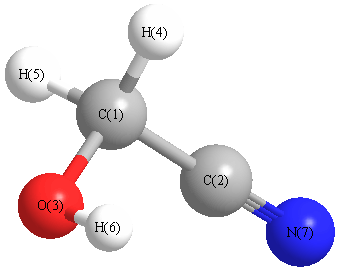 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
177.916 |
|
C1 |
O3 |
H6 |
107.260 |
| C2 |
C1 |
O3 |
112.526 |
|
C2 |
C1 |
H4 |
108.810 |
| C2 |
C1 |
H5 |
109.370 |
|
O3 |
C1 |
H4 |
112.590 |
| O3 |
C1 |
H5 |
105.050 |
|
H4 |
C1 |
H5 |
108.332 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
