return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-191.234083
Energy at 298.15K-191.240309
HF Energy-190.846364
Nuclear repulsion energy114.951650
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3508 3337 1.20      
2 A 3261 3102 15.98      
3 A 3208 3052 3.01      
4 A 3172 3018 8.16      
5 A 3062 2912 57.45      
6 A 3027 2880 40.96      
7 A 1698 1615 0.77      
8 A 1617 1539 1.62      
9 A 1537 1462 6.14      
10 A 1480 1408 5.86      
11 A 1363 1297 0.12      
12 A 1301 1237 33.18      
13 A 1272 1210 32.14      
14 A 1179 1122 11.47      
15 A 1051 1000 17.32      
16 A 1042 991 11.53      
17 A 999 950 52.50      
18 A 968 921 41.95      
19 A 919 874 7.68      
20 A 644 613 4.00      
21 A 463 440 4.43      
22 A 315 299 12.45      
23 A 283 269 149.38      
24 A 98 94 1.06      

Unscaled Zero Point Vibrational Energy (zpe) 18732.3 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 17820.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.93896 0.14008 0.13198

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.573 0.510 0.232
C2 -0.665 -0.359 0.228
C3 -1.872 0.049 -0.189
O4 1.671 -0.342 -0.268
H5 0.408 1.395 -0.401
H6 0.778 0.860 1.256
H7 -0.512 -1.361 0.620
H8 -2.741 -0.599 -0.149
H9 -2.032 1.046 -0.593
H10 2.505 0.194 -0.217

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.51262.52361.47691.10021.10092.19773.51592.78472.0083
C21.51261.34062.38782.14982.15011.08692.12402.12533.2483
C32.52361.34063.56512.65593.12502.11951.08521.08764.3792
O41.47692.38783.56512.15142.13612.56724.42133.96770.9928
H51.10022.14982.65592.15141.78013.07943.73602.47222.4230
H61.10092.15013.12502.13611.78012.64594.06023.36872.3657
H72.19771.08692.11952.56723.07942.64592.47813.09443.4959
H83.51592.12401.08524.42133.73604.06022.47811.84595.3063
H92.78472.12531.08763.96772.47223.36873.09441.84594.6312
H102.00833.24834.37920.99282.42302.36573.49595.30634.6312

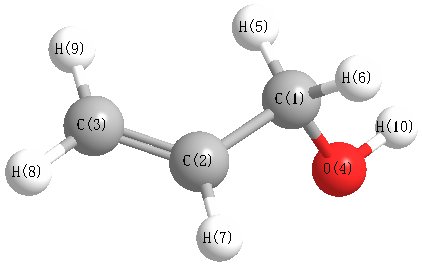
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.257 C1 C2 H7 114.439
C1 O4 H10 107.185 C2 C1 O4 106.006
C2 C1 H5 109.721 C2 C1 H6 109.704
C2 C3 H8 121.871 C2 C3 H9 121.801
C3 C2 H7 121.289 O4 C1 H5 112.367
O4 C1 H6 111.084 H5 C1 H6 107.941
H8 C3 H9 116.326
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability