return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H14 (Pentane, 2-methyl-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-234.626716
Energy at 298.15K-234.641994
HF Energy-234.069505
Nuclear repulsion energy251.714892
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3161 3007 36.56      
2 A 3155 3001 58.71      
3 A 3152 2998 27.18      
4 A 3146 2993 57.41      
5 A 3145 2992 25.55      
6 A 3139 2986 1.78      
7 A 3128 2976 9.77      
8 A 3095 2944 15.22      
9 A 3081 2931 25.39      
10 A 3078 2928 18.05      
11 A 3073 2924 16.10      
12 A 3069 2919 21.90      
13 A 3067 2918 5.08      
14 A 3050 2901 13.08      
15 A 1611 1533 13.50      
16 A 1606 1528 2.79      
17 A 1605 1526 1.81      
18 A 1603 1525 9.64      
19 A 1596 1518 2.82      
20 A 1593 1516 0.31      
21 A 1592 1514 0.68      
22 A 1577 1500 0.24      
23 A 1497 1424 6.42      
24 A 1492 1419 2.84      
25 A 1479 1407 6.59      
26 A 1435 1366 1.95      
27 A 1424 1355 2.41      
28 A 1415 1347 1.22      
29 A 1394 1326 0.30      
30 A 1365 1298 0.13      
31 A 1339 1274 1.16      
32 A 1264 1203 0.60      
33 A 1237 1177 4.33      
34 A 1202 1143 2.50      
35 A 1108 1054 3.62      
36 A 1084 1031 0.44      
37 A 1044 994 1.05      
38 A 1004 955 0.52      
39 A 981 933 0.73      
40 A 970 923 3.67      
41 A 923 878 1.41      
42 A 898 854 0.81      
43 A 820 780 0.47      
44 A 770 732 3.85      
45 A 459 436 0.12      
46 A 452 430 0.27      
47 A 388 369 0.11      
48 A 331 314 0.02      
49 A 271 258 0.00      
50 A 258 246 0.00      
51 A 244 232 0.01      
52 A 187 178 0.05      
53 A 116 110 0.01      
54 A 80 76 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 42624.2 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 40548.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.21767 0.05608 0.04835

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.855 -0.216 0.165
H2 3.694 0.391 -0.191
H3 2.883 -0.236 1.261
H4 2.987 -1.240 -0.201
C5 -1.443 1.387 0.205
H6 -0.711 2.143 -0.098
H7 -2.429 1.698 -0.160
H8 -1.475 1.350 1.301
C9 1.501 0.360 -0.326
H10 1.396 1.391 0.029
H11 1.490 0.387 -1.424
C12 0.304 -0.491 0.167
H13 0.286 -0.492 1.267
C14 -1.078 -0.012 -0.351
H15 -1.029 0.046 -1.448
C16 -2.175 -1.036 0.040
H17 -1.944 -2.027 -0.370
H18 -2.235 -1.116 1.132
H19 -3.154 -0.720 -0.340
H20 0.461 -1.530 -0.159

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 H8 C9 H10 H11 C12 H13 C14 H15 C16 H17 H18 H19 H20
C11.09571.09591.09594.58714.28335.62924.74211.55082.17492.18002.56552.80903.97224.21405.09815.15755.25836.05112.7506
H21.09571.77791.77815.24804.74176.26155.46532.19732.51662.52573.52153.81144.79254.90016.04516.13806.25936.93963.7617
H31.09591.77791.77654.73914.51845.82864.63722.18662.52483.08802.81262.60954.28214.76645.26435.40065.19416.26393.0913
H41.09591.77811.77655.16635.01276.16165.37322.18693.08362.52662.80943.16384.24924.39755.17164.99655.39066.16442.5431
C54.58715.24804.73915.16631.09471.09651.09733.16332.84403.50092.56562.76571.54942.16832.53713.49822.78422.76863.5027
H64.28334.74174.51845.01271.09471.77611.78032.85042.24023.11232.83523.13042.20092.51423.50284.35673.80173.77143.8561
H75.62926.26155.82866.16161.09651.77611.77954.15543.84184.32163.51713.76872.18762.51922.75313.76183.10212.53054.3328
H84.74215.46534.63725.37321.09731.78031.77953.53373.14004.14062.80022.54842.17743.07492.78833.79642.58543.12983.7647
C91.55082.19732.18662.18693.16332.85044.15543.53371.09601.09821.54962.17782.60672.78593.94984.19204.27384.77912.1646
H102.17492.51662.52483.08362.84402.24023.84183.14001.09601.76932.18042.51232.86983.14214.31794.79574.54775.02933.0735
H112.18002.52573.08802.52663.50093.11234.32164.14061.09821.76932.16983.07642.81212.54244.19544.32814.76094.89572.5173
C122.56553.52152.81262.80942.56562.83523.51712.80021.54962.18042.16981.10031.55192.16182.54172.77522.78723.50251.1001
H132.80903.81142.60953.16382.76573.13043.76872.54842.17782.51233.07641.10032.16973.06402.80313.16332.59993.80321.7720
C143.97224.79254.28214.24921.54942.20092.18762.17742.60672.86982.81211.55192.16971.09971.55042.19282.18032.19302.1704
H154.21404.90014.76644.39752.16832.51422.51923.07492.78593.14212.54242.16183.06401.09972.16712.50913.07552.51572.5233
C165.09816.04515.26435.17162.53713.50282.75312.78833.94984.31794.19542.54172.80311.55042.16711.09631.09701.09642.6891
H175.15756.13805.40064.99653.49824.35673.76183.79644.19204.79574.32812.77523.16332.19282.50911.09631.78051.78092.4649
H185.25836.25935.19415.39062.78423.80173.10212.58544.27384.54774.76092.78722.59992.18033.07551.09701.78051.77993.0173
H196.05116.93966.26396.16442.76863.77142.53053.12984.77915.02934.89573.50253.80322.19302.51571.09641.78091.77993.7089
H202.75063.76173.09132.54313.50273.85614.33283.76472.16463.07352.51731.10011.77202.17042.52332.68912.46493.01733.7089

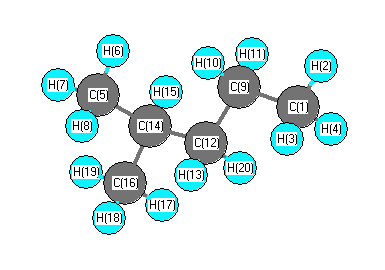
picture of Pentane, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C9 H10 109.306 C1 C9 H11 109.581
C1 C9 C12 111.683 H2 C1 H3 108.442
H2 C1 H4 108.454 H2 C1 C9 111.090
H3 C1 H4 108.295 H3 C1 C9 110.232
H4 C1 C9 110.251 C5 C14 C12 111.640
C5 C14 H15 108.689 C5 C14 C16 109.869
H6 C5 H7 108.303 H6 C5 H8 108.624
H6 C5 C14 111.532 H7 C5 H8 108.416
H7 C5 C14 110.366 H8 C5 C14 109.526
C9 C12 H13 109.368 C9 C12 C14 114.376
C9 C12 H20 108.356 H10 C9 H11 107.487
H10 C9 C12 109.821 H11 C9 C12 108.867
C12 C14 H15 108.011 C12 C14 C16 110.025
H13 C12 C14 108.586 H13 C12 H20 107.281
C14 C12 H20 108.644 C14 C16 H17 110.716
C14 C16 H18 109.699 C14 C16 H19 110.727
H15 C14 C16 108.526 H17 C16 H18 108.542
H17 C16 H19 108.616 H18 C16 H19 108.479
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability