return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Pentane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-195.713575
Energy at 298.15K-195.726566
HF Energy-195.250266
Nuclear repulsion energy183.321169
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3152 2999 39.24      
2 A1 3081 2931 68.70      
3 A1 3079 2929 0.25      
4 A1 3062 2913 4.02      
5 A1 1609 1531 6.78      
6 A1 1597 1519 0.43      
7 A1 1583 1506 0.00      
8 A1 1492 1419 4.33      
9 A1 1425 1355 0.60      
10 A1 1200 1141 1.62      
11 A1 1029 979 0.33      
12 A1 889 846 1.18      
13 A1 404 384 0.00      
14 A1 183 174 0.06      
15 A2 3154 3000 0.00      
16 A2 3113 2961 0.00      
17 A2 1604 1526 0.00      
18 A2 1397 1329 0.00      
19 A2 1339 1274 0.00      
20 A2 1060 1009 0.00      
21 A2 802 763 0.00      
22 A2 260 247 0.00      
23 A2 117 112 0.00      
24 B1 3156 3003 103.33      
25 B1 3128 2976 20.98      
26 B1 3095 2945 0.31      
27 B1 1603 1525 13.29      
28 B1 1394 1326 0.60      
29 B1 1275 1213 0.21      
30 B1 920 875 1.83      
31 B1 755 718 5.55      
32 B1 255 242 0.00      
33 B1 115 109 0.00      
34 B2 3152 2999 23.88      
35 B2 3079 2929 31.12      
36 B2 3075 2925 0.01      
37 B2 1606 1527 2.14      
38 B2 1587 1510 0.38      
39 B2 1493 1420 2.41      
40 B2 1434 1364 1.19      
41 B2 1360 1293 0.39      
42 B2 1097 1044 7.75      
43 B2 1050 998 1.32      
44 B2 922 877 1.62      
45 B2 410 390 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 36295.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 34527.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.55505 0.06369 0.06037

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.319
C2 0.000 1.290 -0.538
C3 0.000 -1.290 -0.538
C4 0.000 2.572 0.335
C5 0.000 -2.572 0.335
H6 0.885 0.000 0.971
H7 -0.885 0.000 0.971
H8 0.885 1.287 -1.188
H9 -0.885 1.287 -1.188
H10 -0.885 -1.287 -1.188
H11 0.885 -1.287 -1.188
H12 0.000 3.473 -0.287
H13 0.000 -3.473 -0.287
H14 -0.888 2.593 0.976
H15 0.888 2.593 0.976
H16 0.888 -2.593 0.976
H17 -0.888 -2.593 0.976

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.54921.54922.57162.57161.09911.09912.17062.17062.17062.17063.52593.52592.81882.81882.81882.8188
C21.54922.58081.55033.95942.17362.17361.09781.09782.80152.80152.19724.77032.18622.18624.26204.2620
C31.54922.58083.95941.55032.17362.17362.80152.80151.09781.09784.77032.19724.26204.26202.18622.1862
C42.57161.55033.95945.14322.79302.79302.18002.18004.24164.24161.09566.07691.09581.09585.27985.2798
C52.57163.95941.55035.14322.79302.79304.24164.24162.18002.18006.07691.09565.27985.27981.09581.0958
H61.09912.17362.17362.79302.79301.77072.51333.07393.07392.51333.79873.79873.14172.59332.59333.1417
H71.09912.17362.17362.79302.79301.77073.07392.51332.51333.07393.79873.79872.59333.14173.14172.5933
H82.17061.09782.80152.18004.24162.51333.07391.76903.12332.57412.52464.92493.08782.52824.44324.7838
H92.17061.09782.80152.18004.24163.07392.51331.76902.57413.12332.52464.92492.52823.08784.78384.4432
H102.17062.80151.09784.24162.18003.07392.51333.12332.57411.76904.92492.52464.44324.78383.08782.5282
H112.17062.80151.09784.24162.18002.51333.07392.57413.12331.76904.92492.52464.78384.44322.52823.0878
H123.52592.19724.77031.09566.07693.79873.79872.52462.52464.92494.92496.94661.77781.77786.26016.2601
H133.52594.77032.19726.07691.09563.79873.79874.92494.92492.52462.52466.94666.26016.26011.77781.7778
H142.81882.18624.26201.09585.27983.14172.59333.08782.52824.44324.78381.77786.26011.77645.48235.1865
H152.81882.18624.26201.09585.27982.59333.14172.52823.08784.78384.44321.77786.26011.77645.18655.4823
H162.81884.26202.18625.27981.09582.59333.14174.44324.78383.08782.52826.26011.77785.48235.18651.7764
H172.81884.26202.18625.27981.09583.14172.59334.78384.44322.52823.08786.26011.77785.18655.48231.7764

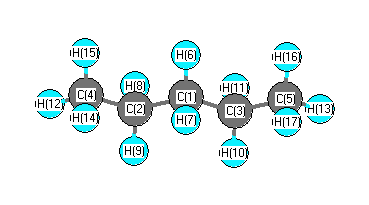
picture of Pentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.136 C1 C2 H8 108.978
C1 C2 H9 108.978 C1 C3 C5 112.136
C1 C3 H10 108.978 C1 C3 H11 108.978
C2 C1 C3 112.806 C2 C1 H6 109.141
C2 C1 H7 109.141 C2 C4 H12 111.122
C2 C4 H14 110.238 C2 C4 H15 110.238
C3 C1 H6 109.141 C3 C1 H7 109.141
C3 C5 H13 111.122 C3 C5 H16 110.238
C3 C5 H17 110.238 C4 C2 H8 109.636
C4 C2 H9 109.636 C5 C3 H10 109.636
C5 C3 H11 109.636 H6 C1 H7 107.323
H8 C2 H9 107.355 H10 C3 H11 107.355
H12 C4 H14 108.434 H12 C4 H15 108.434
H13 C5 H16 108.434 H13 C5 H17 108.434
H14 C4 H15 108.297 H16 C5 H17 108.297
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability