Jump to
S1C2
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -194.516150 |
| Energy at 298.15K | -194.526429 |
| HF Energy | -194.060438 |
| Nuclear repulsion energy | 168.778637 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3255 |
3096 |
19.03 |
|
|
|
| 2 |
A |
3173 |
3018 |
4.23 |
|
|
|
| 3 |
A |
3161 |
3007 |
38.48 |
|
|
|
| 4 |
A |
3156 |
3002 |
30.80 |
|
|
|
| 5 |
A |
3154 |
3000 |
29.16 |
|
|
|
| 6 |
A |
3129 |
2977 |
7.41 |
|
|
|
| 7 |
A |
3103 |
2952 |
4.02 |
|
|
|
| 8 |
A |
3085 |
2934 |
28.20 |
|
|
|
| 9 |
A |
3079 |
2929 |
18.13 |
|
|
|
| 10 |
A |
3062 |
2913 |
12.74 |
|
|
|
| 11 |
A |
1691 |
1609 |
3.95 |
|
|
|
| 12 |
A |
1606 |
1528 |
4.73 |
|
|
|
| 13 |
A |
1603 |
1525 |
6.64 |
|
|
|
| 14 |
A |
1591 |
1514 |
1.61 |
|
|
|
| 15 |
A |
1575 |
1498 |
2.35 |
|
|
|
| 16 |
A |
1533 |
1458 |
0.92 |
|
|
|
| 17 |
A |
1492 |
1419 |
3.20 |
|
|
|
| 18 |
A |
1424 |
1355 |
0.97 |
|
|
|
| 19 |
A |
1394 |
1326 |
0.38 |
|
|
|
| 20 |
A |
1376 |
1309 |
0.19 |
|
|
|
| 21 |
A |
1363 |
1296 |
0.54 |
|
|
|
| 22 |
A |
1332 |
1267 |
0.10 |
|
|
|
| 23 |
A |
1242 |
1182 |
0.80 |
|
|
|
| 24 |
A |
1153 |
1097 |
2.26 |
|
|
|
| 25 |
A |
1086 |
1033 |
6.40 |
|
|
|
| 26 |
A |
1050 |
999 |
15.42 |
|
|
|
| 27 |
A |
1035 |
985 |
1.27 |
|
|
|
| 28 |
A |
960 |
913 |
41.23 |
|
|
|
| 29 |
A |
949 |
903 |
0.59 |
|
|
|
| 30 |
A |
924 |
879 |
2.83 |
|
|
|
| 31 |
A |
903 |
859 |
5.02 |
|
|
|
| 32 |
A |
771 |
733 |
3.27 |
|
|
|
| 33 |
A |
652 |
621 |
10.00 |
|
|
|
| 34 |
A |
453 |
431 |
0.95 |
|
|
|
| 35 |
A |
389 |
370 |
0.18 |
|
|
|
| 36 |
A |
256 |
244 |
0.03 |
|
|
|
| 37 |
A |
243 |
232 |
0.17 |
|
|
|
| 38 |
A |
107 |
102 |
0.00 |
|
|
|
| 39 |
A |
90 |
86 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30798.8 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29298.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.465 |
-0.240 |
-0.327 |
| C2 |
1.429 |
0.357 |
0.283 |
| C3 |
0.069 |
-0.285 |
0.498 |
| C4 |
-1.041 |
0.435 |
-0.320 |
| C5 |
-2.435 |
-0.199 |
-0.087 |
| H6 |
3.414 |
0.266 |
-0.480 |
| H7 |
2.397 |
-1.263 |
-0.691 |
| H8 |
1.528 |
1.385 |
0.634 |
| H9 |
0.111 |
-1.341 |
0.196 |
| H10 |
-0.198 |
-0.253 |
1.564 |
| H11 |
-1.063 |
1.495 |
-0.039 |
| H12 |
-0.779 |
0.382 |
-1.384 |
| H13 |
-3.203 |
0.316 |
-0.673 |
| H14 |
-2.426 |
-1.255 |
-0.379 |
| H15 |
-2.708 |
-0.138 |
0.973 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3426 | 2.5343 | 3.5700 | 4.9063 | 1.0859 | 1.0876 | 2.1079 | 2.6507 | 3.2659 | 3.9422 | 3.4675 | 5.7062 | 4.9954 | 5.3350 |
C2 | 1.3426 | | 1.5191 | 2.5433 | 3.9214 | 2.1284 | 2.1240 | 1.0905 | 2.1513 | 2.1584 | 2.7582 | 2.7661 | 4.7301 | 4.2306 | 4.2233 | C3 | 2.5343 | 1.5191 | | 1.5557 | 2.5733 | 3.5278 | 2.7909 | 2.2216 | 1.0988 | 1.0991 | 2.1775 | 2.1692 | 3.5277 | 2.8172 | 2.8216 | C4 | 3.5700 | 2.5433 | 1.5557 | | 1.5496 | 4.4603 | 3.8521 | 2.9001 | 2.1790 | 2.1755 | 1.0976 | 1.0966 | 2.1946 | 2.1858 | 2.1866 | C5 | 4.9063 | 3.9214 | 2.5733 | 1.5496 | | 5.8805 | 4.9851 | 4.3283 | 2.8053 | 2.7807 | 2.1810 | 2.1829 | 1.0955 | 1.0958 | 1.0959 | H6 | 1.0859 | 2.1284 | 3.5278 | 4.4603 | 5.8805 | | 1.8476 | 2.4598 | 3.7343 | 4.1820 | 4.6633 | 4.2900 | 6.6201 | 6.0352 | 6.3048 | H7 | 1.0876 | 2.1240 | 2.7909 | 3.8521 | 4.9851 | 1.8476 | | 3.0859 | 2.4535 | 3.5832 | 4.4729 | 3.6430 | 5.8193 | 4.8334 | 5.4865 | H8 | 2.1079 | 1.0905 | 2.2216 | 2.9001 | 4.3283 | 2.4598 | 3.0859 | | 3.1031 | 2.5542 | 2.6791 | 3.2245 | 5.0235 | 4.8610 | 4.5140 | H9 | 2.6507 | 2.1513 | 1.0988 | 2.1790 | 2.8053 | 3.7343 | 2.4535 | 3.1031 | | 1.7748 | 3.0790 | 2.5015 | 3.8068 | 2.6033 | 3.1625 | H10 | 3.2659 | 2.1584 | 1.0991 | 2.1755 | 2.7807 | 4.1820 | 3.5832 | 2.5542 | 1.7748 | | 2.5245 | 3.0706 | 3.7897 | 3.1218 | 2.5815 | H11 | 3.9422 | 2.7582 | 2.1775 | 1.0976 | 2.1810 | 4.6633 | 4.4729 | 2.6791 | 3.0790 | 2.5245 | | 1.7690 | 2.5247 | 3.0884 | 2.5294 | H12 | 3.4675 | 2.7661 | 2.1692 | 1.0966 | 2.1829 | 4.2900 | 3.6430 | 3.2245 | 2.5015 | 3.0706 | 1.7690 | | 2.5278 | 2.5302 | 3.0900 | H13 | 5.7062 | 4.7301 | 3.5277 | 2.1946 | 1.0955 | 6.6201 | 5.8193 | 5.0235 | 3.8068 | 3.7897 | 2.5247 | 2.5278 | | 1.7776 | 1.7781 | H14 | 4.9954 | 4.2306 | 2.8172 | 2.1858 | 1.0958 | 6.0352 | 4.8334 | 4.8610 | 2.6033 | 3.1218 | 3.0884 | 2.5302 | 1.7776 | | 1.7769 | H15 | 5.3350 | 4.2233 | 2.8216 | 2.1866 | 1.0959 | 6.3048 | 5.4865 | 4.5140 | 3.1625 | 2.5815 | 2.5294 | 3.0900 | 1.7781 | 1.7769 | |
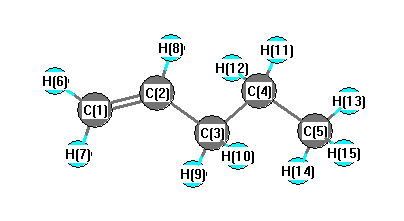 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.527 |
|
C1 |
C2 |
H8 |
119.711 |
| C2 |
C1 |
H6 |
122.068 |
|
C2 |
C1 |
H7 |
121.495 |
| C2 |
C3 |
C4 |
111.602 |
|
C2 |
C3 |
H9 |
109.470 |
| C2 |
C3 |
H10 |
110.015 |
|
C3 |
C2 |
H8 |
115.737 |
| C3 |
C4 |
C5 |
111.923 |
|
C3 |
C4 |
H11 |
109.083 |
| C3 |
C4 |
H12 |
108.503 |
|
C4 |
C3 |
H9 |
109.123 |
| C4 |
C3 |
H10 |
108.842 |
|
C4 |
C5 |
H13 |
110.971 |
| C4 |
C5 |
H14 |
110.255 |
|
C4 |
C5 |
H15 |
110.310 |
| C5 |
C4 |
H11 |
109.773 |
|
C5 |
C4 |
H12 |
109.980 |
| H6 |
C1 |
H7 |
116.436 |
|
H9 |
C3 |
H10 |
107.700 |
| H11 |
C4 |
H12 |
107.464 |
|
H13 |
C5 |
H14 |
108.431 |
| H13 |
C5 |
H15 |
108.466 |
|
H14 |
C5 |
H15 |
108.335 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/3-21G*
| | hartrees |
|---|
| Energy at 0K | -194.516587 |
| Energy at 298.15K | -194.526961 |
| HF Energy | -194.060422 |
| Nuclear repulsion energy | 172.688652 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3254 |
3096 |
17.90 |
|
|
|
| 2 |
A |
3172 |
3017 |
4.81 |
|
|
|
| 3 |
A |
3171 |
3016 |
26.21 |
|
|
|
| 4 |
A |
3157 |
3004 |
28.86 |
|
|
|
| 5 |
A |
3152 |
2998 |
26.19 |
|
|
|
| 6 |
A |
3123 |
2970 |
17.35 |
|
|
|
| 7 |
A |
3109 |
2958 |
20.80 |
|
|
|
| 8 |
A |
3083 |
2933 |
16.83 |
|
|
|
| 9 |
A |
3079 |
2929 |
19.09 |
|
|
|
| 10 |
A |
3062 |
2913 |
18.73 |
|
|
|
| 11 |
A |
1690 |
1608 |
3.31 |
|
|
|
| 12 |
A |
1609 |
1530 |
3.88 |
|
|
|
| 13 |
A |
1601 |
1523 |
7.25 |
|
|
|
| 14 |
A |
1589 |
1511 |
1.93 |
|
|
|
| 15 |
A |
1573 |
1496 |
5.19 |
|
|
|
| 16 |
A |
1532 |
1458 |
1.30 |
|
|
|
| 17 |
A |
1491 |
1419 |
5.73 |
|
|
|
| 18 |
A |
1434 |
1364 |
2.29 |
|
|
|
| 19 |
A |
1406 |
1338 |
0.78 |
|
|
|
| 20 |
A |
1378 |
1311 |
0.72 |
|
|
|
| 21 |
A |
1350 |
1284 |
0.44 |
|
|
|
| 22 |
A |
1315 |
1251 |
1.34 |
|
|
|
| 23 |
A |
1238 |
1178 |
0.63 |
|
|
|
| 24 |
A |
1146 |
1090 |
4.14 |
|
|
|
| 25 |
A |
1087 |
1034 |
5.28 |
|
|
|
| 26 |
A |
1051 |
999 |
14.98 |
|
|
|
| 27 |
A |
1048 |
997 |
1.16 |
|
|
|
| 28 |
A |
959 |
913 |
45.36 |
|
|
|
| 29 |
A |
936 |
890 |
0.57 |
|
|
|
| 30 |
A |
926 |
881 |
1.30 |
|
|
|
| 31 |
A |
869 |
826 |
1.99 |
|
|
|
| 32 |
A |
798 |
759 |
3.97 |
|
|
|
| 33 |
A |
656 |
624 |
9.07 |
|
|
|
| 34 |
A |
443 |
421 |
0.34 |
|
|
|
| 35 |
A |
408 |
388 |
2.57 |
|
|
|
| 36 |
A |
310 |
295 |
0.16 |
|
|
|
| 37 |
A |
237 |
225 |
0.21 |
|
|
|
| 38 |
A |
126 |
120 |
0.02 |
|
|
|
| 39 |
A |
87 |
83 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30826.8 cm
-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 29325.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/3-21G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.060 |
0.593 |
0.114 |
| C2 |
-1.186 |
-0.290 |
-0.395 |
| C3 |
-0.053 |
-0.936 |
0.384 |
| C4 |
1.341 |
-0.518 |
-0.172 |
| C5 |
1.605 |
0.996 |
0.017 |
| H6 |
-2.848 |
1.036 |
-0.487 |
| H7 |
-2.018 |
0.897 |
1.158 |
| H8 |
-1.255 |
-0.573 |
-1.446 |
| H9 |
-0.150 |
-2.030 |
0.334 |
| H10 |
-0.122 |
-0.642 |
1.441 |
| H11 |
2.119 |
-1.095 |
0.342 |
| H12 |
1.390 |
-0.773 |
-1.239 |
| H13 |
2.582 |
1.277 |
-0.393 |
| H14 |
0.829 |
1.582 |
-0.484 |
| H15 |
1.592 |
1.249 |
1.083 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3429 | 2.5376 | 3.5891 | 3.6891 | 1.0860 | 1.0876 | 2.1081 | 3.2522 | 2.6533 | 4.5129 | 3.9503 | 4.7194 | 3.1124 | 3.8351 |
C2 | 1.3429 | | 1.5191 | 2.5462 | 3.1005 | 2.1288 | 2.1239 | 1.0909 | 2.1522 | 2.1506 | 3.4803 | 2.7536 | 4.0799 | 2.7517 | 3.5026 | C3 | 2.5376 | 1.5191 | | 1.5580 | 2.5727 | 3.5303 | 2.7958 | 2.2194 | 1.0989 | 1.0990 | 2.1783 | 2.1783 | 3.5275 | 2.8060 | 2.8231 | C4 | 3.5891 | 2.5462 | 1.5580 | | 1.5484 | 4.4790 | 3.8791 | 2.8914 | 2.1823 | 2.1811 | 1.0972 | 1.0976 | 2.1930 | 2.1835 | 2.1823 | C5 | 3.6891 | 3.1005 | 2.5727 | 1.5484 | | 4.4823 | 3.7996 | 3.5752 | 3.5121 | 2.7736 | 2.1775 | 2.1802 | 1.0955 | 1.0939 | 1.0962 | H6 | 1.0860 | 2.1288 | 3.5303 | 4.4790 | 4.4823 | | 1.8479 | 2.4598 | 4.1663 | 3.7369 | 5.4684 | 4.6698 | 5.4360 | 3.7179 | 4.7145 | H7 | 1.0876 | 2.1239 | 2.7958 | 3.8791 | 3.7996 | 1.8479 | | 3.0859 | 3.5682 | 2.4575 | 4.6628 | 4.4885 | 4.8685 | 3.3572 | 3.6274 | H8 | 2.1081 | 1.0909 | 2.2194 | 2.8914 | 3.5752 | 2.4598 | 3.0859 | | 2.5517 | 3.1019 | 3.8538 | 2.6607 | 4.3870 | 3.1484 | 4.2215 | H9 | 3.2522 | 2.1522 | 1.0989 | 2.1823 | 3.5121 | 4.1663 | 3.5682 | 2.5517 | | 1.7757 | 2.4535 | 2.5344 | 4.3497 | 3.8303 | 3.7874 | H10 | 2.6533 | 2.1506 | 1.0990 | 2.1811 | 2.7736 | 3.7369 | 2.4575 | 3.1019 | 1.7757 | | 2.5365 | 3.0798 | 3.7886 | 3.0911 | 2.5767 | H11 | 4.5129 | 3.4803 | 2.1783 | 1.0972 | 2.1775 | 5.4684 | 4.6628 | 3.8538 | 2.4535 | 2.5365 | | 1.7704 | 2.5259 | 3.0841 | 2.5142 | H12 | 3.9503 | 2.7536 | 2.1783 | 1.0976 | 2.1802 | 4.6698 | 4.4885 | 2.6607 | 2.5344 | 3.0798 | 1.7704 | | 2.5171 | 2.5360 | 3.0859 | H13 | 4.7194 | 4.0799 | 3.5275 | 2.1930 | 1.0955 | 5.4360 | 4.8685 | 4.3870 | 4.3497 | 3.7886 | 2.5259 | 2.5171 | | 1.7809 | 1.7780 | H14 | 3.1124 | 2.7517 | 2.8060 | 2.1835 | 1.0939 | 3.7179 | 3.3572 | 3.1484 | 3.8303 | 3.0911 | 3.0841 | 2.5360 | 1.7809 | | 1.7748 | H15 | 3.8351 | 3.5026 | 2.8231 | 2.1823 | 1.0962 | 4.7145 | 3.6274 | 4.2215 | 3.7874 | 2.5767 | 2.5142 | 3.0859 | 1.7780 | 1.7748 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.793 |
|
C1 |
C2 |
H8 |
119.673 |
| C2 |
C1 |
H6 |
122.073 |
|
C2 |
C1 |
H7 |
121.463 |
| C2 |
C3 |
C4 |
111.677 |
|
C2 |
C3 |
H9 |
109.542 |
| C2 |
C3 |
H10 |
109.415 |
|
C3 |
C2 |
H8 |
115.526 |
| C3 |
C4 |
C5 |
111.826 |
|
C3 |
C4 |
H11 |
109.010 |
| C3 |
C4 |
H12 |
108.985 |
|
C4 |
C3 |
H9 |
109.220 |
| C4 |
C3 |
H10 |
109.124 |
|
C4 |
C5 |
H13 |
110.925 |
| C4 |
C5 |
H14 |
110.263 |
|
C4 |
C5 |
H15 |
110.031 |
| C5 |
C4 |
H11 |
109.602 |
|
C5 |
C4 |
H12 |
109.786 |
| H6 |
C1 |
H7 |
116.462 |
|
H9 |
C3 |
H10 |
107.780 |
| H11 |
C4 |
H12 |
107.530 |
|
H13 |
C5 |
H14 |
108.864 |
| H13 |
C5 |
H15 |
108.434 |
|
H14 |
C5 |
H15 |
108.257 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
