return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2ClCH2CH2CH3 (Butane, 1-chloro-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-613.756947
Energy at 298.15K-613.766910
HF Energy-613.255400
Nuclear repulsion energy216.881137
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3158 3004 26.69      
2 A' 3136 2984 15.78      
3 A' 3098 2947 13.72      
4 A' 3083 2933 18.29      
5 A' 3078 2928 11.63      
6 A' 1607 1529 6.13      
7 A' 1596 1518 3.71      
8 A' 1587 1509 0.39      
9 A' 1578 1501 2.50      
10 A' 1495 1422 3.30      
11 A' 1434 1364 3.22      
12 A' 1405 1337 17.01      
13 A' 1334 1269 9.98      
14 A' 1156 1099 6.77      
15 A' 1054 1002 2.06      
16 A' 1007 958 5.06      
17 A' 918 873 0.31      
18 A' 733 697 40.00      
19 A' 408 388 1.67      
20 A' 337 321 3.18      
21 A' 161 153 1.55      
22 A" 3201 3045 12.67      
23 A" 3161 3007 42.45      
24 A" 3146 2993 2.11      
25 A" 3115 2963 4.28      
26 A" 1603 1525 7.52      
27 A" 1396 1328 0.10      
28 A" 1381 1314 0.94      
29 A" 1302 1238 0.42      
30 A" 1167 1110 0.17      
31 A" 990 941 1.44      
32 A" 834 793 0.00      
33 A" 768 730 5.46      
34 A" 258 245 0.04      
35 A" 123 117 0.77      
36 A" 108 102 0.88      

Unscaled Zero Point Vibrational Energy (zpe) 27956.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 26594.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.54978 0.04313 0.04123

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.195 -1.005 0.000
H2 0.727 -1.336 0.893
H3 0.727 -1.336 -0.893
C4 0.000 0.519 0.000
H5 -0.576 0.810 0.886
H6 -0.576 0.810 -0.886
C7 1.365 1.255 0.000
H8 1.939 0.953 -0.885
H9 1.939 0.953 0.885
C10 1.181 2.794 0.000
H11 0.623 3.110 -0.889
H12 0.623 3.110 0.889
H13 2.151 3.302 0.000
Cl14 -1.413 -1.868 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 H9 C10 H11 H12 H13 Cl14
C11.09121.09121.53672.16172.16172.54492.76772.76773.92504.23134.23134.73071.8248
H21.09121.78612.18392.51083.07722.81403.14192.59024.25014.79074.44724.93372.3794
H31.09121.78612.18393.07722.51082.81402.59023.14194.25014.44724.79074.93372.3794
C41.53672.18392.18391.09611.09611.55072.17552.17552.56312.80862.80863.51742.7738
H52.16172.51083.07721.09611.77222.17973.07992.51952.79463.14282.59383.79932.9417
H62.16173.07722.51081.09611.77222.17972.51953.07992.79462.59383.14283.79932.9417
C72.54492.81402.81401.55072.17972.17971.09761.09761.55022.18632.18632.19314.1795
H82.76773.14192.59022.17553.07992.51951.09761.77042.17932.52683.08722.51974.4697
H92.76772.59023.14192.17552.51953.07991.09761.77042.17933.08722.52682.51974.4697
C103.92504.25014.25012.56312.79462.79461.55022.17932.17931.09551.09551.09515.3348
H114.23134.79074.44722.80863.14282.59382.18632.52683.08721.09551.77711.77785.4506
H124.23134.44724.79072.80862.59383.14282.18633.08722.52681.09551.77711.77785.4506
H134.73074.93374.93373.51743.79933.79932.19312.51972.51971.09511.77781.77786.2794
Cl141.82482.37942.37942.77382.94172.94174.17954.46974.46975.33485.45065.45066.2794

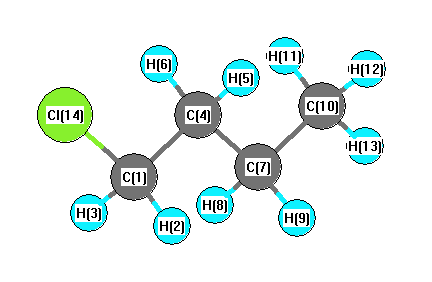
picture of Butane, 1-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 109.241 C1 C4 H6 109.241
C1 C4 C7 111.035 H2 C1 H3 109.859
H2 C1 C4 111.283 H2 C1 Cl14 106.646
H3 C1 C4 111.283 H3 C1 Cl14 106.646
C4 C1 Cl14 110.919 C4 C7 H8 109.273
C4 C7 H9 109.273 C4 C7 C10 111.497
H5 C4 H6 107.884 H5 C4 C7 109.689
H6 C4 C7 109.689 C7 C10 H11 110.274
C7 C10 H12 110.274 C7 C10 H13 110.827
H8 C7 H9 107.505 H8 C7 C10 109.602
H9 C7 C10 109.602 H11 C10 H12 108.407
H11 C10 H13 108.496 H12 C10 H13 108.496
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability