return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8S2 (1,3-Propanedithiol)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-909.455750
Energy at 298.15K-909.464340
HF Energy-908.937146
Nuclear repulsion energy265.761090
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3112 2961 32.37      
2 A1 3098 2947 1.66      
3 A1 2780 2644 4.85      
4 A1 1596 1519 4.55      
5 A1 1585 1508 0.13      
6 A1 1378 1311 34.37      
7 A1 1103 1049 5.00      
8 A1 856 815 0.42      
9 A1 810 771 1.40      
10 A1 304 290 0.38      
11 A1 134 128 0.16      
12 A2 3158 3005 0.00      
13 A2 1398 1330 0.00      
14 A2 1190 1132 0.00      
15 A2 886 843 0.00      
16 A2 194 185 0.00      
17 A2 101 96 0.00      
18 B1 3175 3020 24.63      
19 B1 3141 2988 0.25      
20 B1 1355 1289 2.41      
21 B1 1083 1031 3.94      
22 B1 789 750 4.60      
23 B1 183 174 51.49      
24 B1 100 95 0.00      
25 B2 3104 2953 4.32      
26 B2 2780 2644 15.68      
27 B2 1579 1502 8.86      
28 B2 1431 1362 15.23      
29 B2 1315 1251 6.15      
30 B2 1051 1000 0.19      
31 B2 928 883 0.28      
32 B2 713 678 2.25      
33 B2 348 331 9.06      

Unscaled Zero Point Vibrational Energy (zpe) 23378.8 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 22240.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.47166 0.03004 0.02870

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.177
C2 0.000 1.271 -0.707
C3 0.000 -1.271 -0.707
H4 0.888 0.000 0.819
H5 -0.888 0.000 0.819
S6 0.000 2.751 0.385
S7 0.000 -2.751 0.385
H8 0.000 3.665 -0.593
H9 0.000 -3.665 -0.593
H10 0.892 1.279 -1.340
H11 -0.892 1.279 -1.340
H12 -0.892 -1.279 -1.340
H13 0.892 -1.279 -1.340

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 S6 S7 H8 H9 H10 H11 H12 H13
C11.54851.54851.09571.09572.75872.75873.74533.74532.17562.17562.17562.1756
C21.54852.54252.17572.17571.83914.16782.39674.93781.09391.09392.77492.7749
C31.54852.54252.17572.17574.16781.83914.93782.39672.77492.77491.09391.0939
H41.09572.17572.17571.77582.92302.92304.02704.02702.50953.07673.07672.5095
H51.09572.17572.17571.77582.92302.92304.02704.02703.07672.50952.50953.0767
S62.75871.83914.16782.92302.92305.50171.33906.49022.43692.43694.47344.4734
S72.75874.16781.83912.92302.92305.50176.49021.33904.47344.47342.43692.4369
H83.74532.39674.93784.02704.02701.33906.49027.33042.65482.65485.07925.0792
H93.74534.93782.39674.02704.02706.49021.33907.33045.07925.07922.65482.6548
H102.17561.09392.77492.50953.07672.43694.47342.65485.07921.78443.11882.5579
H112.17561.09392.77493.07672.50952.43694.47342.65485.07921.78442.55793.1188
H122.17562.77491.09393.07672.50954.47342.43695.07922.65483.11882.55791.7844
H132.17562.77491.09392.50953.07674.47342.43695.07922.65482.55793.11881.7844

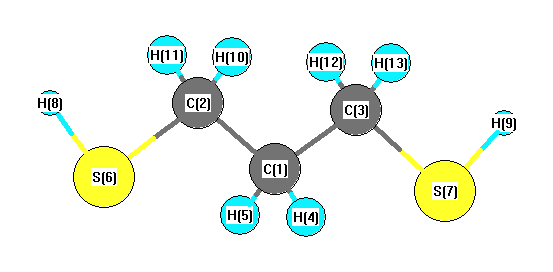
picture of 1,3-Propanedithiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 S6 108.746 C1 C2 H10 109.642
C1 C2 H11 109.642 C1 C3 S7 108.746
C1 C3 H12 109.642 C1 C3 H13 109.642
C2 C1 C3 110.359 C2 C1 H4 109.548
C2 C1 H5 109.548 C2 S6 H8 96.634
C3 C1 H4 109.548 C3 C1 H5 109.548
C3 S7 H9 96.634 H4 C1 H5 108.254
S6 C2 H10 109.748 S6 C2 H11 109.748
S7 C3 H12 109.748 S7 C3 H13 109.748
H10 C2 H11 109.300 H12 C3 H13 109.300
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability