return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-211.612320
Energy at 298.15K-211.625065
HF Energy-211.135832
Nuclear repulsion energy186.174032
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3399 3233 3.19      
2 A' 3181 3026 36.81      
3 A' 3155 3001 55.03      
4 A' 3111 2959 36.17      
5 A' 3087 2936 8.92      
6 A' 2988 2843 130.86      
7 A' 1625 1546 0.41      
8 A' 1606 1528 2.89      
9 A' 1595 1517 12.34      
10 A' 1487 1415 5.57      
11 A' 1460 1389 0.41      
12 A' 1368 1302 1.46      
13 A' 1290 1227 3.72      
14 A' 1191 1133 6.67      
15 A' 1032 982 0.25      
16 A' 938 892 15.15      
17 A' 866 824 5.04      
18 A' 739 703 85.14      
19 A' 422 402 0.32      
20 A' 271 258 1.50      
21 A' 186 177 1.22      
22 A' 105 99 1.67      
23 A" 3181 3026 6.05      
24 A" 3154 3001 15.24      
25 A" 3108 2957 1.36      
26 A" 3086 2936 21.80      
27 A" 2986 2840 6.77      
28 A" 1616 1538 1.60      
29 A" 1604 1526 3.51      
30 A" 1597 1520 3.78      
31 A" 1530 1456 9.06      
32 A" 1485 1412 10.49      
33 A" 1408 1340 32.41      
34 A" 1351 1285 5.08      
35 A" 1165 1108 2.18      
36 A" 1133 1078 55.41      
37 A" 1087 1034 3.36      
38 A" 938 892 0.31      
39 A" 856 814 1.15      
40 A" 424 403 1.36      
41 A" 268 255 1.00      
42 A" 120 115 1.52      

Unscaled Zero Point Vibrational Energy (zpe) 33597.9 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 31961.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.56793 0.06859 0.06447

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.284 0.000
C2 0.017 0.541 1.246
C3 0.017 0.541 -1.246
C4 0.017 -0.388 2.478
C5 0.017 -0.388 -2.478
H6 -0.808 -0.898 0.000
H7 -0.835 1.244 1.302
H8 0.934 1.141 1.235
H9 -0.835 1.244 -1.302
H10 0.934 1.141 -1.235
H11 0.092 0.190 3.404
H12 -0.908 -0.976 2.512
H13 0.864 -1.075 2.406
H14 0.092 0.190 -3.404
H15 -0.908 -0.976 -2.512
H16 0.864 -1.075 -2.406

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.49411.49412.47982.47981.02872.18022.09662.18022.09663.43782.76482.67093.43782.76482.6709
C21.49412.49131.54353.83762.07471.10551.09602.77642.71182.18832.18212.16274.66374.15664.0824
C31.49412.49133.83761.54352.07472.77642.71181.10551.09604.66374.15664.08242.18832.18212.1627
C42.47981.54353.83764.95512.66062.18462.17364.20404.11861.09511.09611.09305.91065.10835.0039
C52.47983.83761.54354.95512.66064.20404.11862.18462.17365.91065.10835.00391.09511.09611.0930
H61.02872.07472.07472.66062.66062.50662.95272.50662.95273.68572.51512.93573.68572.51512.9357
H72.18021.10552.77642.18464.20402.50661.77362.60373.09452.52792.52953.07954.91094.41344.6918
H82.09661.09602.71182.17364.11862.95271.77363.09452.46982.51363.08312.50704.80974.68124.2626
H92.18022.77641.10554.20402.18462.50662.60373.09451.77364.91094.41344.69182.52792.52953.0795
H102.09662.71181.09604.11862.17362.95273.09452.46981.77364.80974.68124.26262.51363.08312.5070
H113.43782.18834.66371.09515.91063.68572.52792.51364.91094.80971.77691.78676.80846.11245.9962
H122.76482.18214.15661.09615.10832.51512.52953.08314.41344.68121.77691.77816.11245.02385.2284
H132.67092.16274.08241.09305.00392.93573.07952.50704.69184.26261.78671.77815.99625.22844.8121
H143.43784.66372.18835.91061.09513.68574.91094.80972.52792.51366.80846.11245.99621.77691.7867
H152.76484.15662.18215.10831.09612.51514.41344.68122.52953.08316.11245.02385.22841.77691.7781
H162.67094.08242.16275.00391.09302.93574.69184.26263.07952.50705.99625.22844.81211.78671.7781

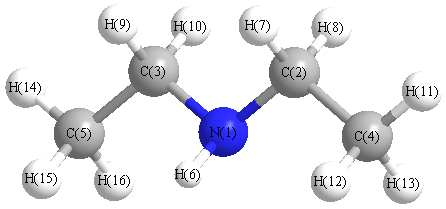
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 109.432 N1 C2 H7 113.156
N1 C2 H8 107.084 N1 C3 C5 109.432
N1 C3 H9 113.156 N1 C3 H10 107.084
C2 N1 C3 112.962 C2 N1 H6 109.258
C2 C4 H11 110.915 C2 C4 H12 110.369
C2 C4 H13 109.033 C3 N1 H6 109.258
C3 C5 H14 110.915 C3 C5 H15 110.369
C3 C5 H16 109.033 C4 C2 H7 110.013
C4 C2 H8 109.712 C5 C3 H8 149.677
C5 C3 H10 109.712 H7 C2 H8 107.347
H9 C3 H10 107.347 H11 C4 H12 108.371
H11 C4 H13 109.482 H12 C4 H13 108.633
H14 C5 H15 108.371 H14 C5 H16 109.482
H15 C5 H16 108.633
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability