return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Pentanenitrile)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-248.220507
Energy at 298.15K-248.230156
HF Energy-247.643326
Nuclear repulsion energy208.771657
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3158 3004 25.75      
2 A' 3097 2946 26.90      
3 A' 3088 2937 2.28      
4 A' 3083 2933 15.48      
5 A' 3076 2927 9.83      
6 A' 2143 2038 2.86      
7 A' 1607 1528 6.19      
8 A' 1594 1517 3.19      
9 A' 1585 1508 0.51      
10 A' 1572 1495 3.54      
11 A' 1496 1423 4.27      
12 A' 1435 1365 0.18      
13 A' 1415 1346 3.01      
14 A' 1347 1282 0.40      
15 A' 1160 1103 6.94      
16 A' 1050 999 0.10      
17 A' 1003 954 0.44      
18 A' 945 899 0.58      
19 A' 894 851 2.65      
20 A' 562 535 2.11      
21 A' 390 371 0.63      
22 A' 299 285 0.70      
23 A' 134 127 3.79      
24 A" 3161 3007 54.61      
25 A" 3150 2996 3.57      
26 A" 3128 2976 0.41      
27 A" 3112 2961 3.26      
28 A" 1603 1525 7.72      
29 A" 1397 1329 0.15      
30 A" 1385 1318 0.53      
31 A" 1316 1252 0.02      
32 A" 1209 1151 0.13      
33 A" 997 948 0.79      
34 A" 837 796 0.14      
35 A" 759 722 5.28      
36 A" 428 407 0.00      
37 A" 255 242 0.06      
38 A" 120 114 0.37      
39 A" 94 90 2.80      

Unscaled Zero Point Vibrational Energy (zpe) 29541.5 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 28102.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.49473 0.04340 0.04113

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.415 0.000
C2 1.449 0.961 0.000
C3 1.477 2.511 0.000
C4 -0.017 -1.143 0.000
N5 -2.498 -2.106 0.000
C6 -1.388 -1.683 0.000
H7 2.507 2.881 0.000
H8 -0.532 0.781 0.886
H9 -0.532 0.781 -0.886
H10 1.976 0.582 0.885
H11 1.976 0.582 -0.885
H12 0.968 2.900 -0.889
H13 0.968 2.900 0.889
H14 0.515 -1.512 -0.886
H15 0.515 -1.512 0.886

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.54812.56371.55833.54932.51563.51701.09651.09652.17152.17152.81102.81102.18272.1827
C21.54811.55012.56414.99853.87772.19292.17732.17731.09781.09782.18662.18662.78802.7880
C32.56371.55013.94736.09235.07881.09512.79512.79512.18002.18001.09551.09554.23004.2300
C41.55832.56413.94732.66181.47374.75052.18022.18022.78032.78034.25524.25521.09731.0973
N53.54934.99856.09232.66181.18827.06623.60383.60385.29415.29416.15366.15363.19653.1965
C62.51563.87775.07881.47371.18826.00052.75492.75494.15084.15085.22925.22922.10612.1061
H73.51702.19291.09514.75057.06626.00053.79923.79922.52052.52051.77751.77754.90484.9048
H81.09652.17732.79512.18023.60382.75493.79921.77272.51553.07683.14492.59623.08152.5207
H91.09652.17732.79512.18023.60382.75493.79921.77273.07682.51552.59623.14492.52073.0815
H102.17151.09782.18002.78035.29414.15082.52052.51553.07681.77063.08802.52763.10772.5533
H112.17151.09782.18002.78035.29414.15082.52053.07682.51551.77062.52763.08802.55333.1077
H122.81102.18661.09554.25526.15365.22921.77753.14492.59623.08802.52761.77724.43534.7773
H132.81102.18661.09554.25526.15365.22921.77752.59623.14492.52763.08801.77724.77734.4353
H142.18272.78804.23001.09733.19652.10614.90483.08152.52073.10772.55334.43534.77731.7725
H152.18272.78804.23001.09733.19652.10614.90482.52073.08152.55333.10774.77734.43531.7725

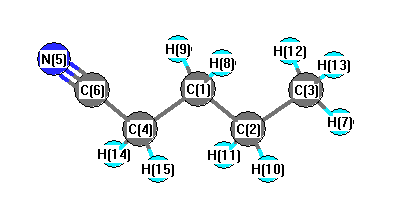
picture of Pentanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.683 C1 C2 H10 109.126
C1 C2 H11 109.126 C1 C4 C6 112.102
C1 C4 H14 109.327 C1 C4 H15 109.327
C2 C1 C4 111.257 C2 C1 H8 109.650
C2 C1 H9 109.650 C2 C3 H7 110.822
C2 C3 H12 110.300 C2 C3 H13 110.300
C3 C2 H10 109.652 C3 C2 H11 109.652
C4 C1 H8 109.176 C4 C1 H9 109.176
C4 C6 N5 179.376 C6 C4 H14 109.126
C6 C4 H15 109.126 H7 C3 H12 108.468
H7 C3 H13 108.468 H8 C1 H9 107.863
H10 C2 H11 107.504 H12 C3 H13 108.412
H14 C4 H15 107.734
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability