Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/3-21G*
Geometric Data calculated at MP2=FULL/3-21G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/3-21G*
| | hartrees |
|---|
| Energy at 0K | -249.371290 |
| Energy at 298.15K | -249.385666 |
| Nuclear repulsion energy | 255.861700 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/3-21G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3402 |
3225 |
3.33 |
|
|
|
| 2 |
A' |
3136 |
2973 |
75.81 |
|
|
|
| 3 |
A' |
3124 |
2962 |
20.36 |
|
|
|
| 4 |
A' |
3116 |
2954 |
21.05 |
|
|
|
| 5 |
A' |
3079 |
2919 |
29.87 |
|
|
|
| 6 |
A' |
3069 |
2910 |
16.93 |
|
|
|
| 7 |
A' |
3069 |
2909 |
21.63 |
|
|
|
| 8 |
A' |
1590 |
1507 |
4.85 |
|
|
|
| 9 |
A' |
1582 |
1500 |
10.90 |
|
|
|
| 10 |
A' |
1572 |
1490 |
2.81 |
|
|
|
| 11 |
A' |
1442 |
1367 |
0.17 |
|
|
|
| 12 |
A' |
1426 |
1352 |
0.43 |
|
|
|
| 13 |
A' |
1373 |
1301 |
1.25 |
|
|
|
| 14 |
A' |
1326 |
1257 |
5.01 |
|
|
|
| 15 |
A' |
1232 |
1168 |
4.53 |
|
|
|
| 16 |
A' |
1076 |
1020 |
3.29 |
|
|
|
| 17 |
A' |
1021 |
968 |
1.93 |
|
|
|
| 18 |
A' |
947 |
898 |
40.38 |
|
|
|
| 19 |
A' |
871 |
825 |
2.84 |
|
|
|
| 20 |
A' |
816 |
773 |
17.15 |
|
|
|
| 21 |
A' |
735 |
697 |
111.56 |
|
|
|
| 22 |
A' |
559 |
530 |
1.42 |
|
|
|
| 23 |
A' |
441 |
418 |
1.72 |
|
|
|
| 24 |
A' |
399 |
378 |
6.15 |
|
|
|
| 25 |
A' |
247 |
234 |
5.46 |
|
|
|
| 26 |
A" |
3133 |
2970 |
8.95 |
|
|
|
| 27 |
A" |
3117 |
2955 |
52.38 |
|
|
|
| 28 |
A" |
3077 |
2917 |
23.22 |
|
|
|
| 29 |
A" |
3068 |
2909 |
19.82 |
|
|
|
| 30 |
A" |
1575 |
1493 |
0.16 |
|
|
|
| 31 |
A" |
1569 |
1488 |
4.19 |
|
|
|
| 32 |
A" |
1525 |
1445 |
4.00 |
|
|
|
| 33 |
A" |
1427 |
1353 |
0.00 |
|
|
|
| 34 |
A" |
1419 |
1345 |
0.01 |
|
|
|
| 35 |
A" |
1413 |
1339 |
2.74 |
|
|
|
| 36 |
A" |
1353 |
1283 |
0.34 |
|
|
|
| 37 |
A" |
1245 |
1180 |
6.27 |
|
|
|
| 38 |
A" |
1174 |
1113 |
1.04 |
|
|
|
| 39 |
A" |
1085 |
1029 |
18.22 |
|
|
|
| 40 |
A" |
1080 |
1024 |
2.01 |
|
|
|
| 41 |
A" |
986 |
935 |
7.61 |
|
|
|
| 42 |
A" |
864 |
819 |
0.01 |
|
|
|
| 43 |
A" |
827 |
784 |
0.11 |
|
|
|
| 44 |
A" |
459 |
435 |
1.36 |
|
|
|
| 45 |
A" |
245 |
232 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35644.3 cm
-1
Scaled (by 0.948) Zero Point Vibrational Energy (zpe) 33790.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/3-21G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.676 |
1.321 |
0.000 |
| H2 |
0.643 |
2.417 |
0.000 |
| H3 |
1.732 |
1.017 |
0.000 |
| C4 |
-0.008 |
0.758 |
1.274 |
| C5 |
-0.008 |
0.758 |
-1.274 |
| C6 |
-0.008 |
-0.794 |
-1.235 |
| C7 |
-0.008 |
-0.794 |
1.235 |
| N8 |
-0.643 |
-1.360 |
0.000 |
| H9 |
-1.647 |
-1.135 |
0.000 |
| H10 |
0.506 |
1.112 |
2.176 |
| H11 |
0.506 |
1.112 |
-2.176 |
| H12 |
-1.047 |
1.114 |
1.313 |
| H13 |
-1.047 |
1.114 |
-1.313 |
| H14 |
1.029 |
-1.152 |
-1.253 |
| H15 |
1.029 |
-1.152 |
1.253 |
| H16 |
-0.528 |
-1.207 |
-2.107 |
| H17 |
-0.528 |
-1.207 |
2.107 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0968 | 1.0991 | 1.5516 | 1.5516 | 2.5432 | 2.5432 | 2.9878 | 3.3808 | 2.1930 | 2.1930 | 2.1761 | 2.1761 | 2.7950 | 2.7950 | 3.5043 | 3.5043 |
H2 | 1.0968 | | 1.7742 | 2.1909 | 2.1909 | 3.5019 | 3.5019 | 3.9900 | 4.2266 | 2.5417 | 2.5417 | 2.5056 | 2.5056 | 3.8028 | 3.8028 | 4.3527 | 4.3527 | H3 | 1.0991 | 1.7742 | | 2.1723 | 2.1723 | 2.7991 | 2.7991 | 3.3603 | 4.0064 | 2.4999 | 2.4999 | 3.0753 | 3.0753 | 2.6021 | 2.6021 | 3.8072 | 3.8072 | C4 | 1.5516 | 2.1909 | 2.1723 | | 2.5471 | 2.9499 | 1.5525 | 2.5512 | 2.8090 | 1.0975 | 3.5059 | 1.0988 | 2.8098 | 3.3331 | 2.1735 | 3.9444 | 2.1966 | C5 | 1.5516 | 2.1909 | 2.1723 | 2.5471 | | 1.5525 | 2.9499 | 2.5512 | 2.8090 | 3.5059 | 1.0975 | 2.8098 | 1.0988 | 2.1735 | 3.3331 | 2.1966 | 3.9444 | C6 | 2.5432 | 3.5019 | 2.7991 | 2.9499 | 1.5525 | | 2.4702 | 1.4995 | 2.0802 | 3.9414 | 2.1869 | 3.3485 | 2.1741 | 1.0972 | 2.7196 | 1.0958 | 3.4073 | C7 | 2.5432 | 3.5019 | 2.7991 | 1.5525 | 2.9499 | 2.4702 | | 1.4995 | 2.0802 | 2.1869 | 3.9414 | 2.1741 | 3.3485 | 2.7196 | 1.0972 | 3.4073 | 1.0958 | N8 | 2.9878 | 3.9900 | 3.3603 | 2.5512 | 2.5512 | 1.4995 | 1.4995 | | 1.0285 | 3.4877 | 3.4877 | 2.8294 | 2.8294 | 2.1000 | 2.1000 | 2.1156 | 2.1156 | H9 | 3.3808 | 4.2266 | 4.0064 | 2.8090 | 2.8090 | 2.0802 | 2.0802 | 1.0285 | | 3.7973 | 3.7973 | 2.6725 | 2.6725 | 2.9549 | 2.9549 | 2.3865 | 2.3865 | H10 | 2.1930 | 2.5417 | 2.4999 | 1.0975 | 3.5059 | 3.9414 | 2.1869 | 3.4877 | 3.7973 | | 4.3527 | 1.7768 | 3.8191 | 4.1427 | 2.5000 | 4.9791 | 2.5397 | H11 | 2.1930 | 2.5417 | 2.4999 | 3.5059 | 1.0975 | 2.1869 | 3.9414 | 3.4877 | 3.7973 | 4.3527 | | 3.8191 | 1.7768 | 2.5000 | 4.1427 | 2.5397 | 4.9791 | H12 | 2.1761 | 2.5056 | 3.0753 | 1.0988 | 2.8098 | 3.3485 | 2.1741 | 2.8294 | 2.6725 | 1.7768 | 3.8191 | | 2.6256 | 4.0038 | 3.0738 | 4.1654 | 2.5073 | H13 | 2.1761 | 2.5056 | 3.0753 | 2.8098 | 1.0988 | 2.1741 | 3.3485 | 2.8294 | 2.6725 | 3.8191 | 1.7768 | 2.6256 | | 3.0738 | 4.0038 | 2.5073 | 4.1654 | H14 | 2.7950 | 3.8028 | 2.6021 | 3.3331 | 2.1735 | 1.0972 | 2.7196 | 2.1000 | 2.9549 | 4.1427 | 2.5000 | 4.0038 | 3.0738 | | 2.5069 | 1.7765 | 3.7040 | H15 | 2.7950 | 3.8028 | 2.6021 | 2.1735 | 3.3331 | 2.7196 | 1.0972 | 2.1000 | 2.9549 | 2.5000 | 4.1427 | 3.0738 | 4.0038 | 2.5069 | | 3.7040 | 1.7765 | H16 | 3.5043 | 4.3527 | 3.8072 | 3.9444 | 2.1966 | 1.0958 | 3.4073 | 2.1156 | 2.3865 | 4.9791 | 2.5397 | 4.1654 | 2.5073 | 1.7765 | 3.7040 | | 4.2138 | H17 | 3.5043 | 4.3527 | 3.8072 | 2.1966 | 3.9444 | 3.4073 | 1.0958 | 2.1156 | 2.3865 | 2.5397 | 4.9791 | 2.5073 | 4.1654 | 3.7040 | 1.7765 | 4.2138 | |
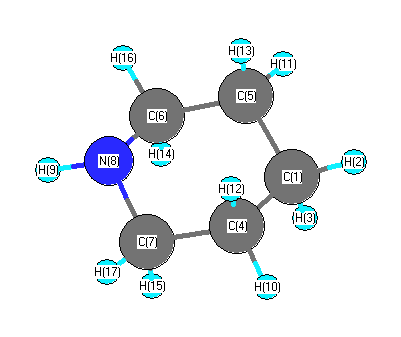 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.030 |
|
C1 |
C4 |
H10 |
110.577 |
| C1 |
C4 |
H12 |
109.185 |
|
C1 |
C5 |
C6 |
110.030 |
| C1 |
C5 |
H11 |
110.577 |
|
C1 |
C5 |
H13 |
109.185 |
| H2 |
C1 |
H3 |
107.792 |
|
H2 |
C1 |
C4 |
110.457 |
| H2 |
C1 |
C5 |
110.457 |
|
H3 |
C1 |
C4 |
108.872 |
| H3 |
C1 |
C5 |
108.872 |
|
C4 |
C1 |
C5 |
110.325 |
| C4 |
C7 |
N8 |
113.414 |
|
C4 |
C7 |
H15 |
109.010 |
| C4 |
C7 |
H17 |
110.901 |
|
C5 |
C6 |
N8 |
113.414 |
| C5 |
C6 |
H14 |
109.010 |
|
C5 |
C6 |
H16 |
110.901 |
| C6 |
C5 |
H11 |
110.039 |
|
C6 |
C5 |
H13 |
108.969 |
| C6 |
N8 |
C7 |
110.907 |
|
C6 |
N8 |
H9 |
109.328 |
| C7 |
C4 |
H10 |
110.039 |
|
C7 |
C4 |
H12 |
108.969 |
| C7 |
N8 |
H9 |
109.328 |
|
N8 |
C6 |
H14 |
106.922 |
| N8 |
C6 |
H16 |
108.196 |
|
N8 |
C7 |
H15 |
106.922 |
| N8 |
C7 |
H17 |
108.196 |
|
H10 |
C4 |
H12 |
107.993 |
| H11 |
C5 |
H13 |
107.993 |
|
H14 |
C6 |
H16 |
108.203 |
| H15 |
C7 |
H17 |
108.203 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
