return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for SiH2(CH3)2 (dimethylsilane)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-367.723531
Energy at 298.15K-367.731607
Nuclear repulsion energy113.369796
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3142 2989 13.37      
2 A1 3062 2913 2.89      
3 A1 2310 2198 117.44      
4 A1 1567 1490 3.27      
5 A1 1410 1341 15.78      
6 A1 1016 967 138.50      
7 A1 940 894 26.82      
8 A1 673 640 4.06      
9 A1 212 202 0.68      
10 A2 3138 2985 0.00      
11 A2 1558 1482 0.00      
12 A2 953 906 0.00      
13 A2 632 602 0.00      
14 A2 164 156 0.00      
15 B1 3139 2986 22.16      
16 B1 2309 2197 169.02      
17 B1 1570 1494 9.95      
18 B1 962 915 91.90      
19 B1 483 459 12.81      
20 B1 178 169 0.01      
21 B2 3142 2989 6.46      
22 B2 3063 2914 4.57      
23 B2 1561 1485 1.98      
24 B2 1406 1337 35.12      
25 B2 988 939 264.66      
26 B2 748 712 13.37      
27 B2 698 664 16.66      

Unscaled Zero Point Vibrational Energy (zpe) 20511.1 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 19512.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.55437 0.19356 0.16002

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.551
H2 -1.199 0.000 1.432
H3 1.199 0.000 1.432
C4 0.000 1.562 -0.513
C5 0.000 -1.562 -0.513
H6 0.000 2.464 0.111
H7 0.000 -2.464 0.111
H8 0.888 1.595 -1.158
H9 -0.888 1.595 -1.158
H10 -0.888 -1.595 -1.158
H11 0.888 -1.595 -1.158

Atom - Atom Distances (Å)
  Si1 H2 H3 C4 C5 H6 H7 H8 H9 H10 H11
Si11.48811.48811.88971.88972.50302.50302.50002.50002.50002.5000
H21.48812.39882.76772.76773.04213.04213.68843.05693.05693.6884
H31.48812.39882.76772.76773.04213.04213.05693.68843.68843.0569
C41.88972.76772.76773.12321.09724.07371.09761.09763.34143.3414
C51.88972.76772.76773.12324.07371.09723.34143.34141.09761.0976
H62.50303.04213.04211.09724.07374.92801.77581.77584.34394.3439
H72.50303.04213.04214.07371.09724.92804.34394.34391.77581.7758
H82.50003.68843.05691.09763.34141.77584.34391.77593.65033.1892
H92.50003.05693.68841.09763.34141.77584.34391.77593.18923.6503
H102.50003.05693.68843.34141.09764.34391.77583.65033.18921.7759
H112.50003.68843.05693.34141.09764.34391.77583.18923.65031.7759

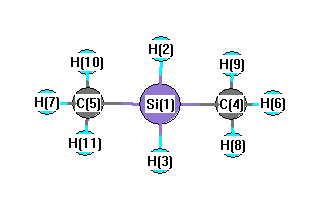
picture of dimethylsilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Si1 C4 H6 111.059 Si1 C4 H8 110.820
Si1 C4 H9 110.820 Si1 C5 H7 111.059
Si1 C5 H10 110.820 Si1 C5 H11 110.820
H2 Si1 H3 107.409 H2 Si1 C4 109.471
H2 Si1 C5 109.471 H3 Si1 C4 109.471
H3 Si1 C5 109.471 C4 Si1 C5 111.459
H6 C4 H8 108.013 H6 C4 H9 108.013
H7 C5 H10 108.013 H7 C5 H11 108.013
H8 C4 H9 107.991 H10 C5 H11 107.991
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability