return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Vinyl sulfoxide)

using model chemistry: CID/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/3-21G*
 hartrees
Energy at 0K-624.643144
Energy at 298.15K-624.649798
Nuclear repulsion energy292.021511
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3334 3103 0.91      
2 A' 3284 3055 3.85      
3 A' 3242 3016 1.91      
4 A' 1751 1629 1.35      
5 A' 1522 1416 18.48      
6 A' 1377 1282 0.43      
7 A' 1119 1041 15.28      
8 A' 1091 1015 93.93      
9 A' 1081 1006 33.08      
10 A' 1065 991 8.60      
11 A' 782 728 99.64      
12 A' 680 633 0.70      
13 A' 554 516 2.08      
14 A' 343 319 3.60      
15 A' 238 222 3.64      
16 A' 102 95 0.62      
17 A" 3334 3102 2.58      
18 A" 3279 3051 3.68      
19 A" 3240 3015 3.14      
20 A" 1741 1620 1.09      
21 A" 1518 1412 8.86      
22 A" 1362 1267 7.40      
23 A" 1093 1017 5.09      
24 A" 1081 1006 52.90      
25 A" 1059 986 13.35      
26 A" 695 647 27.74      
27 A" 631 587 7.51      
28 A" 527 490 6.39      
29 A" 289 269 10.91      
30 A" 178 165 0.81      

Unscaled Zero Point Vibrational Energy (zpe) 20795.3 cm-1
Scaled (by 0.9305) Zero Point Vibrational Energy (zpe) 19350.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/3-21G*
ABC
0.16284 0.07727 0.06916

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/3-21G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.581 -0.546 0.000
O2 1.307 0.773 0.000
C3 -0.616 -0.480 1.321
C4 -0.616 -0.480 -1.321
C5 -0.616 0.574 2.121
C6 -0.616 0.574 -2.121
H7 -1.267 -1.337 1.427
H8 -1.267 -1.337 -1.427
H9 -1.293 0.664 2.958
H10 -1.293 0.664 -2.958
H11 0.083 1.380 1.941
H12 0.083 1.380 -1.941

Atom - Atom Distances (Å)
  S1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
S11.50521.78391.78392.68062.68062.46462.46463.70443.70442.77872.7787
O21.50522.64822.64822.87012.87013.62063.62063.93943.93942.37342.3734
C31.78392.64822.64201.32313.59991.08092.95062.10844.48052.08103.8191
C41.78392.64822.64203.59991.32312.95061.08094.48052.10843.81912.0810
C52.68062.87011.32313.59994.24252.13444.08171.07975.12471.08194.1997
C62.68062.87013.59991.32314.24254.08172.13445.12471.07974.19971.0819
H72.46463.62061.08092.95062.13444.08172.85312.51964.81933.07634.5319
H82.46463.62062.95061.08094.08172.13442.85314.81932.51964.53193.0763
H93.70443.93942.10844.48051.07975.12472.51964.81935.91551.85435.1380
H103.70443.93944.48052.10845.12471.07974.81932.51965.91555.13801.8543
H112.77872.37342.08103.81911.08194.19973.07634.53191.85435.13803.8814
H122.77872.37343.81912.08104.19971.08194.53193.07635.13801.85433.8814

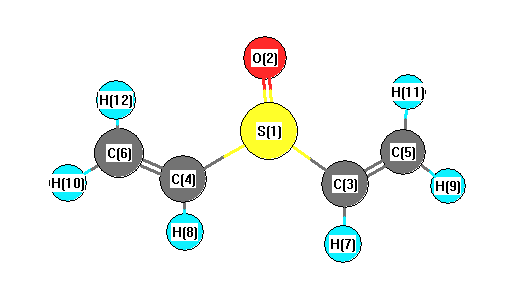
picture of Vinyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 118.504 S1 C3 H7 116.546
S1 C4 C6 118.504 S1 C4 H8 116.546
O2 S1 C3 106.941 O2 S1 C4 106.941
C3 S1 C4 95.552 C3 C5 H9 122.359
C3 C5 H11 119.487 C4 C6 H10 122.359
C4 C6 H12 119.487 C5 C3 H7 124.903
C6 C4 H8 124.903 H9 C5 H11 118.154
H10 C6 H12 118.154
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability