return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6S (Thiophene, 2,3-dihydro-)

using model chemistry: MP2/3-21G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/3-21G*
 hartrees
Energy at 0K-550.229736
Energy at 298.15K-550.236641
HF Energy-549.748654
Nuclear repulsion energy214.860575
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/3-21G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3263 3104 6.54      
2 A 3230 3073 0.57      
3 A 3188 3033 7.98      
4 A 3142 2989 16.15      
5 A 3121 2969 11.05      
6 A 3080 2930 20.81      
7 A 1611 1532 5.00      
8 A 1577 1500 2.46      
9 A 1564 1488 1.58      
10 A 1387 1319 1.32      
11 A 1371 1304 3.42      
12 A 1349 1284 16.90      
13 A 1253 1192 1.00      
14 A 1209 1150 0.19      
15 A 1188 1130 1.14      
16 A 1055 1004 3.85      
17 A 982 934 5.05      
18 A 938 893 1.39      
19 A 912 868 2.50      
20 A 876 833 1.05      
21 A 799 760 17.95      
22 A 711 676 39.14      
23 A 693 659 28.65      
24 A 659 627 3.96      
25 A 519 493 0.70      
26 A 407 387 2.85      
27 A 148 141 1.31      

Unscaled Zero Point Vibrational Energy (zpe) 20115.0 cm-1
Scaled (by 0.9513) Zero Point Vibrational Energy (zpe) 19135.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/3-21G*
ABC
0.22514 0.15286 0.09590

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/3-21G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.064 -1.276 0.201
H2 -0.058 -1.550 1.259
H3 0.125 -2.159 -0.413
C4 -1.413 -0.574 -0.169
H5 -2.223 -0.969 0.453
H6 -1.660 -0.757 -1.224
C7 0.115 1.287 0.076
H8 0.488 2.300 0.166
C9 -1.181 0.921 0.057
H10 -2.003 1.625 0.118
S11 1.287 -0.040 -0.085

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 C9 H10 S11
C11.09301.09171.56522.19562.20152.57263.61842.46933.49091.8533
H21.09301.78892.19722.38253.05933.07854.03882.96863.89442.4281
H31.09171.78892.22152.77102.40993.48084.51103.37844.37392.4394
C41.56522.19722.22151.09491.09842.42043.46211.53042.29552.7532
H52.19562.38252.77101.09491.78143.27034.25622.19442.62483.6702
H62.20153.05932.40991.09841.78143.00263.98582.16472.75573.2390
C72.57263.07853.48082.42043.27033.00261.08301.34602.14471.7774
H83.61844.03884.51103.46214.25623.98581.08302.16722.58112.4844
C92.46932.96863.37841.53042.19442.16471.34602.16721.08422.6514
H103.49093.89444.37392.29552.62482.75572.14472.58111.08423.6923
S111.85332.42812.43942.75323.67023.23901.77742.48442.65143.6923

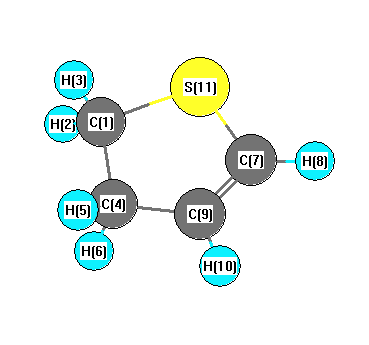
picture of Thiophene, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 109.998 C1 C4 H6 110.250
C1 C4 C9 105.816 C1 S11 C7 90.210
H2 C1 H3 109.940 H2 C1 C4 110.232
H2 C1 S11 108.211 H3 C1 C4 112.239
H3 C1 S11 109.099 C4 C1 S11 106.992
C4 C9 C7 114.441 C4 C9 H10 121.862
H5 C4 H6 108.627 H5 C4 C9 112.359
H6 C4 C9 109.772 C7 C9 H10 123.538
H8 C7 C9 125.956 H8 C7 S11 118.549
C9 C7 S11 115.489
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability